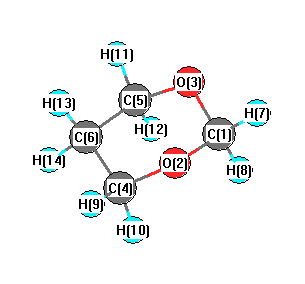
Vibrational Frequencies calculated at HF/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3406 |
3065 |
34.59 |
|
|
|
| 2 |
A' |
3344 |
3009 |
41.41 |
|
|
|
| 3 |
A' |
3295 |
2965 |
58.45 |
|
|
|
| 4 |
A' |
3233 |
2910 |
43.59 |
|
|
|
| 5 |
A' |
3209 |
2888 |
202.39 |
|
|
|
| 6 |
A' |
3196 |
2876 |
38.94 |
|
|
|
| 7 |
A' |
1671 |
1504 |
6.93 |
|
|
|
| 8 |
A' |
1657 |
1491 |
0.30 |
|
|
|
| 9 |
A' |
1630 |
1466 |
7.60 |
|
|
|
| 10 |
A' |
1539 |
1385 |
19.87 |
|
|
|
| 11 |
A' |
1421 |
1279 |
6.38 |
|
|
|
| 12 |
A' |
1310 |
1179 |
53.49 |
|
|
|
| 13 |
A' |
1281 |
1153 |
168.94 |
|
|
|
| 14 |
A' |
1198 |
1078 |
34.32 |
|
|
|
| 15 |
A' |
1073 |
966 |
57.50 |
|
|
|
| 16 |
A' |
971 |
874 |
6.64 |
|
|
|
| 17 |
A' |
888 |
799 |
17.89 |
|
|
|
| 18 |
A' |
691 |
622 |
10.11 |
|
|
|
| 19 |
A' |
520 |
468 |
1.47 |
|
|
|
| 20 |
A' |
405 |
364 |
19.83 |
|
|
|
| 21 |
A' |
259 |
233 |
6.34 |
|
|
|
| 22 |
A" |
3340 |
3006 |
53.39 |
|
|
|
| 23 |
A" |
3200 |
2880 |
31.90 |
|
|
|
| 24 |
A" |
1653 |
1488 |
3.91 |
|
|
|
| 25 |
A" |
1565 |
1408 |
11.14 |
|
|
|
| 26 |
A" |
1523 |
1371 |
2.22 |
|
|
|
| 27 |
A" |
1510 |
1359 |
0.34 |
|
|
|
| 28 |
A" |
1425 |
1283 |
1.30 |
|
|
|
| 29 |
A" |
1367 |
1231 |
36.49 |
|
|
|
| 30 |
A" |
1333 |
1200 |
4.56 |
|
|
|
| 31 |
A" |
1187 |
1068 |
57.99 |
|
|
|
| 32 |
A" |
1124 |
1012 |
79.54 |
|
|
|
| 33 |
A" |
996 |
896 |
17.08 |
|
|
|
| 34 |
A" |
938 |
844 |
21.60 |
|
|
|
| 35 |
A" |
480 |
432 |
13.95 |
|
|
|
| 36 |
A" |
245 |
221 |
3.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29042.3 cm
-1
Scaled (by 0.8999) Zero Point Vibrational Energy (zpe) 26135.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.089 |
|
|
|
| 2 |
O |
-0.484 |
|
|
|
| 3 |
O |
-0.484 |
|
|
|
| 4 |
C |
-0.121 |
|
|
|
| 5 |
C |
-0.121 |
|
|
|
| 6 |
C |
-0.308 |
|
|
|
| 7 |
H |
0.206 |
|
|
|
| 8 |
H |
0.157 |
|
|
|
| 9 |
H |
0.204 |
|
|
|
| 10 |
H |
0.148 |
|
|
|
| 11 |
H |
0.204 |
|
|
|
| 12 |
H |
0.148 |
|
|
|
| 13 |
H |
0.197 |
|
|
|
| 14 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.261 |
2.824 |
0.000 |
3.093 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.676 |
2.255 |
0.000 |
| y |
2.255 |
-35.006 |
0.000 |
| z |
0.000 |
0.000 |
-41.179 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.417 |
2.255 |
0.000 |
| y |
2.255 |
3.421 |
0.000 |
| z |
0.000 |
0.000 |
-5.838 |
|
| Polar |
|---|
| 3z2-r2 | -11.675 |
|---|
| x2-y2 | -0.669 |
|---|
| xy | 2.255 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.683 |
0.378 |
0.000 |
| y |
0.378 |
6.761 |
0.000 |
| z |
0.000 |
0.000 |
6.202 |
<r2> (average value of r
2) Å
2
| <r2> |
142.239 |
| (<r2>)1/2 |
11.926 |