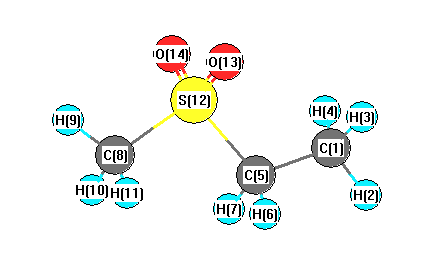
Vibrational Frequencies calculated at HF/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3412 |
3071 |
2.02 |
|
|
|
| 2 |
A |
3403 |
3063 |
0.50 |
|
|
|
| 3 |
A |
3373 |
3036 |
3.77 |
|
|
|
| 4 |
A |
3333 |
2999 |
6.95 |
|
|
|
| 5 |
A |
3302 |
2972 |
32.46 |
|
|
|
| 6 |
A |
3290 |
2961 |
6.51 |
|
|
|
| 7 |
A |
3270 |
2943 |
6.69 |
|
|
|
| 8 |
A |
3221 |
2898 |
27.88 |
|
|
|
| 9 |
A |
1656 |
1490 |
10.70 |
|
|
|
| 10 |
A |
1643 |
1478 |
14.25 |
|
|
|
| 11 |
A |
1618 |
1456 |
30.15 |
|
|
|
| 12 |
A |
1612 |
1451 |
4.24 |
|
|
|
| 13 |
A |
1595 |
1435 |
10.20 |
|
|
|
| 14 |
A |
1580 |
1422 |
10.61 |
|
|
|
| 15 |
A |
1531 |
1378 |
0.56 |
|
|
|
| 16 |
A |
1468 |
1321 |
14.77 |
|
|
|
| 17 |
A |
1398 |
1258 |
1.80 |
|
|
|
| 18 |
A |
1236 |
1112 |
10.08 |
|
|
|
| 19 |
A |
1194 |
1074 |
18.32 |
|
|
|
| 20 |
A |
1152 |
1036 |
6.60 |
|
|
|
| 21 |
A |
1135 |
1021 |
13.38 |
|
|
|
| 22 |
A |
1075 |
968 |
7.98 |
|
|
|
| 23 |
A |
876 |
788 |
12.07 |
|
|
|
| 24 |
A |
828 |
745 |
1.19 |
|
|
|
| 25 |
A |
771 |
694 |
2.05 |
|
|
|
| 26 |
A |
717 |
645 |
2.21 |
|
|
|
| 27 |
A |
586 |
528 |
3.45 |
|
|
|
| 28 |
A |
409 |
368 |
5.72 |
|
|
|
| 29 |
A |
320 |
288 |
49.61 |
|
|
|
| 30 |
A |
302 |
272 |
43.92 |
|
|
|
| 31 |
A |
268 |
241 |
7.12 |
|
|
|
| 32 |
A |
254 |
228 |
2.58 |
|
|
|
| 33 |
A |
208 |
187 |
4.57 |
|
|
|
| 34 |
A |
185 |
167 |
1.56 |
|
|
|
| 35 |
A |
166 |
150 |
0.23 |
|
|
|
| 36 |
A |
76 |
68 |
4.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26230.2 cm
-1
Scaled (by 0.8999) Zero Point Vibrational Energy (zpe) 23604.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.521 |
|
|
|
| 2 |
H |
0.245 |
|
|
|
| 3 |
H |
0.205 |
|
|
|
| 4 |
H |
0.174 |
|
|
|
| 5 |
C |
-0.464 |
|
|
|
| 6 |
H |
0.279 |
|
|
|
| 7 |
H |
0.231 |
|
|
|
| 8 |
C |
-0.644 |
|
|
|
| 9 |
H |
0.235 |
|
|
|
| 10 |
H |
0.282 |
|
|
|
| 11 |
H |
0.239 |
|
|
|
| 12 |
S |
1.185 |
|
|
|
| 13 |
O |
-0.728 |
|
|
|
| 14 |
O |
-0.720 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.619 |
3.212 |
-4.693 |
7.326 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.610 |
-8.947 |
0.562 |
| y |
-8.947 |
-44.132 |
-3.453 |
| z |
0.562 |
-3.453 |
-51.429 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.170 |
-8.947 |
0.562 |
| y |
-8.947 |
4.888 |
-3.453 |
| z |
0.562 |
-3.453 |
-6.058 |
|
| Polar |
|---|
| 3z2-r2 | -12.115 |
|---|
| x2-y2 | -2.479 |
|---|
| xy | -8.947 |
|---|
| xz | 0.562 |
|---|
| yz | -3.453 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.848 |
0.140 |
0.346 |
| y |
0.140 |
8.014 |
0.337 |
| z |
0.346 |
0.337 |
8.024 |
<r2> (average value of r
2) Å
2
| <r2> |
205.471 |
| (<r2>)1/2 |
14.334 |