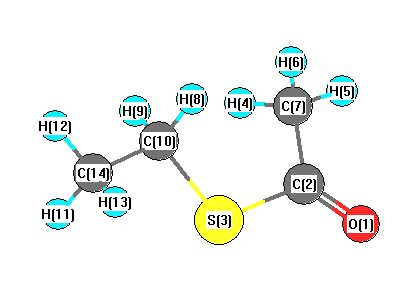
Vibrational Frequencies calculated at HF/LANL2DZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3350 |
3015 |
47.51 |
|
|
|
| 2 |
A |
3344 |
3009 |
14.74 |
|
|
|
| 3 |
A |
3314 |
2983 |
9.47 |
|
|
|
| 4 |
A |
3306 |
2975 |
24.12 |
|
|
|
| 5 |
A |
3297 |
2967 |
53.63 |
|
|
|
| 6 |
A |
3274 |
2946 |
20.05 |
|
|
|
| 7 |
A |
3226 |
2903 |
6.78 |
|
|
|
| 8 |
A |
3210 |
2889 |
46.22 |
|
|
|
| 9 |
A |
1849 |
1664 |
412.35 |
|
|
|
| 10 |
A |
1650 |
1485 |
6.23 |
|
|
|
| 11 |
A |
1641 |
1477 |
13.16 |
|
|
|
| 12 |
A |
1639 |
1475 |
1.00 |
|
|
|
| 13 |
A |
1633 |
1469 |
22.04 |
|
|
|
| 14 |
A |
1619 |
1457 |
37.21 |
|
|
|
| 15 |
A |
1576 |
1418 |
9.15 |
|
|
|
| 16 |
A |
1561 |
1405 |
20.60 |
|
|
|
| 17 |
A |
1455 |
1309 |
36.04 |
|
|
|
| 18 |
A |
1404 |
1264 |
0.09 |
|
|
|
| 19 |
A |
1266 |
1139 |
135.55 |
|
|
|
| 20 |
A |
1187 |
1068 |
4.46 |
|
|
|
| 21 |
A |
1172 |
1055 |
0.01 |
|
|
|
| 22 |
A |
1156 |
1040 |
3.73 |
|
|
|
| 23 |
A |
1077 |
969 |
11.05 |
|
|
|
| 24 |
A |
1051 |
946 |
46.68 |
|
|
|
| 25 |
A |
878 |
790 |
6.53 |
|
|
|
| 26 |
A |
707 |
636 |
2.98 |
|
|
|
| 27 |
A |
666 |
599 |
70.90 |
|
|
|
| 28 |
A |
556 |
500 |
7.83 |
|
|
|
| 29 |
A |
488 |
439 |
6.28 |
|
|
|
| 30 |
A |
392 |
353 |
3.72 |
|
|
|
| 31 |
A |
323 |
291 |
0.74 |
|
|
|
| 32 |
A |
254 |
229 |
0.11 |
|
|
|
| 33 |
A |
169 |
152 |
0.00 |
|
|
|
| 34 |
A |
169 |
152 |
1.59 |
|
|
|
| 35 |
A |
66 |
60 |
0.52 |
|
|
|
| 36 |
A |
55 |
49 |
0.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26989.8 cm
-1
Scaled (by 0.8999) Zero Point Vibrational Energy (zpe) 24288.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/LANL2DZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.339 |
|
|
|
| 2 |
C |
0.136 |
|
|
|
| 3 |
S |
0.173 |
|
|
|
| 4 |
H |
0.202 |
|
|
|
| 5 |
H |
0.236 |
|
|
|
| 6 |
H |
0.202 |
|
|
|
| 7 |
C |
-0.576 |
|
|
|
| 8 |
H |
0.217 |
|
|
|
| 9 |
H |
0.217 |
|
|
|
| 10 |
C |
-0.517 |
|
|
|
| 11 |
H |
0.195 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
| 13 |
H |
0.195 |
|
|
|
| 14 |
C |
-0.525 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.221 |
4.520 |
0.000 |
5.550 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.488 |
5.522 |
-0.002 |
| y |
5.522 |
-47.696 |
0.000 |
| z |
-0.002 |
0.000 |
-44.284 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.498 |
5.522 |
-0.002 |
| y |
5.522 |
-1.310 |
0.000 |
| z |
-0.002 |
0.000 |
3.807 |
|
| Polar |
|---|
| 3z2-r2 | 7.615 |
|---|
| x2-y2 | -0.792 |
|---|
| xy | 5.522 |
|---|
| xz | -0.002 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.077 |
-0.712 |
0.001 |
| y |
-0.712 |
8.229 |
0.001 |
| z |
0.001 |
0.001 |
5.469 |
<r2> (average value of r
2) Å
2
| <r2> |
249.635 |
| (<r2>)1/2 |
15.800 |