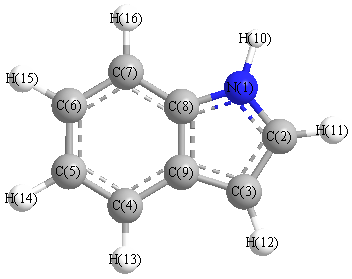
Vibrational Frequencies calculated at HF/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3947 |
3554 |
93.29 |
|
|
|
| 2 |
A' |
3485 |
3138 |
10.45 |
|
|
|
| 3 |
A' |
3454 |
3110 |
3.06 |
|
|
|
| 4 |
A' |
3414 |
3074 |
37.39 |
|
|
|
| 5 |
A' |
3397 |
3059 |
51.48 |
|
|
|
| 6 |
A' |
3383 |
3046 |
6.29 |
|
|
|
| 7 |
A' |
3375 |
3038 |
1.39 |
|
|
|
| 8 |
A' |
1810 |
1630 |
3.95 |
|
|
|
| 9 |
A' |
1773 |
1596 |
1.31 |
|
|
|
| 10 |
A' |
1691 |
1523 |
18.65 |
|
|
|
| 11 |
A' |
1651 |
1487 |
2.61 |
|
|
|
| 12 |
A' |
1616 |
1455 |
48.39 |
|
|
|
| 13 |
A' |
1562 |
1407 |
41.71 |
|
|
|
| 14 |
A' |
1508 |
1358 |
2.46 |
|
|
|
| 15 |
A' |
1423 |
1282 |
16.36 |
|
|
|
| 16 |
A' |
1402 |
1262 |
23.81 |
|
|
|
| 17 |
A' |
1387 |
1249 |
9.84 |
|
|
|
| 18 |
A' |
1330 |
1198 |
5.50 |
|
|
|
| 19 |
A' |
1280 |
1152 |
4.21 |
|
|
|
| 20 |
A' |
1243 |
1119 |
4.75 |
|
|
|
| 21 |
A' |
1207 |
1086 |
28.44 |
|
|
|
| 22 |
A' |
1170 |
1054 |
36.51 |
|
|
|
| 23 |
A' |
1106 |
996 |
7.53 |
|
|
|
| 24 |
A' |
988 |
890 |
7.50 |
|
|
|
| 25 |
A' |
959 |
864 |
1.09 |
|
|
|
| 26 |
A' |
824 |
742 |
3.46 |
|
|
|
| 27 |
A' |
672 |
605 |
1.60 |
|
|
|
| 28 |
A' |
593 |
534 |
0.19 |
|
|
|
| 29 |
A' |
434 |
391 |
5.63 |
|
|
|
| 30 |
A" |
1137 |
1024 |
0.00 |
|
|
|
| 31 |
A" |
1100 |
991 |
6.16 |
|
|
|
| 32 |
A" |
1042 |
938 |
1.32 |
|
|
|
| 33 |
A" |
996 |
897 |
0.46 |
|
|
|
| 34 |
A" |
898 |
808 |
79.70 |
|
|
|
| 35 |
A" |
869 |
783 |
137.04 |
|
|
|
| 36 |
A" |
841 |
757 |
9.58 |
|
|
|
| 37 |
A" |
672 |
605 |
5.85 |
|
|
|
| 38 |
A" |
644 |
580 |
0.00 |
|
|
|
| 39 |
A" |
569 |
513 |
141.60 |
|
|
|
| 40 |
A" |
481 |
433 |
18.40 |
|
|
|
| 41 |
A" |
275 |
248 |
0.48 |
|
|
|
| 42 |
A" |
239 |
216 |
14.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30923.0 cm
-1
Scaled (by 0.9004) Zero Point Vibrational Energy (zpe) 27843.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.655 |
|
|
|
| 2 |
C |
-0.110 |
|
|
|
| 3 |
C |
-0.374 |
|
|
|
| 4 |
C |
-0.344 |
|
|
|
| 5 |
C |
-0.251 |
|
|
|
| 6 |
C |
-0.194 |
|
|
|
| 7 |
C |
-0.322 |
|
|
|
| 8 |
C |
0.307 |
|
|
|
| 9 |
C |
0.258 |
|
|
|
| 10 |
H |
0.371 |
|
|
|
| 11 |
H |
0.230 |
|
|
|
| 12 |
H |
0.234 |
|
|
|
| 13 |
H |
0.219 |
|
|
|
| 14 |
H |
0.211 |
|
|
|
| 15 |
H |
0.209 |
|
|
|
| 16 |
H |
0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.760 |
1.855 |
0.000 |
2.005 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.021 |
-2.100 |
0.000 |
| y |
-2.100 |
-42.618 |
0.000 |
| z |
0.000 |
0.000 |
-60.448 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.512 |
-2.100 |
0.000 |
| y |
-2.100 |
11.117 |
0.000 |
| z |
0.000 |
0.000 |
-15.629 |
|
| Polar |
|---|
| 3z2-r2 | -31.258 |
|---|
| x2-y2 | -4.403 |
|---|
| xy | -2.100 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
16.252 |
-1.424 |
0.000 |
| y |
-1.424 |
13.946 |
0.000 |
| z |
0.000 |
0.000 |
5.382 |
<r2> (average value of r
2) Å
2
| <r2> |
282.829 |
| (<r2>)1/2 |
16.818 |