Jump to
S1C2
Energy calculated at HF/SDD
| | hartrees |
|---|
| Energy at 0K | -133.029081 |
| Energy at 298.15K | -133.034658 |
| HF Energy | -133.029081 |
| Nuclear repulsion energy | 70.895576 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3716 |
3346 |
4.29 |
|
|
|
| 2 |
A' |
3333 |
3001 |
27.71 |
|
|
|
| 3 |
A' |
3246 |
2923 |
67.65 |
|
|
|
| 4 |
A' |
3197 |
2879 |
28.74 |
|
|
|
| 5 |
A' |
1863 |
1677 |
72.92 |
|
|
|
| 6 |
A' |
1626 |
1464 |
17.08 |
|
|
|
| 7 |
A' |
1569 |
1413 |
15.46 |
|
|
|
| 8 |
A' |
1537 |
1384 |
38.93 |
|
|
|
| 9 |
A' |
1373 |
1236 |
44.50 |
|
|
|
| 10 |
A' |
1162 |
1046 |
37.06 |
|
|
|
| 11 |
A' |
1002 |
903 |
9.08 |
|
|
|
| 12 |
A' |
521 |
469 |
24.07 |
|
|
|
| 13 |
A" |
3267 |
2942 |
44.84 |
|
|
|
| 14 |
A" |
1632 |
1469 |
16.42 |
|
|
|
| 15 |
A" |
1257 |
1131 |
5.27 |
|
|
|
| 16 |
A" |
1195 |
1076 |
23.36 |
|
|
|
| 17 |
A" |
762 |
686 |
102.73 |
|
|
|
| 18 |
A" |
184 |
166 |
3.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16220.4 cm
-1
Scaled (by 0.9004) Zero Point Vibrational Energy (zpe) 14604.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/SDD
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.045 |
-0.628 |
0.000 |
| C2 |
0.000 |
0.451 |
0.000 |
| N3 |
1.245 |
0.184 |
0.000 |
| H4 |
-0.576 |
-1.601 |
0.000 |
| H5 |
-1.681 |
-0.539 |
0.875 |
| H6 |
-1.681 |
-0.539 |
-0.875 |
| H7 |
-0.376 |
1.469 |
0.000 |
| H8 |
1.864 |
0.981 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5023 | 2.4300 | 1.0797 | 1.0857 | 1.0857 | 2.2017 | 3.3242 |
C2 | 1.5023 | | 1.2738 | 2.1318 | 2.1387 | 2.1387 | 1.0853 | 1.9377 | N3 | 2.4300 | 1.2738 | | 2.5507 | 3.1393 | 3.1393 | 2.0690 | 1.0088 | H4 | 1.0797 | 2.1318 | 2.5507 | | 1.7646 | 1.7646 | 3.0770 | 3.5528 | H5 | 1.0857 | 2.1387 | 3.1393 | 1.7646 | | 1.7498 | 2.5504 | 3.9556 | H6 | 1.0857 | 2.1387 | 3.1393 | 1.7646 | 1.7498 | | 2.5504 | 3.9556 | H7 | 2.2017 | 1.0853 | 2.0690 | 3.0770 | 2.5504 | 2.5504 | | 2.2925 | H8 | 3.3242 | 1.9377 | 1.0088 | 3.5528 | 3.9556 | 3.9556 | 2.2925 | |
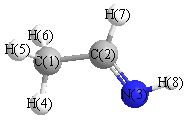 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.953 |
|
C1 |
C2 |
H7 |
115.671 |
| C2 |
C1 |
H4 |
110.229 |
|
C2 |
C1 |
H5 |
110.424 |
| C2 |
C1 |
H6 |
110.424 |
|
C2 |
N3 |
H8 |
115.704 |
| N3 |
C2 |
H7 |
122.376 |
|
H4 |
C1 |
H5 |
109.158 |
| H4 |
C1 |
H6 |
109.158 |
|
H5 |
C1 |
H6 |
107.386 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.577 |
|
|
|
| 2 |
C |
-0.024 |
|
|
|
| 3 |
N |
-0.413 |
|
|
|
| 4 |
H |
0.221 |
|
|
|
| 5 |
H |
0.187 |
|
|
|
| 6 |
H |
0.187 |
|
|
|
| 7 |
H |
0.160 |
|
|
|
| 8 |
H |
0.259 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.694 |
1.919 |
0.000 |
2.560 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.014 |
3.106 |
0.000 |
| y |
3.106 |
-17.726 |
0.000 |
| z |
0.000 |
0.000 |
-20.211 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.045 |
3.106 |
0.000 |
| y |
3.106 |
2.386 |
0.000 |
| z |
0.000 |
0.000 |
-1.341 |
|
| Polar |
|---|
| 3z2-r2 | -2.682 |
|---|
| x2-y2 | -2.288 |
|---|
| xy | 3.106 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.665 |
0.127 |
0.000 |
| y |
0.127 |
3.569 |
0.000 |
| z |
0.000 |
0.000 |
2.638 |
<r2> (average value of r
2) Å
2
| <r2> |
50.699 |
| (<r2>)1/2 |
7.120 |
Jump to
S1C1
Energy calculated at HF/SDD
| | hartrees |
|---|
| Energy at 0K | -133.028378 |
| Energy at 298.15K | -133.033944 |
| HF Energy | -133.028378 |
| Nuclear repulsion energy | 70.825878 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3690 |
3322 |
4.76 |
|
|
|
| 2 |
A' |
3346 |
3013 |
56.26 |
|
|
|
| 3 |
A' |
3294 |
2966 |
21.92 |
|
|
|
| 4 |
A' |
3198 |
2879 |
21.67 |
|
|
|
| 5 |
A' |
1849 |
1665 |
75.53 |
|
|
|
| 6 |
A' |
1633 |
1470 |
32.60 |
|
|
|
| 7 |
A' |
1568 |
1412 |
17.04 |
|
|
|
| 8 |
A' |
1536 |
1383 |
3.89 |
|
|
|
| 9 |
A' |
1377 |
1240 |
91.15 |
|
|
|
| 10 |
A' |
1162 |
1046 |
41.03 |
|
|
|
| 11 |
A' |
989 |
891 |
0.59 |
|
|
|
| 12 |
A' |
519 |
467 |
11.19 |
|
|
|
| 13 |
A" |
3278 |
2951 |
34.61 |
|
|
|
| 14 |
A" |
1626 |
1464 |
15.08 |
|
|
|
| 15 |
A" |
1261 |
1136 |
57.59 |
|
|
|
| 16 |
A" |
1201 |
1081 |
28.04 |
|
|
|
| 17 |
A" |
778 |
701 |
18.49 |
|
|
|
| 18 |
A" |
174 |
157 |
0.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16239.3 cm
-1
Scaled (by 0.9004) Zero Point Vibrational Energy (zpe) 14621.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/SDD
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.032 |
-0.616 |
0.000 |
| C2 |
0.000 |
0.487 |
0.000 |
| N3 |
1.267 |
0.362 |
0.000 |
| H4 |
-0.571 |
-1.597 |
0.000 |
| H5 |
-1.668 |
-0.533 |
0.875 |
| H6 |
-1.668 |
-0.533 |
-0.875 |
| H7 |
-0.384 |
1.493 |
0.000 |
| H8 |
1.612 |
-0.588 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5104 | 2.4988 | 1.0834 | 1.0848 | 1.0848 | 2.2068 | 2.6440 |
C2 | 1.5104 | | 1.2734 | 2.1606 | 2.1416 | 2.1416 | 1.0773 | 1.9372 | N3 | 2.4988 | 1.2734 | | 2.6869 | 3.1910 | 3.1910 | 2.0014 | 1.0105 | H4 | 1.0834 | 2.1606 | 2.6869 | | 1.7610 | 1.7610 | 3.0959 | 2.4054 | H5 | 1.0848 | 2.1416 | 3.1910 | 1.7610 | | 1.7493 | 2.5530 | 3.3952 | H6 | 1.0848 | 2.1416 | 3.1910 | 1.7610 | 1.7493 | | 2.5530 | 3.3952 | H7 | 2.2068 | 1.0773 | 2.0014 | 3.0959 | 2.5530 | 2.5530 | | 2.8833 | H8 | 2.6440 | 1.9372 | 1.0105 | 2.4054 | 3.3952 | 3.3952 | 2.8833 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.486 |
|
C1 |
C2 |
H7 |
116.028 |
| C2 |
C1 |
H4 |
111.751 |
|
C2 |
C1 |
H5 |
110.137 |
| C2 |
C1 |
H6 |
110.137 |
|
C2 |
N3 |
H8 |
115.549 |
| N3 |
C2 |
H7 |
116.486 |
|
H4 |
C1 |
H5 |
108.618 |
| H4 |
C1 |
H6 |
108.618 |
|
H5 |
C1 |
H6 |
107.464 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.598 |
|
|
|
| 2 |
C |
-0.024 |
|
|
|
| 3 |
N |
-0.400 |
|
|
|
| 4 |
H |
0.187 |
|
|
|
| 5 |
H |
0.202 |
|
|
|
| 6 |
H |
0.202 |
|
|
|
| 7 |
H |
0.185 |
|
|
|
| 8 |
H |
0.245 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.457 |
-1.801 |
0.000 |
3.046 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.062 |
-4.164 |
0.000 |
| y |
-4.164 |
-19.336 |
0.000 |
| z |
0.000 |
0.000 |
-20.164 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.312 |
-4.164 |
0.000 |
| y |
-4.164 |
1.777 |
0.000 |
| z |
0.000 |
0.000 |
0.534 |
|
| Polar |
|---|
| 3z2-r2 | 1.069 |
|---|
| x2-y2 | -2.726 |
|---|
| xy | -4.164 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.460 |
-0.260 |
0.000 |
| y |
-0.260 |
3.667 |
0.000 |
| z |
0.000 |
0.000 |
2.648 |
<r2> (average value of r
2) Å
2
| <r2> |
50.928 |
| (<r2>)1/2 |
7.136 |
