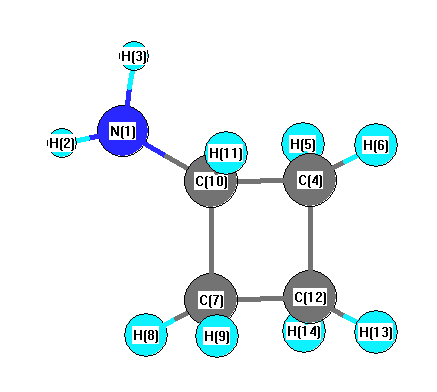
Vibrational Frequencies calculated at HF/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3774 |
3398 |
0.34 |
|
|
|
| 2 |
A' |
3321 |
2990 |
119.72 |
|
|
|
| 3 |
A' |
3291 |
2963 |
17.16 |
|
|
|
| 4 |
A' |
3256 |
2931 |
44.00 |
|
|
|
| 5 |
A' |
3234 |
2911 |
69.74 |
|
|
|
| 6 |
A' |
3176 |
2860 |
98.94 |
|
|
|
| 7 |
A' |
1828 |
1646 |
47.72 |
|
|
|
| 8 |
A' |
1651 |
1487 |
4.20 |
|
|
|
| 9 |
A' |
1628 |
1466 |
3.06 |
|
|
|
| 10 |
A' |
1520 |
1368 |
29.25 |
|
|
|
| 11 |
A' |
1437 |
1294 |
5.70 |
|
|
|
| 12 |
A' |
1358 |
1222 |
1.26 |
|
|
|
| 13 |
A' |
1282 |
1155 |
10.14 |
|
|
|
| 14 |
A' |
1172 |
1055 |
11.85 |
|
|
|
| 15 |
A' |
1025 |
923 |
0.38 |
|
|
|
| 16 |
A' |
957 |
862 |
4.96 |
|
|
|
| 17 |
A' |
898 |
809 |
9.60 |
|
|
|
| 18 |
A' |
773 |
696 |
45.91 |
|
|
|
| 19 |
A' |
697 |
628 |
222.69 |
|
|
|
| 20 |
A' |
438 |
394 |
8.19 |
|
|
|
| 21 |
A' |
130 |
117 |
1.05 |
|
|
|
| 22 |
A" |
3897 |
3509 |
4.16 |
|
|
|
| 23 |
A" |
3297 |
2968 |
15.95 |
|
|
|
| 24 |
A" |
3231 |
2910 |
112.90 |
|
|
|
| 25 |
A" |
1623 |
1462 |
5.61 |
|
|
|
| 26 |
A" |
1473 |
1327 |
2.00 |
|
|
|
| 27 |
A" |
1430 |
1288 |
0.14 |
|
|
|
| 28 |
A" |
1394 |
1255 |
2.37 |
|
|
|
| 29 |
A" |
1330 |
1197 |
2.63 |
|
|
|
| 30 |
A" |
1284 |
1156 |
0.35 |
|
|
|
| 31 |
A" |
1121 |
1009 |
0.07 |
|
|
|
| 32 |
A" |
1028 |
926 |
1.71 |
|
|
|
| 33 |
A" |
991 |
893 |
4.79 |
|
|
|
| 34 |
A" |
850 |
766 |
0.04 |
|
|
|
| 35 |
A" |
420 |
378 |
6.23 |
|
|
|
| 36 |
A" |
259 |
234 |
39.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30236.1 cm
-1
Scaled (by 0.9004) Zero Point Vibrational Energy (zpe) 27224.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.790 |
|
|
|
| 2 |
H |
0.305 |
|
|
|
| 3 |
H |
0.305 |
|
|
|
| 4 |
C |
-0.378 |
|
|
|
| 5 |
H |
0.184 |
|
|
|
| 6 |
H |
0.172 |
|
|
|
| 7 |
C |
-0.378 |
|
|
|
| 8 |
H |
0.184 |
|
|
|
| 9 |
H |
0.172 |
|
|
|
| 10 |
C |
0.020 |
|
|
|
| 11 |
H |
0.176 |
|
|
|
| 12 |
C |
-0.345 |
|
|
|
| 13 |
H |
0.182 |
|
|
|
| 14 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.357 |
1.168 |
0.000 |
1.222 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.738 |
-2.228 |
0.000 |
| y |
-2.228 |
-31.309 |
0.000 |
| z |
0.000 |
0.000 |
-31.544 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.311 |
-2.228 |
0.000 |
| y |
-2.228 |
0.831 |
0.000 |
| z |
0.000 |
0.000 |
0.479 |
|
| Polar |
|---|
| 3z2-r2 | 0.959 |
|---|
| x2-y2 | -1.428 |
|---|
| xy | -2.228 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.468 |
-0.472 |
0.000 |
| y |
-0.472 |
6.547 |
0.000 |
| z |
0.000 |
0.000 |
6.871 |
<r2> (average value of r
2) Å
2
| <r2> |
115.508 |
| (<r2>)1/2 |
10.747 |