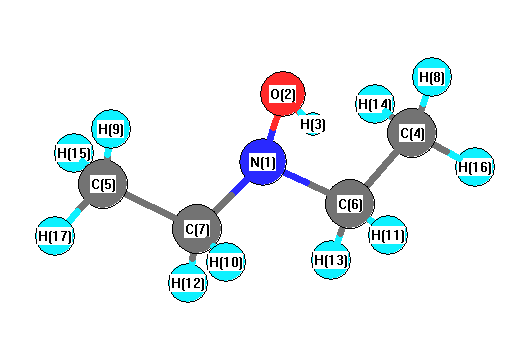
Vibrational Frequencies calculated at HF/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4005 |
3646 |
5.23 |
|
|
|
| 2 |
A |
3252 |
2960 |
83.05 |
|
|
|
| 3 |
A |
3224 |
2935 |
66.35 |
|
|
|
| 4 |
A |
3215 |
2927 |
26.30 |
|
|
|
| 5 |
A |
3168 |
2884 |
2.38 |
|
|
|
| 6 |
A |
3058 |
2784 |
164.33 |
|
|
|
| 7 |
A |
1635 |
1489 |
10.25 |
|
|
|
| 8 |
A |
1614 |
1469 |
1.40 |
|
|
|
| 9 |
A |
1603 |
1459 |
12.58 |
|
|
|
| 10 |
A |
1566 |
1425 |
17.19 |
|
|
|
| 11 |
A |
1533 |
1396 |
42.16 |
|
|
|
| 12 |
A |
1520 |
1384 |
2.20 |
|
|
|
| 13 |
A |
1428 |
1300 |
8.90 |
|
|
|
| 14 |
A |
1326 |
1207 |
3.60 |
|
|
|
| 15 |
A |
1271 |
1157 |
12.98 |
|
|
|
| 16 |
A |
1169 |
1064 |
3.26 |
|
|
|
| 17 |
A |
1065 |
970 |
17.77 |
|
|
|
| 18 |
A |
889 |
810 |
2.40 |
|
|
|
| 19 |
A |
860 |
783 |
17.09 |
|
|
|
| 20 |
A |
494 |
450 |
4.17 |
|
|
|
| 21 |
A |
451 |
411 |
0.05 |
|
|
|
| 22 |
A |
226 |
206 |
0.84 |
|
|
|
| 23 |
A |
225 |
204 |
2.14 |
|
|
|
| 24 |
A |
108 |
99 |
0.43 |
|
|
|
| 25 |
A |
3250 |
2959 |
0.19 |
|
|
|
| 26 |
A |
3224 |
2935 |
17.76 |
|
|
|
| 27 |
A |
3197 |
2911 |
0.09 |
|
|
|
| 28 |
A |
3167 |
2883 |
70.06 |
|
|
|
| 29 |
A |
3044 |
2772 |
3.40 |
|
|
|
| 30 |
A |
1627 |
1481 |
0.20 |
|
|
|
| 31 |
A |
1605 |
1462 |
3.96 |
|
|
|
| 32 |
A |
1601 |
1458 |
0.01 |
|
|
|
| 33 |
A |
1541 |
1403 |
27.01 |
|
|
|
| 34 |
A |
1512 |
1377 |
21.21 |
|
|
|
| 35 |
A |
1413 |
1286 |
1.41 |
|
|
|
| 36 |
A |
1314 |
1196 |
4.15 |
|
|
|
| 37 |
A |
1190 |
1084 |
16.34 |
|
|
|
| 38 |
A |
1170 |
1065 |
10.40 |
|
|
|
| 39 |
A |
1008 |
917 |
0.02 |
|
|
|
| 40 |
A |
866 |
788 |
0.64 |
|
|
|
| 41 |
A |
586 |
534 |
23.05 |
|
|
|
| 42 |
A |
432 |
394 |
116.75 |
|
|
|
| 43 |
A |
369 |
336 |
4.84 |
|
|
|
| 44 |
A |
263 |
239 |
0.04 |
|
|
|
| 45 |
A |
133 |
121 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35707.2 cm
-1
Scaled (by 0.9104) Zero Point Vibrational Energy (zpe) 32507.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.102 |
|
|
|
| 2 |
O |
-0.501 |
|
|
|
| 3 |
H |
0.161 |
|
|
|
| 4 |
C |
-0.997 |
|
|
|
| 5 |
C |
-0.997 |
|
|
|
| 6 |
C |
-0.276 |
|
|
|
| 7 |
C |
-0.276 |
|
|
|
| 8 |
H |
0.294 |
|
|
|
| 9 |
H |
0.294 |
|
|
|
| 10 |
H |
0.263 |
|
|
|
| 11 |
H |
0.263 |
|
|
|
| 12 |
H |
0.355 |
|
|
|
| 13 |
H |
0.355 |
|
|
|
| 14 |
H |
0.270 |
|
|
|
| 15 |
H |
0.270 |
|
|
|
| 16 |
H |
0.313 |
|
|
|
| 17 |
H |
0.313 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.112 |
2.391 |
0.000 |
2.393 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.041 |
1.164 |
0.000 |
| y |
1.164 |
-41.367 |
0.000 |
| z |
0.000 |
0.000 |
-38.233 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.760 |
1.164 |
0.000 |
| y |
1.164 |
-3.730 |
0.000 |
| z |
0.000 |
0.000 |
0.971 |
|
| Polar |
|---|
| 3z2-r2 | 1.941 |
|---|
| x2-y2 | 4.327 |
|---|
| xy | 1.164 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.677 |
-0.116 |
0.000 |
| y |
-0.116 |
8.517 |
0.000 |
| z |
0.000 |
0.000 |
10.439 |
<r2> (average value of r
2) Å
2
| <r2> |
210.066 |
| (<r2>)1/2 |
14.494 |