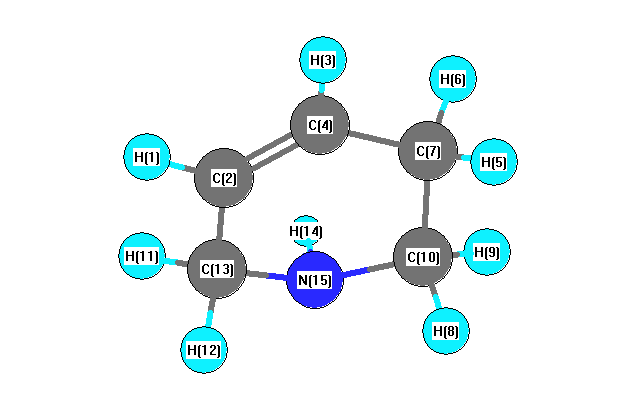
Vibrational Frequencies calculated at HF/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3746 |
3411 |
1.42 |
|
|
|
| 2 |
A |
3307 |
3011 |
36.62 |
|
|
|
| 3 |
A |
3280 |
2987 |
14.90 |
|
|
|
| 4 |
A |
3208 |
2920 |
59.35 |
|
|
|
| 5 |
A |
3188 |
2903 |
37.55 |
|
|
|
| 6 |
A |
3176 |
2891 |
82.17 |
|
|
|
| 7 |
A |
3157 |
2874 |
18.74 |
|
|
|
| 8 |
A |
3139 |
2858 |
51.36 |
|
|
|
| 9 |
A |
3131 |
2851 |
41.46 |
|
|
|
| 10 |
A |
1851 |
1685 |
3.73 |
|
|
|
| 11 |
A |
1628 |
1482 |
10.87 |
|
|
|
| 12 |
A |
1622 |
1477 |
1.89 |
|
|
|
| 13 |
A |
1610 |
1466 |
3.08 |
|
|
|
| 14 |
A |
1601 |
1458 |
3.54 |
|
|
|
| 15 |
A |
1537 |
1399 |
0.32 |
|
|
|
| 16 |
A |
1526 |
1389 |
12.65 |
|
|
|
| 17 |
A |
1493 |
1359 |
4.64 |
|
|
|
| 18 |
A |
1473 |
1341 |
0.20 |
|
|
|
| 19 |
A |
1439 |
1310 |
0.94 |
|
|
|
| 20 |
A |
1364 |
1242 |
4.03 |
|
|
|
| 21 |
A |
1310 |
1193 |
18.39 |
|
|
|
| 22 |
A |
1295 |
1179 |
5.40 |
|
|
|
| 23 |
A |
1225 |
1115 |
20.59 |
|
|
|
| 24 |
A |
1211 |
1103 |
1.81 |
|
|
|
| 25 |
A |
1117 |
1017 |
2.44 |
|
|
|
| 26 |
A |
1114 |
1014 |
1.17 |
|
|
|
| 27 |
A |
1089 |
991 |
9.78 |
|
|
|
| 28 |
A |
1062 |
967 |
10.69 |
|
|
|
| 29 |
A |
975 |
887 |
11.32 |
|
|
|
| 30 |
A |
949 |
864 |
4.15 |
|
|
|
| 31 |
A |
907 |
826 |
23.66 |
|
|
|
| 32 |
A |
863 |
786 |
61.44 |
|
|
|
| 33 |
A |
816 |
742 |
54.66 |
|
|
|
| 34 |
A |
717 |
653 |
25.31 |
|
|
|
| 35 |
A |
565 |
515 |
1.04 |
|
|
|
| 36 |
A |
520 |
474 |
0.52 |
|
|
|
| 37 |
A |
434 |
395 |
0.62 |
|
|
|
| 38 |
A |
308 |
280 |
8.10 |
|
|
|
| 39 |
A |
185 |
169 |
1.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31568.3 cm
-1
Scaled (by 0.9104) Zero Point Vibrational Energy (zpe) 28739.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.469 |
|
|
|
| 2 |
C |
-0.603 |
|
|
|
| 3 |
H |
0.334 |
|
|
|
| 4 |
C |
-0.679 |
|
|
|
| 5 |
H |
0.321 |
|
|
|
| 6 |
H |
0.258 |
|
|
|
| 7 |
C |
-0.231 |
|
|
|
| 8 |
H |
0.290 |
|
|
|
| 9 |
H |
0.285 |
|
|
|
| 10 |
C |
-0.469 |
|
|
|
| 11 |
H |
0.321 |
|
|
|
| 12 |
H |
0.315 |
|
|
|
| 13 |
C |
-0.172 |
|
|
|
| 14 |
H |
-0.004 |
|
|
|
| 15 |
N |
-0.435 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.510 |
0.749 |
-0.481 |
1.025 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.792 |
-1.882 |
0.208 |
| y |
-1.882 |
-40.811 |
0.955 |
| z |
0.208 |
0.955 |
-37.320 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.273 |
-1.882 |
0.208 |
| y |
-1.882 |
-4.255 |
0.955 |
| z |
0.208 |
0.955 |
0.982 |
|
| Polar |
|---|
| 3z2-r2 | 1.964 |
|---|
| x2-y2 | 5.018 |
|---|
| xy | -1.882 |
|---|
| xz | 0.208 |
|---|
| yz | 0.955 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.009 |
0.084 |
-0.172 |
| y |
0.084 |
9.244 |
0.008 |
| z |
-0.172 |
0.008 |
7.747 |
<r2> (average value of r
2) Å
2
| <r2> |
146.031 |
| (<r2>)1/2 |
12.084 |