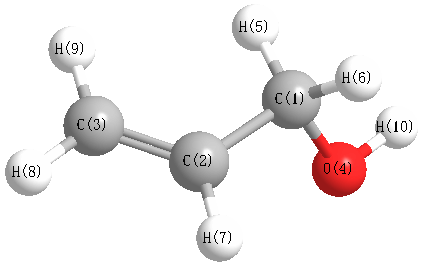
Vibrational Frequencies calculated at HF/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4226 |
3451 |
27.19 |
|
|
|
| 2 |
A |
3809 |
3110 |
0.02 |
|
|
|
| 3 |
A |
3717 |
3035 |
3.84 |
|
|
|
| 4 |
A |
3652 |
2982 |
30.27 |
|
|
|
| 5 |
A |
3646 |
2977 |
9.20 |
|
|
|
| 6 |
A |
3530 |
2883 |
7.10 |
|
|
|
| 7 |
A |
2070 |
1690 |
0.15 |
|
|
|
| 8 |
A |
1829 |
1494 |
0.82 |
|
|
|
| 9 |
A |
1778 |
1452 |
15.78 |
|
|
|
| 10 |
A |
1705 |
1392 |
4.09 |
|
|
|
| 11 |
A |
1542 |
1259 |
16.39 |
|
|
|
| 12 |
A |
1516 |
1238 |
34.47 |
|
|
|
| 13 |
A |
1449 |
1183 |
7.11 |
|
|
|
| 14 |
A |
1326 |
1083 |
5.01 |
|
|
|
| 15 |
A |
1317 |
1075 |
2.76 |
|
|
|
| 16 |
A |
1242 |
1014 |
3.79 |
|
|
|
| 17 |
A |
1163 |
949 |
16.25 |
|
|
|
| 18 |
A |
1118 |
913 |
3.26 |
|
|
|
| 19 |
A |
1054 |
861 |
3.21 |
|
|
|
| 20 |
A |
751 |
613 |
1.51 |
|
|
|
| 21 |
A |
484 |
395 |
1.71 |
|
|
|
| 22 |
A |
364 |
297 |
11.99 |
|
|
|
| 23 |
A |
336 |
274 |
74.43 |
|
|
|
| 24 |
A |
90 |
73 |
0.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21856.0 cm
-1
Scaled (by 0.8165) Zero Point Vibrational Energy (zpe) 17845.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.001 |
|
|
|
| 2 |
C |
-0.052 |
|
|
|
| 3 |
C |
-0.134 |
|
|
|
| 4 |
O |
-0.283 |
|
|
|
| 5 |
H |
0.052 |
|
|
|
| 6 |
H |
0.053 |
|
|
|
| 7 |
H |
0.066 |
|
|
|
| 8 |
H |
0.064 |
|
|
|
| 9 |
H |
0.060 |
|
|
|
| 10 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.234 |
1.312 |
0.645 |
1.481 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.853 |
2.575 |
0.939 |
| y |
2.575 |
-22.923 |
-0.864 |
| z |
0.939 |
-0.864 |
-23.984 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.601 |
2.575 |
0.939 |
| y |
2.575 |
-0.504 |
-0.864 |
| z |
0.939 |
-0.864 |
-2.096 |
|
| Polar |
|---|
| 3z2-r2 | -4.192 |
|---|
| x2-y2 | 2.070 |
|---|
| xy | 2.575 |
|---|
| xz | 0.939 |
|---|
| yz | -0.864 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.603 |
-0.111 |
0.606 |
| y |
-0.111 |
2.442 |
-0.424 |
| z |
0.606 |
-0.424 |
1.641 |
<r2> (average value of r
2) Å
2
| <r2> |
92.765 |
| (<r2>)1/2 |
9.631 |