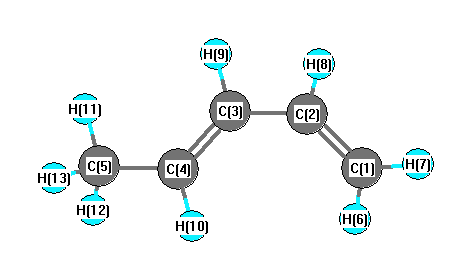
Vibrational Frequencies calculated at HF/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3812 |
3113 |
0.01 |
|
|
|
| 2 |
A' |
3769 |
3077 |
0.73 |
|
|
|
| 3 |
A' |
3729 |
3044 |
6.86 |
|
|
|
| 4 |
A' |
3716 |
3034 |
0.95 |
|
|
|
| 5 |
A' |
3690 |
3013 |
32.30 |
|
|
|
| 6 |
A' |
3653 |
2983 |
55.17 |
|
|
|
| 7 |
A' |
3564 |
2910 |
2.03 |
|
|
|
| 8 |
A' |
2089 |
1705 |
4.33 |
|
|
|
| 9 |
A' |
2013 |
1643 |
0.94 |
|
|
|
| 10 |
A' |
1828 |
1493 |
2.89 |
|
|
|
| 11 |
A' |
1743 |
1423 |
0.75 |
|
|
|
| 12 |
A' |
1722 |
1406 |
8.50 |
|
|
|
| 13 |
A' |
1644 |
1342 |
8.97 |
|
|
|
| 14 |
A' |
1540 |
1258 |
7.55 |
|
|
|
| 15 |
A' |
1486 |
1213 |
0.37 |
|
|
|
| 16 |
A' |
1367 |
1116 |
1.09 |
|
|
|
| 17 |
A' |
1234 |
1007 |
0.61 |
|
|
|
| 18 |
A' |
1115 |
911 |
3.01 |
|
|
|
| 19 |
A' |
1022 |
835 |
0.01 |
|
|
|
| 20 |
A' |
695 |
567 |
4.28 |
|
|
|
| 21 |
A' |
434 |
354 |
2.64 |
|
|
|
| 22 |
A' |
247 |
202 |
0.23 |
|
|
|
| 23 |
A" |
3723 |
3040 |
0.63 |
|
|
|
| 24 |
A" |
1821 |
1487 |
2.51 |
|
|
|
| 25 |
A" |
1275 |
1041 |
0.17 |
|
|
|
| 26 |
A" |
1250 |
1020 |
3.28 |
|
|
|
| 27 |
A" |
1211 |
989 |
0.35 |
|
|
|
| 28 |
A" |
1147 |
936 |
16.96 |
|
|
|
| 29 |
A" |
940 |
767 |
3.34 |
|
|
|
| 30 |
A" |
745 |
608 |
9.34 |
|
|
|
| 31 |
A" |
417 |
341 |
0.09 |
|
|
|
| 32 |
A" |
114 |
93 |
0.00 |
|
|
|
| 33 |
A" |
96 |
78 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29423.7 cm
-1
Scaled (by 0.8165) Zero Point Vibrational Energy (zpe) 24024.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.132 |
|
|
|
| 2 |
C |
-0.059 |
|
|
|
| 3 |
C |
-0.071 |
|
|
|
| 4 |
C |
-0.049 |
|
|
|
| 5 |
C |
-0.187 |
|
|
|
| 6 |
H |
0.060 |
|
|
|
| 7 |
H |
0.062 |
|
|
|
| 8 |
H |
0.061 |
|
|
|
| 9 |
H |
0.058 |
|
|
|
| 10 |
H |
0.057 |
|
|
|
| 11 |
H |
0.066 |
|
|
|
| 12 |
H |
0.067 |
|
|
|
| 13 |
H |
0.067 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.303 |
-0.128 |
0.000 |
0.328 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.480 |
0.077 |
0.000 |
| y |
0.077 |
-28.668 |
0.000 |
| z |
0.000 |
0.000 |
-31.226 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.466 |
0.077 |
0.000 |
| y |
0.077 |
1.185 |
0.000 |
| z |
0.000 |
0.000 |
-2.651 |
|
| Polar |
|---|
| 3z2-r2 | -5.303 |
|---|
| x2-y2 | 0.187 |
|---|
| xy | 0.077 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.877 |
0.688 |
0.000 |
| y |
0.688 |
3.874 |
0.000 |
| z |
0.000 |
0.000 |
1.465 |
<r2> (average value of r
2) Å
2
| <r2> |
154.754 |
| (<r2>)1/2 |
12.440 |