Jump to
S1C2
Vibrational Frequencies calculated at HF/STO-3G
Geometric Data calculated at HF/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HF/STO-3G
| | hartrees |
|---|
| Energy at 0K | -231.437890 |
| Energy at 298.15K | -231.451448 |
| Nuclear repulsion energy | 253.698188 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3751 |
3063 |
2.63 |
|
|
|
| 2 |
A' |
3750 |
3062 |
4.86 |
|
|
|
| 3 |
A' |
3747 |
3060 |
0.95 |
|
|
|
| 4 |
A' |
3740 |
3054 |
0.20 |
|
|
|
| 5 |
A' |
3623 |
2958 |
1.12 |
|
|
|
| 6 |
A' |
3611 |
2949 |
0.83 |
|
|
|
| 7 |
A' |
3563 |
2909 |
2.23 |
|
|
|
| 8 |
A' |
3563 |
2909 |
2.68 |
|
|
|
| 9 |
A' |
1841 |
1503 |
0.96 |
|
|
|
| 10 |
A' |
1835 |
1499 |
0.17 |
|
|
|
| 11 |
A' |
1833 |
1497 |
0.46 |
|
|
|
| 12 |
A' |
1806 |
1475 |
0.35 |
|
|
|
| 13 |
A' |
1742 |
1423 |
0.65 |
|
|
|
| 14 |
A' |
1736 |
1417 |
0.07 |
|
|
|
| 15 |
A' |
1651 |
1348 |
0.23 |
|
|
|
| 16 |
A' |
1569 |
1281 |
7.91 |
|
|
|
| 17 |
A' |
1489 |
1215 |
14.19 |
|
|
|
| 18 |
A' |
1438 |
1174 |
0.76 |
|
|
|
| 19 |
A' |
1201 |
981 |
1.21 |
|
|
|
| 20 |
A' |
1169 |
954 |
0.31 |
|
|
|
| 21 |
A' |
1151 |
940 |
0.27 |
|
|
|
| 22 |
A' |
1114 |
909 |
0.15 |
|
|
|
| 23 |
A' |
1003 |
819 |
0.12 |
|
|
|
| 24 |
A' |
858 |
701 |
0.94 |
|
|
|
| 25 |
A' |
728 |
595 |
0.34 |
|
|
|
| 26 |
A' |
438 |
358 |
0.10 |
|
|
|
| 27 |
A' |
383 |
313 |
0.00 |
|
|
|
| 28 |
A' |
67 |
55 |
0.00 |
|
|
|
| 29 |
A" |
3747 |
3059 |
3.20 |
|
|
|
| 30 |
A" |
3746 |
3058 |
0.22 |
|
|
|
| 31 |
A" |
3742 |
3055 |
0.12 |
|
|
|
| 32 |
A" |
3614 |
2951 |
2.76 |
|
|
|
| 33 |
A" |
1844 |
1506 |
1.59 |
|
|
|
| 34 |
A" |
1834 |
1497 |
0.00 |
|
|
|
| 35 |
A" |
1810 |
1478 |
0.06 |
|
|
|
| 36 |
A" |
1587 |
1296 |
3.16 |
|
|
|
| 37 |
A" |
1535 |
1254 |
0.58 |
|
|
|
| 38 |
A" |
1474 |
1204 |
3.41 |
|
|
|
| 39 |
A" |
1461 |
1193 |
0.91 |
|
|
|
| 40 |
A" |
1296 |
1058 |
0.00 |
|
|
|
| 41 |
A" |
1190 |
972 |
0.00 |
|
|
|
| 42 |
A" |
1173 |
958 |
0.03 |
|
|
|
| 43 |
A" |
1090 |
890 |
0.61 |
|
|
|
| 44 |
A" |
944 |
771 |
0.14 |
|
|
|
| 45 |
A" |
442 |
361 |
0.04 |
|
|
|
| 46 |
A" |
347 |
283 |
0.00 |
|
|
|
| 47 |
A" |
274 |
224 |
0.02 |
|
|
|
| 48 |
A" |
240 |
196 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43894.6 cm
-1
Scaled (by 0.8165) Zero Point Vibrational Energy (zpe) 35839.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.046 |
0.406 |
0.000 |
| C2 |
0.167 |
-1.786 |
0.000 |
| C3 |
-0.046 |
-0.709 |
1.096 |
| C4 |
-0.046 |
-0.709 |
-1.096 |
| C5 |
1.250 |
1.248 |
0.000 |
| C6 |
-1.286 |
1.324 |
0.000 |
| H7 |
1.168 |
-2.206 |
0.000 |
| H8 |
-0.553 |
-2.599 |
0.000 |
| H9 |
0.744 |
-0.633 |
1.838 |
| H10 |
0.744 |
-0.633 |
-1.838 |
| H11 |
-0.999 |
-0.803 |
1.608 |
| H12 |
-0.999 |
-0.803 |
-1.608 |
| H13 |
2.125 |
0.606 |
0.000 |
| H14 |
-2.199 |
0.737 |
0.000 |
| H15 |
1.292 |
1.882 |
-0.880 |
| H16 |
1.292 |
1.882 |
0.880 |
| H17 |
-1.290 |
1.961 |
0.880 |
| H18 |
-1.290 |
1.961 |
-0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.2022 | 1.5634 | 1.5634 | 1.5443 | 1.5429 | 2.8810 | 3.0483 | 2.2543 | 2.2543 | 2.2266 | 2.2266 | 2.1801 | 2.1790 | 2.1781 | 2.1781 | 2.1769 | 2.1769 |
C2 | 2.2022 | | 1.5507 | 1.5507 | 3.2206 | 3.4322 | 1.0865 | 1.0865 | 2.2453 | 2.2453 | 2.2159 | 2.2159 | 3.0911 | 3.4587 | 3.9366 | 3.9366 | 4.1147 | 4.1147 | C3 | 1.5634 | 1.5507 | | 2.1912 | 2.5895 | 2.6214 | 2.2173 | 2.2432 | 1.0866 | 3.0391 | 1.0865 | 2.8685 | 2.7644 | 2.8161 | 3.5228 | 2.9243 | 2.9532 | 3.5467 | C4 | 1.5634 | 1.5507 | 2.1912 | | 2.5895 | 2.6214 | 2.2173 | 2.2432 | 3.0391 | 1.0866 | 2.8685 | 1.0865 | 2.7644 | 2.8161 | 2.9243 | 3.5228 | 3.5467 | 2.9532 | C5 | 1.5443 | 3.2206 | 2.5895 | 2.5895 | | 2.5367 | 3.4549 | 4.2484 | 2.6775 | 2.6775 | 3.4419 | 3.4419 | 1.0857 | 3.4865 | 1.0863 | 1.0863 | 2.7807 | 2.7807 | C6 | 1.5429 | 3.4322 | 2.6214 | 2.6214 | 2.5367 | | 4.2997 | 3.9912 | 3.3657 | 3.3657 | 2.6819 | 2.6819 | 3.4861 | 1.0857 | 2.7811 | 2.7811 | 1.0864 | 1.0864 | H7 | 2.8810 | 1.0865 | 2.2173 | 2.2173 | 3.4549 | 4.2997 | | 1.7659 | 2.4568 | 2.4568 | 3.0419 | 3.0419 | 2.9706 | 4.4728 | 4.1843 | 4.1843 | 4.9176 | 4.9176 | H8 | 3.0483 | 1.0865 | 2.2432 | 2.2432 | 4.2484 | 3.9912 | 1.7659 | | 2.9883 | 2.9883 | 2.4519 | 2.4519 | 4.1771 | 3.7205 | 4.9262 | 4.9262 | 4.7023 | 4.7023 | H9 | 2.2543 | 2.2453 | 1.0866 | 3.0391 | 2.6775 | 3.3657 | 2.4568 | 2.9883 | | 3.6763 | 1.7663 | 3.8658 | 2.6116 | 3.7308 | 3.7438 | 2.7465 | 3.4321 | 4.2722 | H10 | 2.2543 | 2.2453 | 3.0391 | 1.0866 | 2.6775 | 3.3657 | 2.4568 | 2.9883 | 3.6763 | | 3.8658 | 1.7663 | 2.6116 | 3.7308 | 2.7465 | 3.7438 | 4.2722 | 3.4321 | H11 | 2.2266 | 2.2159 | 1.0865 | 2.8685 | 3.4419 | 2.6819 | 3.0419 | 2.4519 | 1.7663 | 3.8658 | | 3.2163 | 3.7858 | 2.5297 | 4.3191 | 3.6043 | 2.8727 | 3.7303 | H12 | 2.2266 | 2.2159 | 2.8685 | 1.0865 | 3.4419 | 2.6819 | 3.0419 | 2.4519 | 3.8658 | 1.7663 | 3.2163 | | 3.7858 | 2.5297 | 3.6043 | 4.3191 | 3.7303 | 2.8727 | H13 | 2.1801 | 3.0911 | 2.7644 | 2.7644 | 1.0857 | 3.4861 | 2.9706 | 4.1771 | 2.6116 | 2.6116 | 3.7858 | 3.7858 | | 4.3267 | 1.7603 | 1.7603 | 3.7781 | 3.7781 | H14 | 2.1790 | 3.4587 | 2.8161 | 2.8161 | 3.4865 | 1.0857 | 4.4728 | 3.7205 | 3.7308 | 3.7308 | 2.5297 | 2.5297 | 4.3267 | | 3.7788 | 3.7788 | 1.7604 | 1.7604 | H15 | 2.1781 | 3.9366 | 3.5228 | 2.9243 | 1.0863 | 2.7811 | 4.1843 | 4.9262 | 3.7438 | 2.7465 | 4.3191 | 3.6043 | 1.7603 | 3.7788 | | 1.7608 | 3.1264 | 2.5835 | H16 | 2.1781 | 3.9366 | 2.9243 | 3.5228 | 1.0863 | 2.7811 | 4.1843 | 4.9262 | 2.7465 | 3.7438 | 3.6043 | 4.3191 | 1.7603 | 3.7788 | 1.7608 | | 2.5835 | 3.1264 | H17 | 2.1769 | 4.1147 | 2.9532 | 3.5467 | 2.7807 | 1.0864 | 4.9176 | 4.7023 | 3.4321 | 4.2722 | 2.8727 | 3.7303 | 3.7781 | 1.7604 | 3.1264 | 2.5835 | | 1.7605 | H18 | 2.1769 | 4.1147 | 3.5467 | 2.9532 | 2.7807 | 1.0864 | 4.9176 | 4.7023 | 4.2722 | 3.4321 | 3.7303 | 2.8727 | 3.7781 | 1.7604 | 2.5835 | 3.1264 | 1.7605 | |
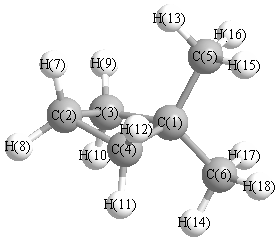 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
90.012 |
|
C1 |
C3 |
H9 |
115.388 |
| C1 |
C3 |
H11 |
113.101 |
|
C1 |
C4 |
C2 |
90.012 |
| C1 |
C4 |
H10 |
115.388 |
|
C1 |
C4 |
H12 |
113.101 |
| C1 |
C5 |
H13 |
110.767 |
|
C1 |
C5 |
H15 |
110.572 |
| C1 |
C5 |
H16 |
110.572 |
|
C1 |
C6 |
H14 |
110.784 |
| C1 |
C6 |
H17 |
110.577 |
|
C1 |
C6 |
H18 |
110.577 |
| C2 |
C3 |
H9 |
115.603 |
|
C2 |
C3 |
H11 |
113.161 |
| C2 |
C4 |
H10 |
115.603 |
|
C2 |
C4 |
H12 |
113.161 |
| C3 |
C1 |
C4 |
88.977 |
|
C3 |
C1 |
C5 |
112.864 |
| C3 |
C1 |
C6 |
115.106 |
|
C3 |
C2 |
C4 |
89.907 |
| C3 |
C2 |
H7 |
113.272 |
|
C3 |
C2 |
H8 |
115.425 |
| C4 |
C1 |
C5 |
112.864 |
|
C4 |
C1 |
C6 |
115.106 |
| C4 |
C2 |
H7 |
113.272 |
|
C4 |
C2 |
H8 |
115.425 |
| C5 |
C1 |
C6 |
110.504 |
|
H7 |
C2 |
H8 |
108.712 |
| H9 |
C3 |
H11 |
108.745 |
|
H10 |
C4 |
H12 |
108.745 |
| H13 |
C5 |
H15 |
108.279 |
|
H13 |
C5 |
H16 |
108.279 |
| H14 |
C6 |
H17 |
108.283 |
|
H14 |
C6 |
H18 |
108.283 |
| H15 |
C5 |
H16 |
108.284 |
|
H17 |
C6 |
H18 |
108.246 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.050 |
|
|
|
| 2 |
C |
-0.108 |
|
|
|
| 3 |
C |
-0.112 |
|
|
|
| 4 |
C |
-0.112 |
|
|
|
| 5 |
C |
-0.181 |
|
|
|
| 6 |
C |
-0.183 |
|
|
|
| 7 |
H |
0.053 |
|
|
|
| 8 |
H |
0.052 |
|
|
|
| 9 |
H |
0.050 |
|
|
|
| 10 |
H |
0.050 |
|
|
|
| 11 |
H |
0.052 |
|
|
|
| 12 |
H |
0.052 |
|
|
|
| 13 |
H |
0.055 |
|
|
|
| 14 |
H |
0.055 |
|
|
|
| 15 |
H |
0.056 |
|
|
|
| 16 |
H |
0.056 |
|
|
|
| 17 |
H |
0.057 |
|
|
|
| 18 |
H |
0.057 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.005 |
0.057 |
0.000 |
0.057 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.587 |
0.120 |
0.000 |
| y |
0.120 |
-38.355 |
0.000 |
| z |
0.000 |
0.000 |
-38.117 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.649 |
0.120 |
0.000 |
| y |
0.120 |
-0.503 |
0.000 |
| z |
0.000 |
0.000 |
-0.146 |
|
| Polar |
|---|
| 3z2-r2 | -0.292 |
|---|
| x2-y2 | 0.768 |
|---|
| xy | 0.120 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.536 |
-0.017 |
0.000 |
| y |
-0.017 |
4.911 |
0.000 |
| z |
0.000 |
0.000 |
4.512 |
<r2> (average value of r
2) Å
2
| <r2> |
168.880 |
| (<r2>)1/2 |
12.995 |
