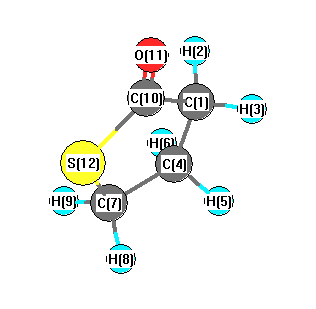
Vibrational Frequencies calculated at HF/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3273 |
2974 |
19.78 |
|
|
|
| 2 |
A |
3258 |
2960 |
15.58 |
|
|
|
| 3 |
A |
3235 |
2939 |
40.27 |
|
|
|
| 4 |
A |
3211 |
2917 |
20.80 |
|
|
|
| 5 |
A |
3189 |
2897 |
30.27 |
|
|
|
| 6 |
A |
3177 |
2886 |
11.34 |
|
|
|
| 7 |
A |
1995 |
1813 |
500.10 |
|
|
|
| 8 |
A |
1628 |
1479 |
0.10 |
|
|
|
| 9 |
A |
1610 |
1463 |
5.24 |
|
|
|
| 10 |
A |
1582 |
1437 |
8.56 |
|
|
|
| 11 |
A |
1482 |
1346 |
0.57 |
|
|
|
| 12 |
A |
1442 |
1310 |
1.88 |
|
|
|
| 13 |
A |
1430 |
1300 |
14.14 |
|
|
|
| 14 |
A |
1353 |
1230 |
1.97 |
|
|
|
| 15 |
A |
1297 |
1178 |
1.59 |
|
|
|
| 16 |
A |
1213 |
1102 |
19.04 |
|
|
|
| 17 |
A |
1167 |
1060 |
90.45 |
|
|
|
| 18 |
A |
1123 |
1020 |
9.63 |
|
|
|
| 19 |
A |
1095 |
995 |
12.42 |
|
|
|
| 20 |
A |
974 |
885 |
11.85 |
|
|
|
| 21 |
A |
939 |
853 |
6.61 |
|
|
|
| 22 |
A |
905 |
822 |
12.01 |
|
|
|
| 23 |
A |
749 |
681 |
4.83 |
|
|
|
| 24 |
A |
678 |
616 |
42.88 |
|
|
|
| 25 |
A |
652 |
593 |
12.95 |
|
|
|
| 26 |
A |
523 |
475 |
0.61 |
|
|
|
| 27 |
A |
510 |
463 |
3.76 |
|
|
|
| 28 |
A |
444 |
404 |
4.09 |
|
|
|
| 29 |
A |
250 |
227 |
1.69 |
|
|
|
| 30 |
A |
130 |
118 |
0.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22257.1 cm
-1
Scaled (by 0.9085) Zero Point Vibrational Energy (zpe) 20220.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.208 |
|
|
|
| 2 |
H |
0.145 |
|
|
|
| 3 |
H |
0.139 |
|
|
|
| 4 |
C |
-0.208 |
|
|
|
| 5 |
H |
0.119 |
|
|
|
| 6 |
H |
0.127 |
|
|
|
| 7 |
C |
-0.329 |
|
|
|
| 8 |
H |
0.140 |
|
|
|
| 9 |
H |
0.148 |
|
|
|
| 10 |
C |
0.248 |
|
|
|
| 11 |
O |
-0.400 |
|
|
|
| 12 |
S |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.049 |
1.981 |
0.424 |
4.527 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.174 |
-0.472 |
-0.118 |
| y |
-0.472 |
-43.288 |
0.123 |
| z |
-0.118 |
0.123 |
-42.815 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.123 |
-0.472 |
-0.118 |
| y |
-0.472 |
2.206 |
0.123 |
| z |
-0.118 |
0.123 |
2.916 |
|
| Polar |
|---|
| 3z2-r2 | 5.833 |
|---|
| x2-y2 | -4.886 |
|---|
| xy | -0.472 |
|---|
| xz | -0.118 |
|---|
| yz | 0.123 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.905 |
-0.219 |
0.165 |
| y |
-0.219 |
9.248 |
0.134 |
| z |
0.165 |
0.134 |
6.124 |
<r2> (average value of r
2) Å
2
| <r2> |
177.216 |
| (<r2>)1/2 |
13.312 |