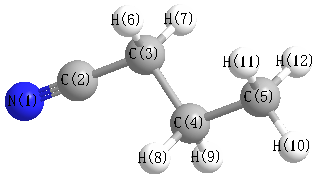
Vibrational Frequencies calculated at HF/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3233 |
2938 |
48.02 |
|
|
|
| 2 |
A' |
3199 |
2906 |
17.95 |
|
|
|
| 3 |
A' |
3183 |
2892 |
11.63 |
|
|
|
| 4 |
A' |
3160 |
2871 |
30.93 |
|
|
|
| 5 |
A' |
2594 |
2357 |
16.97 |
|
|
|
| 6 |
A' |
1627 |
1478 |
6.09 |
|
|
|
| 7 |
A' |
1611 |
1464 |
1.25 |
|
|
|
| 8 |
A' |
1597 |
1451 |
3.36 |
|
|
|
| 9 |
A' |
1544 |
1403 |
1.63 |
|
|
|
| 10 |
A' |
1518 |
1379 |
2.71 |
|
|
|
| 11 |
A' |
1418 |
1289 |
3.19 |
|
|
|
| 12 |
A' |
1204 |
1094 |
2.00 |
|
|
|
| 13 |
A' |
1109 |
1008 |
0.08 |
|
|
|
| 14 |
A' |
1003 |
911 |
4.62 |
|
|
|
| 15 |
A' |
940 |
854 |
2.57 |
|
|
|
| 16 |
A' |
582 |
529 |
1.28 |
|
|
|
| 17 |
A' |
373 |
339 |
0.37 |
|
|
|
| 18 |
A' |
184 |
167 |
7.73 |
|
|
|
| 19 |
A" |
3247 |
2950 |
52.59 |
|
|
|
| 20 |
A" |
3225 |
2930 |
27.80 |
|
|
|
| 21 |
A" |
3204 |
2911 |
2.20 |
|
|
|
| 22 |
A" |
1616 |
1468 |
7.63 |
|
|
|
| 23 |
A" |
1437 |
1305 |
0.08 |
|
|
|
| 24 |
A" |
1366 |
1241 |
0.01 |
|
|
|
| 25 |
A" |
1230 |
1117 |
0.02 |
|
|
|
| 26 |
A" |
943 |
857 |
0.07 |
|
|
|
| 27 |
A" |
797 |
725 |
2.53 |
|
|
|
| 28 |
A" |
436 |
396 |
1.11 |
|
|
|
| 29 |
A" |
258 |
234 |
0.03 |
|
|
|
| 30 |
A" |
106 |
97 |
4.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23971.9 cm
-1
Scaled (by 0.9085) Zero Point Vibrational Energy (zpe) 21778.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.339 |
|
|
|
| 2 |
C |
0.094 |
|
|
|
| 3 |
C |
-0.122 |
|
|
|
| 4 |
C |
-0.202 |
|
|
|
| 5 |
C |
-0.240 |
|
|
|
| 6 |
H |
0.145 |
|
|
|
| 7 |
H |
0.145 |
|
|
|
| 8 |
H |
0.114 |
|
|
|
| 9 |
H |
0.114 |
|
|
|
| 10 |
H |
0.106 |
|
|
|
| 11 |
H |
0.092 |
|
|
|
| 12 |
H |
0.092 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.318 |
0.073 |
0.000 |
4.319 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.782 |
1.016 |
0.000 |
| y |
1.016 |
-30.364 |
0.000 |
| z |
0.000 |
0.000 |
-30.575 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.313 |
1.016 |
0.000 |
| y |
1.016 |
6.814 |
0.000 |
| z |
0.000 |
0.000 |
6.498 |
|
| Polar |
|---|
| 3z2-r2 | 12.996 |
|---|
| x2-y2 | -13.418 |
|---|
| xy | 1.016 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.707 |
-0.215 |
0.000 |
| y |
-0.215 |
5.833 |
0.000 |
| z |
0.000 |
0.000 |
5.474 |
<r2> (average value of r
2) Å
2
| <r2> |
158.732 |
| (<r2>)1/2 |
12.599 |