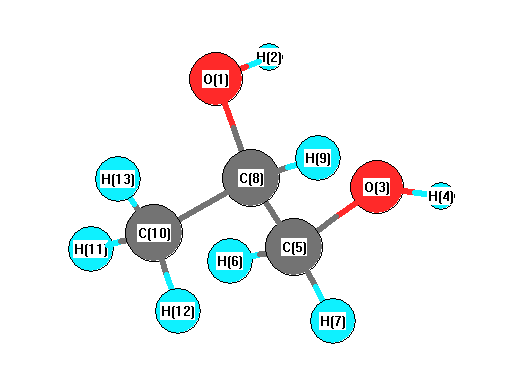
Vibrational Frequencies calculated at HF/6-311G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4174 |
3792 |
50.45 |
|
|
|
| 2 |
A |
4156 |
3776 |
74.22 |
|
|
|
| 3 |
A |
3249 |
2952 |
64.27 |
|
|
|
| 4 |
A |
3246 |
2949 |
39.31 |
|
|
|
| 5 |
A |
3229 |
2934 |
26.47 |
|
|
|
| 6 |
A |
3172 |
2882 |
65.79 |
|
|
|
| 7 |
A |
3164 |
2874 |
12.34 |
|
|
|
| 8 |
A |
3096 |
2812 |
82.86 |
|
|
|
| 9 |
A |
1627 |
1478 |
3.12 |
|
|
|
| 10 |
A |
1618 |
1470 |
1.48 |
|
|
|
| 11 |
A |
1606 |
1459 |
8.35 |
|
|
|
| 12 |
A |
1582 |
1437 |
13.13 |
|
|
|
| 13 |
A |
1546 |
1405 |
38.13 |
|
|
|
| 14 |
A |
1517 |
1378 |
3.13 |
|
|
|
| 15 |
A |
1508 |
1370 |
29.92 |
|
|
|
| 16 |
A |
1472 |
1337 |
45.80 |
|
|
|
| 17 |
A |
1401 |
1273 |
74.66 |
|
|
|
| 18 |
A |
1330 |
1208 |
24.20 |
|
|
|
| 19 |
A |
1283 |
1165 |
22.71 |
|
|
|
| 20 |
A |
1193 |
1084 |
71.48 |
|
|
|
| 21 |
A |
1174 |
1067 |
4.94 |
|
|
|
| 22 |
A |
1147 |
1042 |
105.92 |
|
|
|
| 23 |
A |
1020 |
927 |
6.24 |
|
|
|
| 24 |
A |
1004 |
912 |
18.56 |
|
|
|
| 25 |
A |
914 |
831 |
13.13 |
|
|
|
| 26 |
A |
566 |
514 |
30.85 |
|
|
|
| 27 |
A |
521 |
473 |
46.61 |
|
|
|
| 28 |
A |
485 |
441 |
130.41 |
|
|
|
| 29 |
A |
389 |
354 |
16.27 |
|
|
|
| 30 |
A |
332 |
302 |
85.76 |
|
|
|
| 31 |
A |
271 |
246 |
5.51 |
|
|
|
| 32 |
A |
240 |
218 |
0.58 |
|
|
|
| 33 |
A |
160 |
145 |
3.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26694.9 cm
-1
Scaled (by 0.9085) Zero Point Vibrational Energy (zpe) 24252.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.482 |
|
|
|
| 2 |
H |
0.258 |
|
|
|
| 3 |
O |
-0.487 |
|
|
|
| 4 |
H |
0.245 |
|
|
|
| 5 |
C |
0.100 |
|
|
|
| 6 |
H |
0.095 |
|
|
|
| 7 |
H |
0.075 |
|
|
|
| 8 |
C |
0.085 |
|
|
|
| 9 |
H |
0.055 |
|
|
|
| 10 |
C |
-0.231 |
|
|
|
| 11 |
H |
0.096 |
|
|
|
| 12 |
H |
0.081 |
|
|
|
| 13 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.504 |
-1.777 |
1.663 |
2.486 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.685 |
1.931 |
2.930 |
| y |
1.931 |
-32.110 |
0.196 |
| z |
2.930 |
0.196 |
-29.631 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.814 |
1.931 |
2.930 |
| y |
1.931 |
-0.452 |
0.196 |
| z |
2.930 |
0.196 |
3.266 |
|
| Polar |
|---|
| 3z2-r2 | 6.533 |
|---|
| x2-y2 | -1.575 |
|---|
| xy | 1.931 |
|---|
| xz | 2.930 |
|---|
| yz | 0.196 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.327 |
0.246 |
0.160 |
| y |
0.246 |
5.715 |
-0.046 |
| z |
0.160 |
-0.046 |
5.553 |
<r2> (average value of r
2) Å
2
| <r2> |
131.795 |
| (<r2>)1/2 |
11.480 |