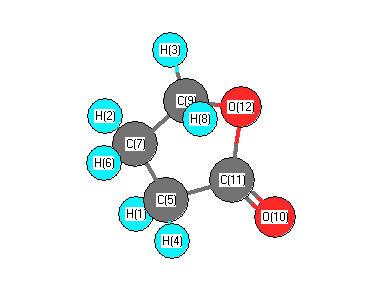
Vibrational Frequencies calculated at HF/cc-pVQZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3272 |
2972 |
17.19 |
79.40 |
0.32 |
0.48 |
| 2 |
A |
3264 |
2965 |
33.43 |
73.41 |
0.66 |
0.80 |
| 3 |
A |
3246 |
2949 |
27.95 |
110.10 |
0.17 |
0.30 |
| 4 |
A |
3194 |
2902 |
31.90 |
36.55 |
0.43 |
0.60 |
| 5 |
A |
3190 |
2898 |
16.34 |
54.92 |
0.21 |
0.35 |
| 6 |
A |
3189 |
2897 |
32.36 |
201.45 |
0.11 |
0.20 |
| 7 |
A |
2028 |
1843 |
567.97 |
10.92 |
0.44 |
0.61 |
| 8 |
A |
1661 |
1509 |
1.20 |
4.14 |
0.65 |
0.79 |
| 9 |
A |
1626 |
1477 |
6.60 |
4.75 |
0.75 |
0.86 |
| 10 |
A |
1590 |
1444 |
8.82 |
5.58 |
0.75 |
0.86 |
| 11 |
A |
1540 |
1399 |
35.26 |
0.82 |
0.26 |
0.41 |
| 12 |
A |
1468 |
1334 |
6.36 |
0.20 |
0.56 |
0.72 |
| 13 |
A |
1433 |
1302 |
19.95 |
0.92 |
0.60 |
0.75 |
| 14 |
A |
1374 |
1248 |
21.85 |
5.04 |
0.72 |
0.84 |
| 15 |
A |
1331 |
1209 |
37.01 |
3.52 |
0.74 |
0.85 |
| 16 |
A |
1308 |
1189 |
17.04 |
0.32 |
0.41 |
0.59 |
| 17 |
A |
1307 |
1187 |
206.03 |
0.78 |
0.69 |
0.82 |
| 18 |
A |
1202 |
1092 |
21.82 |
0.94 |
0.72 |
0.84 |
| 19 |
A |
1189 |
1080 |
92.98 |
1.81 |
0.72 |
0.84 |
| 20 |
A |
1079 |
980 |
6.63 |
3.55 |
0.71 |
0.83 |
| 21 |
A |
996 |
905 |
7.09 |
9.55 |
0.09 |
0.16 |
| 22 |
A |
968 |
879 |
10.38 |
1.55 |
0.31 |
0.47 |
| 23 |
A |
951 |
863 |
9.62 |
8.43 |
0.08 |
0.15 |
| 24 |
A |
873 |
793 |
5.75 |
5.29 |
0.13 |
0.22 |
| 25 |
A |
742 |
674 |
9.72 |
3.28 |
0.33 |
0.50 |
| 26 |
A |
697 |
633 |
6.12 |
1.45 |
0.56 |
0.72 |
| 27 |
A |
584 |
531 |
6.56 |
0.95 |
0.71 |
0.83 |
| 28 |
A |
535 |
486 |
6.32 |
1.19 |
0.42 |
0.59 |
| 29 |
A |
216 |
196 |
2.40 |
0.15 |
0.75 |
0.85 |
| 30 |
A |
153 |
139 |
1.23 |
0.30 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 23102.3 cm
-1
Scaled (by 0.9084) Zero Point Vibrational Energy (zpe) 20986.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/cc-pVQZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.515 |
|
|
|
| 2 |
C |
-0.233 |
|
|
|
| 3 |
C |
-0.175 |
|
|
|
| 4 |
C |
0.116 |
|
|
|
| 5 |
O |
-0.365 |
|
|
|
| 6 |
O |
-0.416 |
|
|
|
| 7 |
H |
0.122 |
|
|
|
| 8 |
H |
0.102 |
|
|
|
| 9 |
H |
0.086 |
|
|
|
| 10 |
H |
0.095 |
|
|
|
| 11 |
H |
0.096 |
|
|
|
| 12 |
H |
0.057 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.736 |
1.589 |
0.424 |
5.013 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.471 |
1.824 |
0.294 |
| y |
1.824 |
-35.294 |
-0.125 |
| z |
0.294 |
-0.125 |
-33.425 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.112 |
1.824 |
0.294 |
| y |
1.824 |
2.155 |
-0.125 |
| z |
0.294 |
-0.125 |
4.957 |
|
| Polar |
|---|
| 3z2-r2 | 9.914 |
|---|
| x2-y2 | -6.178 |
|---|
| xy | 1.824 |
|---|
| xz | 0.294 |
|---|
| yz | -0.125 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.195 |
0.270 |
-0.053 |
| y |
0.270 |
6.820 |
0.006 |
| z |
-0.053 |
0.006 |
5.759 |
<r2> (average value of r
2) Å
2
| <r2> |
137.515 |
| (<r2>)1/2 |
11.727 |