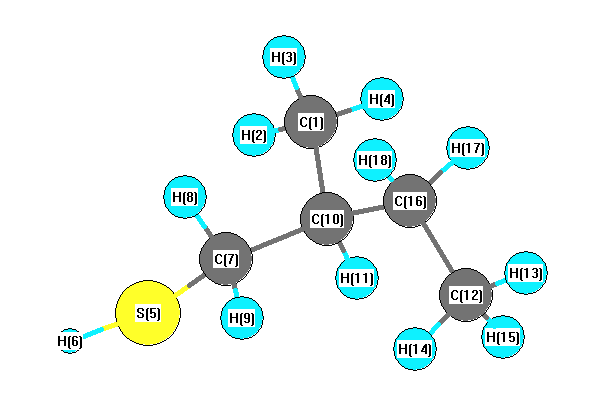
Vibrational Frequencies calculated at HF/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3267 |
2955 |
15.85 |
|
|
|
| 2 |
A |
3233 |
2924 |
41.61 |
|
|
|
| 3 |
A |
3221 |
2914 |
99.41 |
|
|
|
| 4 |
A |
3212 |
2905 |
74.11 |
|
|
|
| 5 |
A |
3209 |
2903 |
16.42 |
|
|
|
| 6 |
A |
3196 |
2891 |
18.18 |
|
|
|
| 7 |
A |
3174 |
2871 |
14.41 |
|
|
|
| 8 |
A |
3157 |
2856 |
24.71 |
|
|
|
| 9 |
A |
3156 |
2855 |
56.13 |
|
|
|
| 10 |
A |
3147 |
2847 |
7.86 |
|
|
|
| 11 |
A |
3138 |
2839 |
8.75 |
|
|
|
| 12 |
A |
2853 |
2581 |
4.46 |
|
|
|
| 13 |
A |
1626 |
1471 |
7.46 |
|
|
|
| 14 |
A |
1621 |
1466 |
7.01 |
|
|
|
| 15 |
A |
1619 |
1464 |
4.06 |
|
|
|
| 16 |
A |
1616 |
1462 |
1.29 |
|
|
|
| 17 |
A |
1610 |
1456 |
4.27 |
|
|
|
| 18 |
A |
1602 |
1449 |
4.39 |
|
|
|
| 19 |
A |
1543 |
1396 |
4.24 |
|
|
|
| 20 |
A |
1541 |
1394 |
5.15 |
|
|
|
| 21 |
A |
1515 |
1370 |
2.74 |
|
|
|
| 22 |
A |
1502 |
1359 |
1.29 |
|
|
|
| 23 |
A |
1447 |
1309 |
1.57 |
|
|
|
| 24 |
A |
1422 |
1286 |
15.45 |
|
|
|
| 25 |
A |
1386 |
1254 |
6.47 |
|
|
|
| 26 |
A |
1334 |
1206 |
6.98 |
|
|
|
| 27 |
A |
1274 |
1152 |
1.52 |
|
|
|
| 28 |
A |
1235 |
1117 |
2.95 |
|
|
|
| 29 |
A |
1174 |
1062 |
1.05 |
|
|
|
| 30 |
A |
1111 |
1005 |
1.42 |
|
|
|
| 31 |
A |
1099 |
994 |
1.15 |
|
|
|
| 32 |
A |
1059 |
958 |
3.04 |
|
|
|
| 33 |
A |
1006 |
910 |
0.77 |
|
|
|
| 34 |
A |
969 |
876 |
2.60 |
|
|
|
| 35 |
A |
933 |
844 |
0.83 |
|
|
|
| 36 |
A |
858 |
776 |
0.50 |
|
|
|
| 37 |
A |
835 |
755 |
7.47 |
|
|
|
| 38 |
A |
764 |
691 |
1.84 |
|
|
|
| 39 |
A |
487 |
440 |
0.13 |
|
|
|
| 40 |
A |
424 |
383 |
0.27 |
|
|
|
| 41 |
A |
407 |
368 |
0.89 |
|
|
|
| 42 |
A |
275 |
249 |
0.09 |
|
|
|
| 43 |
A |
257 |
233 |
0.47 |
|
|
|
| 44 |
A |
226 |
204 |
0.09 |
|
|
|
| 45 |
A |
215 |
194 |
1.47 |
|
|
|
| 46 |
A |
171 |
154 |
13.50 |
|
|
|
| 47 |
A |
97 |
88 |
0.72 |
|
|
|
| 48 |
A |
53 |
48 |
6.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37135.1 cm
-1
Scaled (by 0.9046) Zero Point Vibrational Energy (zpe) 33592.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-1.602 |
|
|
|
| 2 |
H |
0.462 |
|
|
|
| 3 |
H |
0.397 |
|
|
|
| 4 |
H |
0.336 |
|
|
|
| 5 |
S |
-0.538 |
|
|
|
| 6 |
H |
0.004 |
|
|
|
| 7 |
C |
-0.601 |
|
|
|
| 8 |
H |
0.299 |
|
|
|
| 9 |
H |
0.572 |
|
|
|
| 10 |
C |
0.395 |
|
|
|
| 11 |
H |
0.281 |
|
|
|
| 12 |
C |
-1.280 |
|
|
|
| 13 |
H |
0.244 |
|
|
|
| 14 |
H |
0.401 |
|
|
|
| 15 |
H |
0.308 |
|
|
|
| 16 |
C |
-0.449 |
|
|
|
| 17 |
H |
0.420 |
|
|
|
| 18 |
H |
0.352 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.526 |
-0.540 |
0.851 |
1.829 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.743 |
2.188 |
-1.772 |
| y |
2.188 |
-47.106 |
-1.819 |
| z |
-1.772 |
-1.819 |
-48.285 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.047 |
2.188 |
-1.772 |
| y |
2.188 |
2.407 |
-1.819 |
| z |
-1.772 |
-1.819 |
0.639 |
|
| Polar |
|---|
| 3z2-r2 | 1.279 |
|---|
| x2-y2 | -3.636 |
|---|
| xy | 2.188 |
|---|
| xz | -1.772 |
|---|
| yz | -1.819 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.847 |
0.148 |
0.148 |
| y |
0.148 |
11.622 |
0.056 |
| z |
0.148 |
0.056 |
10.320 |
<r2> (average value of r
2) Å
2
| <r2> |
287.921 |
| (<r2>)1/2 |
16.968 |