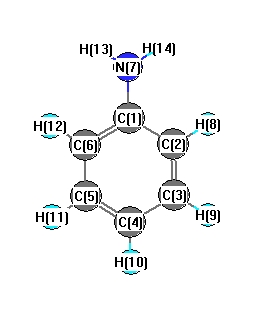
Vibrational Frequencies calculated at HF/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3784 |
3423 |
23.46 |
|
|
|
| 2 |
A' |
3348 |
3028 |
14.03 |
|
|
|
| 3 |
A' |
3323 |
3006 |
7.15 |
|
|
|
| 4 |
A' |
3305 |
2990 |
16.02 |
|
|
|
| 5 |
A' |
1804 |
1632 |
136.47 |
|
|
|
| 6 |
A' |
1778 |
1608 |
45.79 |
|
|
|
| 7 |
A' |
1653 |
1495 |
67.37 |
|
|
|
| 8 |
A' |
1377 |
1246 |
72.41 |
|
|
|
| 9 |
A' |
1285 |
1163 |
11.95 |
|
|
|
| 10 |
A' |
1117 |
1011 |
2.41 |
|
|
|
| 11 |
A' |
1113 |
1007 |
0.47 |
|
|
|
| 12 |
A' |
1078 |
975 |
2.52 |
|
|
|
| 13 |
A' |
995 |
900 |
13.79 |
|
|
|
| 14 |
A' |
884 |
800 |
9.37 |
|
|
|
| 15 |
A' |
847 |
766 |
88.32 |
|
|
|
| 16 |
A' |
772 |
698 |
27.48 |
|
|
|
| 17 |
A' |
670 |
606 |
224.84 |
|
|
|
| 18 |
A' |
572 |
517 |
2.67 |
|
|
|
| 19 |
A' |
553 |
500 |
76.33 |
|
|
|
| 20 |
A' |
243 |
220 |
6.31 |
|
|
|
| 21 |
A" |
3882 |
3511 |
19.96 |
|
|
|
| 22 |
A" |
3330 |
3012 |
40.39 |
|
|
|
| 23 |
A" |
3308 |
2992 |
2.26 |
|
|
|
| 24 |
A" |
1766 |
1598 |
10.88 |
|
|
|
| 25 |
A" |
1620 |
1465 |
2.86 |
|
|
|
| 26 |
A" |
1485 |
1343 |
0.06 |
|
|
|
| 27 |
A" |
1343 |
1215 |
3.94 |
|
|
|
| 28 |
A" |
1239 |
1121 |
1.72 |
|
|
|
| 29 |
A" |
1175 |
1063 |
16.89 |
|
|
|
| 30 |
A" |
1132 |
1024 |
0.49 |
|
|
|
| 31 |
A" |
1098 |
993 |
0.04 |
|
|
|
| 32 |
A" |
925 |
837 |
0.00 |
|
|
|
| 33 |
A" |
678 |
613 |
0.29 |
|
|
|
| 34 |
A" |
457 |
414 |
0.18 |
|
|
|
| 35 |
A" |
410 |
371 |
0.38 |
|
|
|
| 36 |
A" |
277 |
251 |
20.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27312.3 cm
-1
Scaled (by 0.9046) Zero Point Vibrational Energy (zpe) 24706.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.216 |
|
|
|
| 2 |
C |
-0.506 |
|
|
|
| 3 |
C |
-0.500 |
|
|
|
| 4 |
C |
-0.467 |
|
|
|
| 5 |
C |
-0.500 |
|
|
|
| 6 |
C |
-0.506 |
|
|
|
| 7 |
N |
-1.066 |
|
|
|
| 8 |
H |
0.418 |
|
|
|
| 9 |
H |
0.475 |
|
|
|
| 10 |
H |
0.481 |
|
|
|
| 11 |
H |
0.475 |
|
|
|
| 12 |
H |
0.418 |
|
|
|
| 13 |
H |
0.031 |
|
|
|
| 14 |
H |
0.031 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.016 |
1.008 |
0.000 |
1.431 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.360 |
-3.355 |
0.000 |
| y |
-3.355 |
-36.124 |
0.000 |
| z |
0.000 |
0.000 |
-36.743 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.927 |
-3.355 |
0.000 |
| y |
-3.355 |
5.927 |
0.000 |
| z |
0.000 |
0.000 |
4.999 |
|
| Polar |
|---|
| 3z2-r2 | 9.999 |
|---|
| x2-y2 | -11.236 |
|---|
| xy | -3.355 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.354 |
-0.039 |
0.000 |
| y |
-0.039 |
14.248 |
0.000 |
| z |
0.000 |
0.000 |
12.297 |
<r2> (average value of r
2) Å
2
| <r2> |
189.815 |
| (<r2>)1/2 |
13.777 |