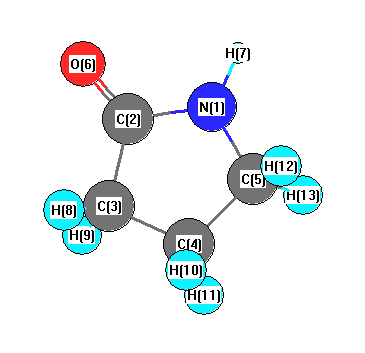
Vibrational Frequencies calculated at HF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3860 |
3505 |
50.53 |
|
|
|
| 2 |
A |
3281 |
2979 |
21.92 |
|
|
|
| 3 |
A |
3264 |
2963 |
33.43 |
|
|
|
| 4 |
A |
3241 |
2943 |
51.26 |
|
|
|
| 5 |
A |
3207 |
2912 |
18.83 |
|
|
|
| 6 |
A |
3203 |
2908 |
21.28 |
|
|
|
| 7 |
A |
3159 |
2868 |
76.39 |
|
|
|
| 8 |
A |
1993 |
1810 |
610.48 |
|
|
|
| 9 |
A |
1645 |
1493 |
3.90 |
|
|
|
| 10 |
A |
1605 |
1458 |
28.16 |
|
|
|
| 11 |
A |
1583 |
1437 |
45.97 |
|
|
|
| 12 |
A |
1566 |
1422 |
10.78 |
|
|
|
| 13 |
A |
1492 |
1355 |
15.95 |
|
|
|
| 14 |
A |
1461 |
1327 |
36.73 |
|
|
|
| 15 |
A |
1432 |
1300 |
43.23 |
|
|
|
| 16 |
A |
1370 |
1244 |
72.05 |
|
|
|
| 17 |
A |
1342 |
1218 |
8.33 |
|
|
|
| 18 |
A |
1314 |
1194 |
0.49 |
|
|
|
| 19 |
A |
1278 |
1160 |
6.33 |
|
|
|
| 20 |
A |
1182 |
1073 |
0.51 |
|
|
|
| 21 |
A |
1160 |
1053 |
20.58 |
|
|
|
| 22 |
A |
1067 |
969 |
11.47 |
|
|
|
| 23 |
A |
985 |
894 |
2.32 |
|
|
|
| 24 |
A |
975 |
886 |
3.20 |
|
|
|
| 25 |
A |
956 |
868 |
4.64 |
|
|
|
| 26 |
A |
884 |
802 |
6.67 |
|
|
|
| 27 |
A |
747 |
678 |
14.49 |
|
|
|
| 28 |
A |
692 |
629 |
23.95 |
|
|
|
| 29 |
A |
609 |
553 |
46.72 |
|
|
|
| 30 |
A |
529 |
480 |
68.29 |
|
|
|
| 31 |
A |
507 |
460 |
19.38 |
|
|
|
| 32 |
A |
214 |
194 |
5.48 |
|
|
|
| 33 |
A |
148 |
134 |
1.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25974.0 cm
-1
Scaled (by 0.908) Zero Point Vibrational Energy (zpe) 23584.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.346 |
-0.641 |
|
|
| 2 |
C |
0.332 |
0.347 |
|
|
| 3 |
C |
-0.074 |
-0.621 |
|
|
| 4 |
C |
-0.073 |
0.277 |
|
|
| 5 |
C |
0.149 |
0.032 |
|
|
| 6 |
O |
-0.384 |
0.731 |
|
|
| 7 |
H |
0.114 |
-0.083 |
|
|
| 8 |
H |
0.064 |
0.053 |
|
|
| 9 |
H |
0.062 |
-0.017 |
|
|
| 10 |
H |
0.037 |
-0.007 |
|
|
| 11 |
H |
0.047 |
-0.041 |
|
|
| 12 |
H |
0.034 |
-0.039 |
|
|
| 13 |
H |
0.040 |
0.009 |
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.088 |
-0.507 |
0.439 |
4.143 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.311 |
-0.216 |
0.170 |
| y |
-0.216 |
-31.725 |
-0.330 |
| z |
0.170 |
-0.330 |
-35.495 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.701 |
-0.216 |
0.170 |
| y |
-0.216 |
8.178 |
-0.330 |
| z |
0.170 |
-0.330 |
2.523 |
|
| Polar |
|---|
| 3z2-r2 | 5.045 |
|---|
| x2-y2 | -12.586 |
|---|
| xy | -0.216 |
|---|
| xz | 0.170 |
|---|
| yz | -0.330 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.987 |
0.332 |
-0.046 |
| y |
0.332 |
7.042 |
0.007 |
| z |
-0.046 |
0.007 |
5.456 |
<r2> (average value of r
2) Å
2
| <r2> |
143.185 |
| (<r2>)1/2 |
11.966 |