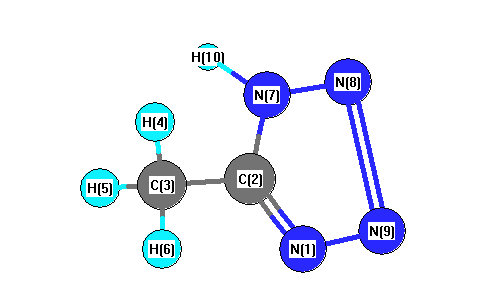
Vibrational Frequencies calculated at HF/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3906 |
3555 |
115.87 |
|
|
|
| 2 |
A |
3285 |
2990 |
4.03 |
|
|
|
| 3 |
A |
3179 |
2893 |
14.38 |
|
|
|
| 4 |
A |
1772 |
1612 |
66.20 |
|
|
|
| 5 |
A |
1624 |
1478 |
2.36 |
|
|
|
| 6 |
A |
1605 |
1461 |
18.76 |
|
|
|
| 7 |
A |
1559 |
1419 |
31.12 |
|
|
|
| 8 |
A |
1542 |
1403 |
4.27 |
|
|
|
| 9 |
A |
1448 |
1318 |
13.11 |
|
|
|
| 10 |
A |
1222 |
1112 |
19.18 |
|
|
|
| 11 |
A |
1198 |
1090 |
0.47 |
|
|
|
| 12 |
A |
1178 |
1072 |
28.25 |
|
|
|
| 13 |
A |
1119 |
1019 |
1.10 |
|
|
|
| 14 |
A |
1078 |
981 |
3.70 |
|
|
|
| 15 |
A |
728 |
663 |
4.65 |
|
|
|
| 16 |
A |
366 |
333 |
4.42 |
|
|
|
| 17 |
A |
3233 |
2942 |
14.11 |
|
|
|
| 18 |
A |
1603 |
1459 |
7.87 |
|
|
|
| 19 |
A |
1172 |
1066 |
1.56 |
|
|
|
| 20 |
A |
817 |
744 |
4.63 |
|
|
|
| 21 |
A |
775 |
705 |
6.77 |
|
|
|
| 22 |
A |
611 |
556 |
92.84 |
|
|
|
| 23 |
A |
303 |
275 |
0.00 |
|
|
|
| 24 |
A |
87 |
80 |
0.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17703.8 cm
-1
Scaled (by 0.9101) Zero Point Vibrational Energy (zpe) 16112.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.187 |
-0.461 |
-0.908 |
-0.468 |
| 2 |
C |
0.112 |
0.588 |
1.335 |
0.606 |
| 3 |
C |
-0.276 |
-0.462 |
0.204 |
-0.556 |
| 4 |
H |
0.111 |
0.122 |
-0.011 |
0.149 |
| 5 |
H |
0.111 |
0.122 |
-0.011 |
0.149 |
| 6 |
H |
0.139 |
0.154 |
0.028 |
0.189 |
| 7 |
N |
-0.072 |
0.035 |
-1.059 |
-0.017 |
| 8 |
N |
-0.083 |
-0.271 |
-0.057 |
-0.264 |
| 9 |
N |
-0.069 |
-0.034 |
-0.040 |
-0.033 |
| 10 |
H |
0.214 |
0.208 |
0.504 |
0.244 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.275 |
4.046 |
0.000 |
5.886 |
| CHELPG |
|
|
|
|
| AIM |
-0.158 |
-1.514 |
0.000 |
1.522 |
| ESP |
4.180 |
3.930 |
0.000 |
5.737 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.248 |
-0.360 |
0.000 |
| y |
-0.360 |
-36.700 |
0.000 |
| z |
0.000 |
0.000 |
-34.497 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.350 |
-0.360 |
0.000 |
| y |
-0.360 |
-2.327 |
0.000 |
| z |
0.000 |
0.000 |
0.977 |
|
| Polar |
|---|
| 3z2-r2 | 1.953 |
|---|
| x2-y2 | 2.451 |
|---|
| xy | -0.360 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.903 |
0.216 |
0.000 |
| y |
0.216 |
7.907 |
0.000 |
| z |
0.000 |
0.000 |
4.412 |
<r2> (average value of r
2) Å
2
| <r2> |
125.836 |
| (<r2>)1/2 |
11.218 |