Jump to
S1C2
Vibrational Frequencies calculated at QCISD(T)/TZVP
Geometric Data calculated at QCISD(T)/TZVP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/TZVP
| | hartrees |
|---|
| Energy at 0K | -553.274356 |
| Energy at 298.15K | -553.279700 |
| HF Energy | -552.491121 |
| Nuclear repulsion energy | 201.172725 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3260 |
3140 |
|
|
|
|
| 2 |
A |
3255 |
3135 |
|
|
|
|
| 3 |
A |
3194 |
3076 |
|
|
|
|
| 4 |
A |
3181 |
3064 |
|
|
|
|
| 5 |
A |
3167 |
3050 |
|
|
|
|
| 6 |
A |
3159 |
3043 |
|
|
|
|
| 7 |
A |
1643 |
1582 |
|
|
|
|
| 8 |
A |
1634 |
1574 |
|
|
|
|
| 9 |
A |
1434 |
1381 |
|
|
|
|
| 10 |
A |
1431 |
1378 |
|
|
|
|
| 11 |
A |
1310 |
1262 |
|
|
|
|
| 12 |
A |
1305 |
1257 |
|
|
|
|
| 13 |
A |
1062 |
1023 |
|
|
|
|
| 14 |
A |
1037 |
999 |
|
|
|
|
| 15 |
A |
975 |
939 |
|
|
|
|
| 16 |
A |
963 |
927 |
|
|
|
|
| 17 |
A |
827 |
797 |
|
|
|
|
| 18 |
A |
809 |
779 |
|
|
|
|
| 19 |
A |
736 |
709 |
|
|
|
|
| 20 |
A |
690 |
665 |
|
|
|
|
| 21 |
A |
619 |
596 |
|
|
|
|
| 22 |
A |
578 |
557 |
|
|
|
|
| 23 |
A |
453 |
436 |
|
|
|
|
| 24 |
A |
368 |
355 |
|
|
|
|
| 25 |
A |
225 |
217 |
|
|
|
|
| 26 |
A |
132 |
127 |
|
|
|
|
| 27 |
A |
83 |
80 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18764.2 cm
-1
Scaled (by 0.9632) Zero Point Vibrational Energy (zpe) 18073.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/TZVP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.012 |
-1.001 |
0.000 |
| C2 |
1.078 |
0.371 |
0.387 |
| C3 |
-1.570 |
-0.187 |
-0.029 |
| C4 |
2.252 |
0.564 |
-0.227 |
| C5 |
-1.800 |
1.130 |
-0.126 |
| H6 |
0.762 |
0.983 |
1.227 |
| H7 |
-2.390 |
-0.893 |
0.057 |
| H8 |
2.927 |
1.345 |
0.107 |
| H9 |
2.562 |
-0.045 |
-1.069 |
| H10 |
-1.002 |
1.854 |
-0.239 |
| H11 |
-2.818 |
1.503 |
-0.112 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7803 | 1.7794 | 2.7425 | 2.8005 | 2.4504 | 2.4050 | 3.7433 | 2.9258 | 3.0398 | 3.7802 |
C2 | 1.7803 | | 2.7374 | 1.3389 | 3.0202 | 1.0864 | 3.7055 | 2.1082 | 2.1197 | 2.6299 | 4.0870 | C3 | 1.7794 | 2.7374 | | 3.9000 | 1.3403 | 2.8946 | 1.0856 | 4.7521 | 4.2628 | 2.1289 | 2.1020 | C4 | 2.7425 | 1.3389 | 3.9000 | | 4.0928 | 2.1238 | 4.8736 | 1.0844 | 1.0842 | 3.5005 | 5.1573 | C5 | 2.8005 | 3.0202 | 1.3403 | 4.0928 | | 2.9007 | 2.1149 | 4.7375 | 4.6147 | 1.0836 | 1.0838 | H6 | 2.4504 | 1.0864 | 2.8946 | 2.1238 | 2.9007 | | 3.8493 | 2.4642 | 3.0930 | 2.4530 | 3.8565 | H7 | 2.4050 | 3.7055 | 1.0856 | 4.8736 | 2.1149 | 3.8493 | | 5.7685 | 5.1486 | 3.0919 | 2.4390 | H8 | 3.7433 | 2.1082 | 4.7521 | 1.0844 | 4.7375 | 2.4642 | 5.7685 | | 1.8566 | 3.9768 | 5.7508 | H9 | 2.9258 | 2.1197 | 4.2628 | 1.0842 | 4.6147 | 3.0930 | 5.1486 | 1.8566 | | 4.1226 | 5.6789 | H10 | 3.0398 | 2.6299 | 2.1289 | 3.5005 | 1.0836 | 2.4530 | 3.0919 | 3.9768 | 4.1226 | | 1.8538 | H11 | 3.7802 | 4.0870 | 2.1020 | 5.1573 | 1.0838 | 3.8565 | 2.4390 | 5.7508 | 5.6789 | 1.8538 | |
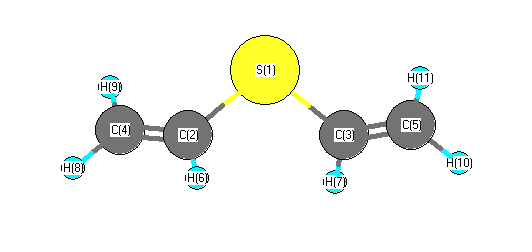 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.471 |
|
S1 |
C2 |
H6 |
115.322 |
| S1 |
C3 |
C5 |
127.143 |
|
S1 |
C3 |
H7 |
111.875 |
| C2 |
S1 |
C3 |
100.526 |
|
C2 |
C4 |
H8 |
120.545 |
| C2 |
C4 |
H9 |
121.690 |
|
C3 |
C5 |
H10 |
122.518 |
| C3 |
C5 |
H11 |
119.879 |
|
C4 |
C2 |
H6 |
121.904 |
| C5 |
C3 |
H7 |
120.977 |
|
H8 |
C4 |
H9 |
117.765 |
| H10 |
C5 |
H11 |
117.594 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
