Jump to
S1C2
Energy calculated at CCSD/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -191.385706 |
| Energy at 298.15K | |
| HF Energy | -190.774747 |
| Nuclear repulsion energy | 102.643538 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3315 |
3096 |
7.39 |
|
|
|
| 2 |
A' |
3262 |
3046 |
1.38 |
|
|
|
| 3 |
A' |
3217 |
3004 |
5.77 |
|
|
|
| 4 |
A' |
3019 |
2819 |
75.09 |
|
|
|
| 5 |
A' |
1807 |
1688 |
196.54 |
|
|
|
| 6 |
A' |
1713 |
1599 |
8.35 |
|
|
|
| 7 |
A' |
1498 |
1398 |
6.60 |
|
|
|
| 8 |
A' |
1426 |
1332 |
2.61 |
|
|
|
| 9 |
A' |
1327 |
1240 |
1.86 |
|
|
|
| 10 |
A' |
1206 |
1126 |
34.55 |
|
|
|
| 11 |
A' |
945 |
882 |
18.26 |
|
|
|
| 12 |
A' |
576 |
538 |
4.76 |
|
|
|
| 13 |
A' |
325 |
304 |
11.46 |
|
|
|
| 14 |
A" |
1027 |
959 |
7.87 |
|
|
|
| 15 |
A" |
1015 |
948 |
28.05 |
|
|
|
| 16 |
A" |
973 |
909 |
32.29 |
|
|
|
| 17 |
A" |
596 |
557 |
12.58 |
|
|
|
| 18 |
A" |
151 |
141 |
5.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13699.9 cm
-1
Scaled (by 0.9338) Zero Point Vibrational Energy (zpe) 12793.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.145 |
-0.750 |
0.000 |
| C2 |
0.000 |
0.722 |
0.000 |
| C3 |
1.216 |
1.292 |
0.000 |
| O4 |
-1.222 |
-1.327 |
0.000 |
| H5 |
0.800 |
-1.322 |
0.000 |
| H6 |
-0.914 |
1.303 |
0.000 |
| H7 |
1.345 |
2.365 |
0.000 |
| H8 |
2.117 |
0.690 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4792 | 2.4539 | 1.2217 | 1.1041 | 2.1923 | 3.4536 | 2.6817 |
C2 | 1.4792 | | 1.3427 | 2.3858 | 2.1945 | 1.0831 | 2.1238 | 2.1174 | C3 | 2.4539 | 1.3427 | | 3.5779 | 2.6465 | 2.1300 | 1.0812 | 1.0839 | O4 | 1.2217 | 2.3858 | 3.5779 | | 2.0215 | 2.6479 | 4.4972 | 3.9010 | H5 | 1.1041 | 2.1945 | 2.6465 | 2.0215 | | 3.1345 | 3.7272 | 2.4046 | H6 | 2.1923 | 1.0831 | 2.1300 | 2.6479 | 3.1345 | | 2.4969 | 3.0927 | H7 | 3.4536 | 2.1238 | 1.0812 | 4.4972 | 3.7272 | 2.4969 | | 1.8447 | H8 | 2.6817 | 2.1174 | 1.0839 | 3.9010 | 2.4046 | 3.0927 | 1.8447 | |
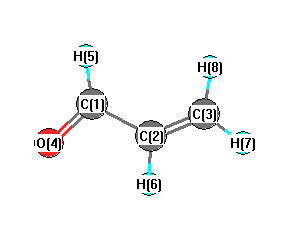 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.745 |
|
C1 |
C2 |
H6 |
116.798 |
| C2 |
C1 |
O4 |
123.816 |
|
C2 |
C1 |
H5 |
115.545 |
| C2 |
C3 |
H7 |
121.999 |
|
C2 |
C3 |
H8 |
121.147 |
| C3 |
C2 |
H6 |
122.457 |
|
O4 |
C1 |
H5 |
120.639 |
| H7 |
C3 |
H8 |
116.855 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/6-31+G**
| | hartrees |
|---|
| Energy at 0K | -191.382724 |
| Energy at 298.15K | |
| HF Energy | -190.771545 |
| Nuclear repulsion energy | 104.199810 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/6-31+G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3328 |
3107 |
3.24 |
|
|
|
| 2 |
A' |
3247 |
3032 |
10.74 |
|
|
|
| 3 |
A' |
3226 |
3013 |
6.02 |
|
|
|
| 4 |
A' |
3038 |
2837 |
112.97 |
|
|
|
| 5 |
A' |
1802 |
1683 |
106.25 |
|
|
|
| 6 |
A' |
1715 |
1601 |
42.86 |
|
|
|
| 7 |
A' |
1480 |
1382 |
31.73 |
|
|
|
| 8 |
A' |
1462 |
1365 |
2.30 |
|
|
|
| 9 |
A' |
1339 |
1251 |
1.83 |
|
|
|
| 10 |
A' |
1095 |
1022 |
3.76 |
|
|
|
| 11 |
A' |
956 |
893 |
56.53 |
|
|
|
| 12 |
A' |
689 |
644 |
13.95 |
|
|
|
| 13 |
A' |
291 |
272 |
7.06 |
|
|
|
| 14 |
A" |
1023 |
955 |
0.12 |
|
|
|
| 15 |
A" |
1014 |
947 |
53.41 |
|
|
|
| 16 |
A" |
977 |
912 |
7.02 |
|
|
|
| 17 |
A" |
551 |
514 |
12.50 |
|
|
|
| 18 |
A" |
121 |
113 |
7.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13676.4 cm
-1
Scaled (by 0.9338) Zero Point Vibrational Energy (zpe) 12771.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/6-31+G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.895 |
-0.289 |
0.000 |
| C2 |
0.000 |
0.901 |
0.000 |
| C3 |
1.337 |
0.775 |
0.000 |
| O4 |
-0.497 |
-1.444 |
0.000 |
| H5 |
-1.977 |
-0.078 |
0.000 |
| H6 |
-0.478 |
1.874 |
0.000 |
| H7 |
1.986 |
1.640 |
0.000 |
| H8 |
1.795 |
-0.205 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4888 | 2.4722 | 1.2221 | 1.1020 | 2.2028 | 3.4668 | 2.6912 |
C2 | 1.4888 | | 1.3425 | 2.3974 | 2.2056 | 1.0841 | 2.1189 | 2.1083 | C3 | 2.4722 | 1.3425 | | 2.8791 | 3.4212 | 2.1212 | 1.0812 | 1.0822 | O4 | 1.2221 | 2.3974 | 2.8791 | | 2.0140 | 3.3186 | 3.9595 | 2.6056 | H5 | 1.1020 | 2.2056 | 3.4212 | 2.0140 | | 2.4610 | 4.3187 | 3.7736 | H6 | 2.2028 | 1.0841 | 2.1212 | 3.3186 | 2.4610 | | 2.4749 | 3.0803 | H7 | 3.4668 | 2.1189 | 1.0812 | 3.9595 | 4.3187 | 2.4749 | | 1.8547 | H8 | 2.6912 | 2.1083 | 1.0822 | 2.6056 | 3.7736 | 3.0803 | 1.8547 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.577 |
|
C1 |
C2 |
H6 |
116.902 |
| C2 |
C1 |
O4 |
124.052 |
|
C2 |
C1 |
H5 |
115.911 |
| C2 |
C3 |
H7 |
121.540 |
|
C2 |
C3 |
H8 |
120.431 |
| C3 |
C2 |
H6 |
121.521 |
|
O4 |
C1 |
H5 |
120.037 |
| H7 |
C3 |
H8 |
118.029 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
