Jump to
S1C2
S1C3
Energy calculated at CCSD/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -100.155716 |
| Energy at 298.15K | -100.154773 |
| HF Energy | -99.808796 |
| Nuclear repulsion energy | 27.595329 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.076 |
| C2 |
0.000 |
0.000 |
-0.149 |
| N3 |
0.000 |
0.000 |
1.017 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9270 | 3.0931 |
C2 | 1.9270 | | 1.1661 | N3 | 3.0931 | 1.1661 | |
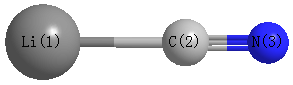 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCSD/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -100.159982 |
| Energy at 298.15K | -100.159208 |
| HF Energy | -99.815598 |
| Nuclear repulsion energy | 29.238816 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.444 |
-0.588 |
0.000 |
| C2 |
-0.722 |
-0.366 |
0.000 |
| N3 |
0.000 |
0.566 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.1777 | 1.8489 |
C2 | 2.1777 | | 1.1788 | N3 | 1.8489 | 1.1788 | |
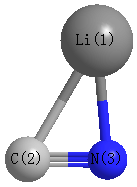 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
58.091 |
|
Li1 |
N3 |
C2 |
89.141 |
| C2 |
Li1 |
N3 |
32.768 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCSD/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -100.160162 |
| Energy at 298.15K | -100.159001 |
| HF Energy | -99.819470 |
| Nuclear repulsion energy | 28.357022 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.887 |
| C2 |
0.000 |
0.000 |
-1.069 |
| N3 |
0.000 |
0.000 |
0.108 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9559 | 1.7793 |
C2 | 2.9559 | | 1.1766 | N3 | 1.7793 | 1.1766 | |
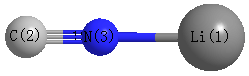 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
