return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SO2NH2 (methanesulfonamide)

using model chemistry: CCSD/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCSD/cc-pVTZ
 hartrees
Energy at 0K-643.628291
Energy at 298.15K-643.636211
HF Energy-642.557021
Nuclear repulsion energy279.734354
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3565 3355 39.46      
2 A' 3196 3008 0.26      
3 A' 3092 2910 0.20      
4 A' 1615 1520 36.91      
5 A' 1477 1390 4.69      
6 A' 1377 1296 17.10      
7 A' 1200 1130 168.46      
8 A' 1004 945 35.52      
9 A' 916 862 105.24      
10 A' 760 716 13.37      
11 A' 698 657 215.07      
12 A' 517 487 52.46      
13 A' 495 466 16.50      
14 A' 303 285 4.72      
15 A" 3671 3455 51.14      
16 A" 3205 3017 0.51      
17 A" 1479 1392 0.27      
18 A" 1419 1335 246.12      
19 A" 1120 1054 2.12      
20 A" 987 929 0.96      
21 A" 409 385 0.04      
22 A" 338 319 2.97      
23 A" 237 224 2.61      
24 A" 182 171 34.02      

Unscaled Zero Point Vibrational Energy (zpe) 16631.2 cm-1
Scaled (by 0.9412) Zero Point Vibrational Energy (zpe) 15653.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD/cc-pVTZ
ABC
0.15953 0.14727 0.14518

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.663 -0.059 0.000
S2 0.107 -0.131 0.000
N3 0.528 1.479 0.000
O4 0.528 -0.694 1.258
O5 0.528 -0.694 -1.258
H6 -2.015 -1.087 0.000
H7 -1.990 0.459 0.897
H8 -1.990 0.459 -0.897
H9 1.058 1.689 0.834
H10 1.058 1.689 -0.834

Atom - Atom Distances (Å)
  C1 S2 N3 O4 O5 H6 H7 H8 H9 H10
C11.77182.67702.60552.60551.08721.08581.08583.33983.3398
S21.77181.66451.44121.44122.32772.35572.35572.21672.2167
N32.67701.66452.51112.51113.61312.86052.86051.01011.0101
O42.60551.44122.51112.51632.86462.79303.50902.47823.2154
O52.60551.44122.51112.51632.86463.50902.79303.21542.4782
H61.08722.32773.61312.86462.86461.78801.78804.22474.2247
H71.08582.35572.86052.79303.50901.78801.79313.28673.7140
H81.08582.35572.86053.50902.79301.78801.79313.71403.2867
H93.33982.21671.01012.47823.21544.22473.28673.71401.6681
H103.33982.21671.01013.21542.47824.22473.71403.28671.6681

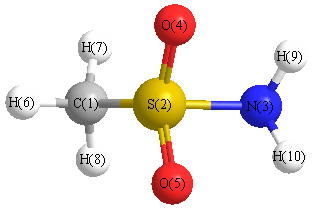
picture of methanesulfonamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 N3 102.302 C1 S2 O4 107.933
C1 S2 O5 107.933 S2 C1 H6 106.541
S2 C1 H7 108.662 S2 C1 H8 108.662
S2 N3 H9 109.512 S2 N3 H10 109.512
N3 S2 O4 107.696 N3 S2 O5 107.696
O4 S2 O5 121.617 H6 C1 H7 110.742
H6 C1 H8 110.742 H7 C1 H8 111.330
H9 N3 H10 111.324
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability