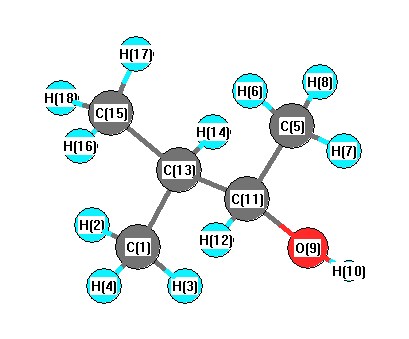
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3301 |
3283 |
8.71 |
|
|
|
| 2 |
A |
3081 |
3064 |
10.79 |
|
|
|
| 3 |
A |
3055 |
3038 |
28.29 |
|
|
|
| 4 |
A |
3038 |
3021 |
40.12 |
|
|
|
| 5 |
A |
3030 |
3013 |
78.45 |
|
|
|
| 6 |
A |
3025 |
3008 |
17.53 |
|
|
|
| 7 |
A |
3020 |
3003 |
4.91 |
|
|
|
| 8 |
A |
2974 |
2958 |
42.89 |
|
|
|
| 9 |
A |
2969 |
2953 |
20.99 |
|
|
|
| 10 |
A |
2966 |
2950 |
9.68 |
|
|
|
| 11 |
A |
2964 |
2948 |
15.83 |
|
|
|
| 12 |
A |
2883 |
2867 |
48.16 |
|
|
|
| 13 |
A |
1548 |
1539 |
9.56 |
|
|
|
| 14 |
A |
1540 |
1531 |
1.72 |
|
|
|
| 15 |
A |
1528 |
1520 |
11.01 |
|
|
|
| 16 |
A |
1525 |
1517 |
5.62 |
|
|
|
| 17 |
A |
1522 |
1513 |
1.78 |
|
|
|
| 18 |
A |
1518 |
1509 |
4.77 |
|
|
|
| 19 |
A |
1432 |
1424 |
4.72 |
|
|
|
| 20 |
A |
1415 |
1407 |
14.31 |
|
|
|
| 21 |
A |
1407 |
1400 |
12.88 |
|
|
|
| 22 |
A |
1384 |
1377 |
0.78 |
|
|
|
| 23 |
A |
1349 |
1342 |
0.56 |
|
|
|
| 24 |
A |
1342 |
1334 |
12.24 |
|
|
|
| 25 |
A |
1329 |
1322 |
0.47 |
|
|
|
| 26 |
A |
1255 |
1248 |
25.33 |
|
|
|
| 27 |
A |
1184 |
1178 |
7.55 |
|
|
|
| 28 |
A |
1156 |
1150 |
14.61 |
|
|
|
| 29 |
A |
1113 |
1107 |
2.29 |
|
|
|
| 30 |
A |
1071 |
1066 |
37.94 |
|
|
|
| 31 |
A |
1017 |
1012 |
18.86 |
|
|
|
| 32 |
A |
992 |
986 |
12.26 |
|
|
|
| 33 |
A |
957 |
952 |
1.68 |
|
|
|
| 34 |
A |
940 |
935 |
0.43 |
|
|
|
| 35 |
A |
899 |
894 |
22.02 |
|
|
|
| 36 |
A |
848 |
843 |
11.14 |
|
|
|
| 37 |
A |
764 |
760 |
2.91 |
|
|
|
| 38 |
A |
495 |
492 |
2.82 |
|
|
|
| 39 |
A |
443 |
441 |
7.34 |
|
|
|
| 40 |
A |
391 |
388 |
0.96 |
|
|
|
| 41 |
A |
354 |
352 |
7.51 |
|
|
|
| 42 |
A |
351 |
349 |
3.94 |
|
|
|
| 43 |
A |
275 |
273 |
9.97 |
|
|
|
| 44 |
A |
269 |
267 |
95.21 |
|
|
|
| 45 |
A |
234 |
233 |
1.51 |
|
|
|
| 46 |
A |
224 |
223 |
0.38 |
|
|
|
| 47 |
A |
218 |
217 |
0.29 |
|
|
|
| 48 |
A |
82 |
82 |
1.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35337.0 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 35142.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.500 |
|
|
|
| 2 |
H |
0.162 |
|
|
|
| 3 |
H |
0.200 |
|
|
|
| 4 |
H |
0.160 |
|
|
|
| 5 |
C |
-0.533 |
|
|
|
| 6 |
H |
0.177 |
|
|
|
| 7 |
H |
0.163 |
|
|
|
| 8 |
H |
0.189 |
|
|
|
| 9 |
O |
-0.502 |
|
|
|
| 10 |
H |
0.312 |
|
|
|
| 11 |
C |
0.041 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
C |
-0.209 |
|
|
|
| 14 |
H |
0.179 |
|
|
|
| 15 |
C |
-0.510 |
|
|
|
| 16 |
H |
0.169 |
|
|
|
| 17 |
H |
0.174 |
|
|
|
| 18 |
H |
0.178 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.941 |
0.886 |
0.875 |
1.561 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.172 |
-1.018 |
2.012 |
| y |
-1.018 |
-41.532 |
-1.321 |
| z |
2.012 |
-1.321 |
-39.621 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.404 |
-1.018 |
2.012 |
| y |
-1.018 |
-3.635 |
-1.321 |
| z |
2.012 |
-1.321 |
-0.769 |
|
| Polar |
|---|
| 3z2-r2 | -1.538 |
|---|
| x2-y2 | 5.359 |
|---|
| xy | -1.018 |
|---|
| xz | 2.012 |
|---|
| yz | -1.321 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
199.714 |
| (<r2>)1/2 |
14.132 |