Jump to
S1C2
Energy calculated at BLYP/STO-3G
| | hartrees |
|---|
| Energy at 0K | -1060.827609 |
| Energy at 298.15K | -1060.829514 |
| HF Energy | -1060.827609 |
| Nuclear repulsion energy | 250.247003 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3257 |
3013 |
1.37 |
|
|
|
| 2 |
A' |
3096 |
2865 |
21.58 |
|
|
|
| 3 |
A' |
1724 |
1595 |
22.25 |
|
|
|
| 4 |
A' |
1390 |
1286 |
6.94 |
|
|
|
| 5 |
A' |
1211 |
1121 |
12.69 |
|
|
|
| 6 |
A' |
957 |
886 |
13.38 |
|
|
|
| 7 |
A' |
759 |
702 |
25.47 |
|
|
|
| 8 |
A' |
404 |
374 |
1.73 |
|
|
|
| 9 |
A' |
283 |
261 |
19.35 |
|
|
|
| 10 |
A' |
226 |
209 |
5.06 |
|
|
|
| 11 |
A" |
1228 |
1136 |
17.67 |
|
|
|
| 12 |
A" |
912 |
844 |
43.40 |
|
|
|
| 13 |
A" |
750 |
694 |
69.85 |
|
|
|
| 14 |
A" |
227 |
210 |
2.93 |
|
|
|
| 15 |
A" |
59 |
55 |
3.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8241.6 cm
-1
Scaled (by 0.9252) Zero Point Vibrational Energy (zpe) 7625.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.395 |
-0.013 |
0.000 |
| C2 |
-0.180 |
1.483 |
0.000 |
| H3 |
1.514 |
-0.038 |
0.000 |
| Cl4 |
-0.180 |
-0.889 |
1.553 |
| Cl5 |
-0.180 |
-0.889 |
-1.553 |
| O6 |
0.579 |
2.492 |
0.000 |
| H7 |
-1.313 |
1.511 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.6023 | 1.1200 | 1.8740 | 1.8740 | 2.5114 | 2.2888 |
C2 | 1.6023 | | 2.2771 | 2.8357 | 2.8357 | 1.2625 | 1.1336 | H3 | 1.1200 | 2.2771 | | 2.4513 | 2.4513 | 2.6976 | 3.2243 | Cl4 | 1.8740 | 2.8357 | 2.4513 | | 3.1069 | 3.7976 | 3.0754 | Cl5 | 1.8740 | 2.8357 | 2.4513 | 3.1069 | | 3.7976 | 3.0754 | O6 | 2.5114 | 1.2625 | 2.6976 | 3.7976 | 3.7976 | | 2.1314 | H7 | 2.2888 | 1.1336 | 3.2243 | 3.0754 | 3.0754 | 2.1314 | |
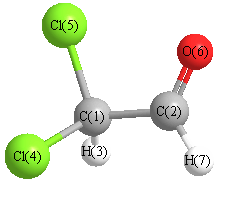 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.031 |
|
C1 |
C2 |
H7 |
112.428 |
| C2 |
C1 |
H3 |
112.326 |
|
C2 |
C1 |
Cl4 |
109.064 |
| C2 |
C1 |
Cl5 |
109.064 |
|
H3 |
C1 |
Cl4 |
107.215 |
| H3 |
C1 |
Cl5 |
107.215 |
|
Cl4 |
C1 |
Cl5 |
111.979 |
| O6 |
C2 |
H7 |
125.541 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.012 |
|
|
|
| 2 |
C |
0.104 |
|
|
|
| 3 |
H |
0.135 |
|
|
|
| 4 |
Cl |
-0.103 |
|
|
|
| 5 |
Cl |
-0.103 |
|
|
|
| 6 |
O |
-0.113 |
|
|
|
| 7 |
H |
0.092 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.322 |
0.988 |
0.000 |
1.039 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.558 |
-2.068 |
0.000 |
| y |
-2.068 |
-42.576 |
0.000 |
| z |
0.000 |
0.000 |
-41.737 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.598 |
-2.068 |
0.000 |
| y |
-2.068 |
-2.928 |
0.000 |
| z |
0.000 |
0.000 |
-1.670 |
|
| Polar |
|---|
| 3z2-r2 | -3.340 |
|---|
| x2-y2 | 5.017 |
|---|
| xy | -2.068 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.254 |
0.752 |
0.000 |
| y |
0.752 |
4.155 |
0.000 |
| z |
0.000 |
0.000 |
4.555 |
<r2> (average value of r
2) Å
2
| <r2> |
208.402 |
| (<r2>)1/2 |
14.436 |
Jump to
S1C1
Energy calculated at BLYP/STO-3G
| | hartrees |
|---|
| Energy at 0K | -1060.825455 |
| Energy at 298.15K | |
| HF Energy | -1060.825455 |
| Nuclear repulsion energy | 253.474047 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3224 |
2983 |
0.37 |
52.34 |
0.26 |
0.41 |
| 2 |
A |
3063 |
2834 |
21.94 |
75.62 |
0.37 |
0.54 |
| 3 |
A |
1742 |
1611 |
28.82 |
9.62 |
0.17 |
0.30 |
| 4 |
A |
1380 |
1276 |
26.75 |
9.47 |
0.72 |
0.84 |
| 5 |
A |
1233 |
1141 |
5.98 |
6.84 |
0.64 |
0.78 |
| 6 |
A |
1210 |
1120 |
9.27 |
10.77 |
0.63 |
0.78 |
| 7 |
A |
937 |
867 |
25.37 |
1.10 |
0.62 |
0.77 |
| 8 |
A |
866 |
801 |
8.80 |
5.37 |
0.54 |
0.70 |
| 9 |
A |
809 |
749 |
62.76 |
8.15 |
0.57 |
0.72 |
| 10 |
A |
626 |
580 |
20.02 |
13.02 |
0.05 |
0.09 |
| 11 |
A |
585 |
542 |
29.24 |
5.65 |
0.75 |
0.86 |
| 12 |
A |
297 |
275 |
1.69 |
4.10 |
0.24 |
0.39 |
| 13 |
A |
235 |
217 |
5.53 |
4.76 |
0.73 |
0.85 |
| 14 |
A |
173 |
160 |
0.62 |
1.66 |
0.68 |
0.81 |
| 15 |
A |
67 |
62 |
4.94 |
1.45 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 8222.4 cm
-1
Scaled (by 0.9252) Zero Point Vibrational Energy (zpe) 7607.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.117 |
-0.030 |
0.557 |
| C2 |
-0.740 |
-1.356 |
0.232 |
| H3 |
0.276 |
0.097 |
1.660 |
| Cl4 |
1.831 |
-0.243 |
-0.185 |
| Cl5 |
-0.745 |
1.498 |
-0.057 |
| O6 |
-1.850 |
-1.355 |
-0.360 |
| H7 |
-0.195 |
-2.281 |
0.604 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.6110 | 1.1220 | 1.8796 | 1.8595 | 2.5432 | 2.2726 |
C2 | 1.6110 | | 2.2766 | 2.8316 | 2.8687 | 1.2590 | 1.1362 | H3 | 1.1220 | 2.2766 | | 2.4367 | 2.4407 | 3.2736 | 2.6446 | Cl4 | 1.8796 | 2.8316 | 2.4367 | | 3.1115 | 3.8495 | 2.9802 | Cl5 | 1.8595 | 2.8687 | 2.4407 | 3.1115 | | 3.0755 | 3.8761 | O6 | 2.5432 | 1.2590 | 3.2736 | 3.8495 | 3.0755 | | 2.1276 | H7 | 2.2726 | 1.1362 | 2.6446 | 2.9802 | 3.8761 | 2.1276 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.325 |
|
C1 |
C2 |
H7 |
110.438 |
| C2 |
C1 |
H3 |
111.562 |
|
C2 |
C1 |
Cl4 |
108.187 |
| C2 |
C1 |
Cl5 |
111.303 |
|
H3 |
C1 |
Cl4 |
105.762 |
| H3 |
C1 |
Cl5 |
107.292 |
|
Cl4 |
C1 |
Cl5 |
112.639 |
| O6 |
C2 |
H7 |
125.235 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.015 |
|
|
|
| 2 |
C |
0.105 |
|
|
|
| 3 |
H |
0.133 |
|
|
|
| 4 |
Cl |
-0.109 |
|
|
|
| 5 |
Cl |
-0.095 |
|
|
|
| 6 |
O |
-0.107 |
|
|
|
| 7 |
H |
0.088 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.036 |
-1.520 |
1.871 |
2.410 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.715 |
-0.017 |
-0.452 |
| y |
-0.017 |
-39.058 |
-0.866 |
| z |
-0.452 |
-0.866 |
-37.170 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.601 |
-0.017 |
-0.452 |
| y |
-0.017 |
1.384 |
-0.866 |
| z |
-0.452 |
-0.866 |
4.217 |
|
| Polar |
|---|
| 3z2-r2 | 8.433 |
|---|
| x2-y2 | -4.657 |
|---|
| xy | -0.017 |
|---|
| xz | -0.452 |
|---|
| yz | -0.866 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.970 |
-0.654 |
0.116 |
| y |
-0.654 |
3.772 |
-0.336 |
| z |
0.116 |
-0.336 |
1.879 |
<r2> (average value of r
2) Å
2
| <r2> |
199.341 |
| (<r2>)1/2 |
14.119 |
