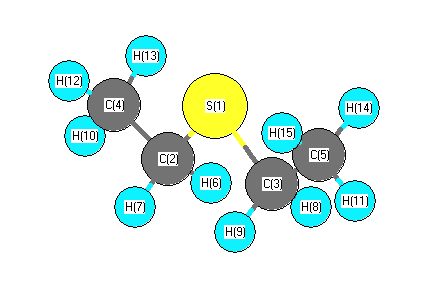
Vibrational Frequencies calculated at BLYP/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3015 |
3008 |
59.69 |
|
|
|
| 2 |
A1 |
2961 |
2953 |
0.30 |
|
|
|
| 3 |
A1 |
2954 |
2946 |
72.51 |
|
|
|
| 4 |
A1 |
1488 |
1484 |
4.81 |
|
|
|
| 5 |
A1 |
1470 |
1466 |
0.28 |
|
|
|
| 6 |
A1 |
1391 |
1388 |
3.00 |
|
|
|
| 7 |
A1 |
1292 |
1289 |
15.86 |
|
|
|
| 8 |
A1 |
1071 |
1068 |
0.13 |
|
|
|
| 9 |
A1 |
962 |
960 |
3.60 |
|
|
|
| 10 |
A1 |
646 |
644 |
0.16 |
|
|
|
| 11 |
A1 |
313 |
312 |
0.99 |
|
|
|
| 12 |
A1 |
132 |
132 |
0.28 |
|
|
|
| 13 |
A2 |
3030 |
3022 |
0.00 |
|
|
|
| 14 |
A2 |
2992 |
2985 |
0.00 |
|
|
|
| 15 |
A2 |
1477 |
1473 |
0.00 |
|
|
|
| 16 |
A2 |
1242 |
1238 |
0.00 |
|
|
|
| 17 |
A2 |
1014 |
1011 |
0.00 |
|
|
|
| 18 |
A2 |
773 |
771 |
0.00 |
|
|
|
| 19 |
A2 |
242 |
242 |
0.00 |
|
|
|
| 20 |
A2 |
76 |
75 |
0.00 |
|
|
|
| 21 |
B1 |
3029 |
3021 |
87.02 |
|
|
|
| 22 |
B1 |
2988 |
2981 |
24.16 |
|
|
|
| 23 |
B1 |
1476 |
1473 |
18.90 |
|
|
|
| 24 |
B1 |
1248 |
1245 |
0.29 |
|
|
|
| 25 |
B1 |
1037 |
1034 |
1.98 |
|
|
|
| 26 |
B1 |
793 |
791 |
13.26 |
|
|
|
| 27 |
B1 |
239 |
239 |
0.18 |
|
|
|
| 28 |
B1 |
51 |
51 |
0.82 |
|
|
|
| 29 |
B2 |
3015 |
3008 |
11.23 |
|
|
|
| 30 |
B2 |
2960 |
2953 |
66.88 |
|
|
|
| 31 |
B2 |
2954 |
2946 |
1.00 |
|
|
|
| 32 |
B2 |
1488 |
1484 |
0.86 |
|
|
|
| 33 |
B2 |
1462 |
1458 |
7.57 |
|
|
|
| 34 |
B2 |
1390 |
1387 |
3.13 |
|
|
|
| 35 |
B2 |
1252 |
1249 |
59.41 |
|
|
|
| 36 |
B2 |
1023 |
1020 |
4.96 |
|
|
|
| 37 |
B2 |
958 |
956 |
5.69 |
|
|
|
| 38 |
B2 |
634 |
632 |
3.02 |
|
|
|
| 39 |
B2 |
326 |
325 |
0.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28430.9 cm
-1
Scaled (by 0.9975) Zero Point Vibrational Energy (zpe) 28359.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.116 |
|
|
|
| 2 |
C |
-0.519 |
|
|
|
| 3 |
C |
-0.519 |
|
|
|
| 4 |
C |
-0.598 |
|
|
|
| 5 |
C |
-0.598 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
| 7 |
H |
0.217 |
|
|
|
| 8 |
H |
0.217 |
|
|
|
| 9 |
H |
0.217 |
|
|
|
| 10 |
H |
0.203 |
|
|
|
| 11 |
H |
0.203 |
|
|
|
| 12 |
H |
0.212 |
|
|
|
| 13 |
H |
0.212 |
|
|
|
| 14 |
H |
0.212 |
|
|
|
| 15 |
H |
0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.703 |
1.703 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
232.855 |
| (<r2>)1/2 |
15.260 |