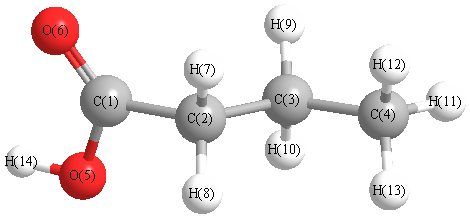Vibrational Frequencies calculated at BLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3431 |
3405 |
7.74 |
|
|
|
| 2 |
A |
3056 |
3033 |
46.17 |
|
|
|
| 3 |
A |
3040 |
3017 |
39.68 |
|
|
|
| 4 |
A |
3035 |
3012 |
30.99 |
|
|
|
| 5 |
A |
3010 |
2987 |
0.07 |
|
|
|
| 6 |
A |
2984 |
2961 |
27.62 |
|
|
|
| 7 |
A |
2968 |
2945 |
9.81 |
|
|
|
| 8 |
A |
2964 |
2941 |
28.84 |
|
|
|
| 9 |
A |
1674 |
1662 |
214.74 |
|
|
|
| 10 |
A |
1518 |
1507 |
5.90 |
|
|
|
| 11 |
A |
1508 |
1496 |
7.03 |
|
|
|
| 12 |
A |
1505 |
1494 |
1.66 |
|
|
|
| 13 |
A |
1484 |
1473 |
3.08 |
|
|
|
| 14 |
A |
1424 |
1413 |
1.41 |
|
|
|
| 15 |
A |
1359 |
1349 |
5.58 |
|
|
|
| 16 |
A |
1328 |
1318 |
26.68 |
|
|
|
| 17 |
A |
1309 |
1299 |
4.23 |
|
|
|
| 18 |
A |
1292 |
1282 |
10.56 |
|
|
|
| 19 |
A |
1248 |
1239 |
0.62 |
|
|
|
| 20 |
A |
1158 |
1149 |
64.92 |
|
|
|
| 21 |
A |
1099 |
1091 |
9.49 |
|
|
|
| 22 |
A |
1034 |
1026 |
143.57 |
|
|
|
| 23 |
A |
1009 |
1001 |
30.85 |
|
|
|
| 24 |
A |
893 |
887 |
0.55 |
|
|
|
| 25 |
A |
888 |
882 |
9.83 |
|
|
|
| 26 |
A |
823 |
817 |
25.90 |
|
|
|
| 27 |
A |
760 |
754 |
3.07 |
|
|
|
| 28 |
A |
686 |
681 |
68.29 |
|
|
|
| 29 |
A |
607 |
602 |
81.44 |
|
|
|
| 30 |
A |
548 |
544 |
41.14 |
|
|
|
| 31 |
A |
415 |
411 |
3.52 |
|
|
|
| 32 |
A |
326 |
324 |
1.35 |
|
|
|
| 33 |
A |
241 |
239 |
0.06 |
|
|
|
| 34 |
A |
188 |
186 |
0.09 |
|
|
|
| 35 |
A |
98 |
98 |
0.17 |
|
|
|
| 36 |
A |
45 |
45 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25476.4 cm
-1
Scaled (by 0.9924) Zero Point Vibrational Energy (zpe) 25282.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.403 |
|
|
|
| 2 |
C |
-0.234 |
|
|
|
| 3 |
C |
-0.207 |
|
|
|
| 4 |
C |
-0.367 |
|
|
|
| 5 |
O |
-0.504 |
|
|
|
| 6 |
O |
-0.375 |
|
|
|
| 7 |
H |
0.143 |
|
|
|
| 8 |
H |
0.146 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.124 |
|
|
|
| 13 |
H |
0.123 |
|
|
|
| 14 |
H |
0.359 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.212 |
1.215 |
0.255 |
1.735 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.847 |
5.727 |
0.219 |
| y |
5.727 |
-39.559 |
-1.712 |
| z |
0.219 |
-1.712 |
-36.247 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.056 |
5.727 |
0.219 |
| y |
5.727 |
-3.512 |
-1.712 |
| z |
0.219 |
-1.712 |
1.456 |
|
| Polar |
|---|
| 3z2-r2 | 2.912 |
|---|
| x2-y2 | 3.712 |
|---|
| xy | 5.727 |
|---|
| xz | 0.219 |
|---|
| yz | -1.712 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.880 |
-0.042 |
-0.411 |
| y |
-0.042 |
7.311 |
0.129 |
| z |
-0.411 |
0.129 |
5.696 |
<r2> (average value of r
2) Å
2
| <r2> |
215.226 |
| (<r2>)1/2 |
14.671 |