Jump to
S1C2
S1C3
Vibrational Frequencies calculated at BLYP/6-31G
Geometric Data calculated at BLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at BLYP/6-31G
Geometric Data calculated at BLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at BLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -210.231525 |
| Energy at 298.15K | -210.240541 |
| HF Energy | -210.231525 |
| Nuclear repulsion energy | 132.546986 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3533 |
3507 |
0.72 |
|
|
|
| 2 |
A |
3417 |
3391 |
0.53 |
|
|
|
| 3 |
A |
3269 |
3244 |
41.47 |
|
|
|
| 4 |
A |
3033 |
3010 |
54.30 |
|
|
|
| 5 |
A |
3020 |
2997 |
28.74 |
|
|
|
| 6 |
A |
2904 |
2882 |
109.20 |
|
|
|
| 7 |
A |
2878 |
2857 |
79.64 |
|
|
|
| 8 |
A |
1660 |
1648 |
25.39 |
|
|
|
| 9 |
A |
1515 |
1504 |
0.16 |
|
|
|
| 10 |
A |
1497 |
1485 |
3.98 |
|
|
|
| 11 |
A |
1412 |
1401 |
85.85 |
|
|
|
| 12 |
A |
1386 |
1375 |
5.40 |
|
|
|
| 13 |
A |
1347 |
1337 |
7.01 |
|
|
|
| 14 |
A |
1298 |
1288 |
4.97 |
|
|
|
| 15 |
A |
1221 |
1211 |
22.56 |
|
|
|
| 16 |
A |
1153 |
1145 |
14.77 |
|
|
|
| 17 |
A |
1053 |
1045 |
12.64 |
|
|
|
| 18 |
A |
991 |
984 |
50.90 |
|
|
|
| 19 |
A |
974 |
966 |
3.09 |
|
|
|
| 20 |
A |
861 |
855 |
15.11 |
|
|
|
| 21 |
A |
847 |
840 |
30.22 |
|
|
|
| 22 |
A |
694 |
688 |
235.00 |
|
|
|
| 23 |
A |
634 |
630 |
109.46 |
|
|
|
| 24 |
A |
534 |
530 |
16.73 |
|
|
|
| 25 |
A |
315 |
313 |
0.50 |
|
|
|
| 26 |
A |
266 |
264 |
11.37 |
|
|
|
| 27 |
A |
176 |
174 |
8.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20943.2 cm
-1
Scaled (by 0.9924) Zero Point Vibrational Energy (zpe) 20784.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.360 |
-0.552 |
0.128 |
| C2 |
-0.641 |
0.693 |
-0.259 |
| C3 |
0.817 |
0.588 |
0.252 |
| O4 |
1.388 |
-0.692 |
-0.163 |
| H5 |
-1.672 |
-0.575 |
1.103 |
| H6 |
-2.106 |
-0.834 |
-0.509 |
| H7 |
-1.118 |
1.625 |
0.112 |
| H8 |
-0.630 |
0.735 |
-1.361 |
| H9 |
1.445 |
1.386 |
-0.175 |
| H10 |
0.832 |
0.695 |
1.360 |
| H11 |
0.611 |
-1.323 |
-0.083 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4886 | 2.4602 | 2.7663 | 1.0241 | 1.0214 | 2.1905 | 2.0991 | 3.4222 | 2.8062 | 2.1270 |
C2 | 1.4886 | | 1.5487 | 2.4581 | 2.1274 | 2.1311 | 1.1109 | 1.1025 | 2.1995 | 2.1891 | 2.3800 | C3 | 2.4602 | 1.5487 | | 1.4616 | 2.8755 | 3.3389 | 2.1994 | 2.1723 | 1.1013 | 1.1127 | 1.9517 | O4 | 2.7663 | 2.4581 | 1.4616 | | 3.3129 | 3.5140 | 3.4235 | 2.7466 | 2.0782 | 2.1327 | 1.0042 | H5 | 1.0241 | 2.1274 | 2.8755 | 3.3129 | | 1.6901 | 2.4759 | 2.9788 | 3.8973 | 2.8188 | 2.6792 | H6 | 1.0214 | 2.1311 | 3.3389 | 3.5140 | 1.6901 | | 2.7225 | 2.3168 | 4.2012 | 3.8033 | 2.7937 | H7 | 2.1905 | 1.1109 | 2.1994 | 3.4235 | 2.4759 | 2.7225 | | 1.7886 | 2.5895 | 2.4951 | 3.4236 | H8 | 2.0991 | 1.1025 | 2.1723 | 2.7466 | 2.9788 | 2.3168 | 1.7886 | | 2.4770 | 3.0892 | 2.7225 | H9 | 3.4222 | 2.1995 | 1.1013 | 2.0782 | 3.8973 | 4.2012 | 2.5895 | 2.4770 | | 1.7913 | 2.8360 | H10 | 2.8062 | 2.1891 | 1.1127 | 2.1327 | 2.8188 | 3.8033 | 2.4951 | 3.0892 | 1.7913 | | 2.4905 | H11 | 2.1270 | 2.3800 | 1.9517 | 1.0042 | 2.6792 | 2.7937 | 3.4236 | 2.7225 | 2.8360 | 2.4905 | |
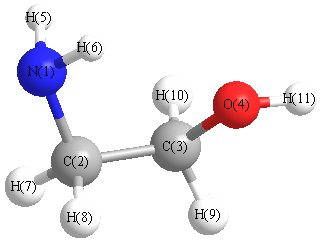 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.178 |
|
N1 |
C2 |
H7 |
114.058 |
| N1 |
C2 |
H8 |
107.273 |
|
C2 |
N1 |
H5 |
114.435 |
| C2 |
N1 |
H6 |
114.945 |
|
C2 |
C3 |
O4 |
109.450 |
| C2 |
C3 |
H9 |
111.075 |
|
C2 |
C3 |
H10 |
109.597 |
| C3 |
C2 |
H7 |
110.498 |
|
C3 |
C2 |
H8 |
108.874 |
| C3 |
O4 |
H11 |
103.078 |
|
O4 |
C3 |
H9 |
107.531 |
| O4 |
C3 |
H10 |
111.159 |
|
H5 |
N1 |
H6 |
111.432 |
| H7 |
C2 |
H8 |
107.812 |
|
H9 |
C3 |
H10 |
108.011 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.664 |
|
|
|
| 2 |
C |
-0.130 |
|
|
|
| 3 |
C |
0.016 |
|
|
|
| 4 |
O |
-0.577 |
|
|
|
| 5 |
H |
0.272 |
|
|
|
| 6 |
H |
0.277 |
|
|
|
| 7 |
H |
0.102 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.094 |
|
|
|
| 11 |
H |
0.347 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.524 |
1.067 |
0.545 |
3.722 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.705 |
3.211 |
0.509 |
| y |
3.211 |
-25.208 |
-0.351 |
| z |
0.509 |
-0.351 |
-24.392 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.905 |
3.211 |
0.509 |
| y |
3.211 |
0.340 |
-0.351 |
| z |
0.509 |
-0.351 |
1.565 |
|
| Polar |
|---|
| 3z2-r2 | 3.130 |
|---|
| x2-y2 | -1.497 |
|---|
| xy | 3.211 |
|---|
| xz | 0.509 |
|---|
| yz | -0.351 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.263 |
0.301 |
-0.053 |
| y |
0.301 |
5.175 |
0.129 |
| z |
-0.053 |
0.129 |
4.526 |
<r2> (average value of r
2) Å
2
| <r2> |
88.351 |
| (<r2>)1/2 |
9.400 |
