Jump to
S1C2
S1C3
Energy calculated at CID/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -254.534935 |
| Energy at 298.15K | -254.534313 |
| HF Energy | -254.215429 |
| Nuclear repulsion energy | 46.517868 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CID/cc-pCVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.702 |
| N2 |
0.000 |
0.000 |
-1.859 |
| Na3 |
0.000 |
0.000 |
1.566 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1566 | 2.2675 |
N2 | 1.1566 | | 3.4242 | Na3 | 2.2675 | 3.4242 | |
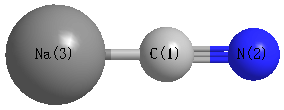 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
0.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CID/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -254.534935 |
| Energy at 298.15K | -254.534313 |
| HF Energy | -254.215429 |
| Nuclear repulsion energy | 46.517868 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/cc-pCVTZ
Geometric Data calculated at CID/cc-pCVTZ
Point Group is C∞v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CID/cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -254.537223 |
| Energy at 298.15K | -254.536347 |
| HF Energy | -254.221565 |
| Nuclear repulsion energy | 48.795897 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CID/cc-pCVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.850 |
| N2 |
0.000 |
0.000 |
-0.684 |
| Na3 |
0.000 |
0.000 |
1.445 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1663 | 3.2951 |
N2 | 1.1663 | | 2.1288 | Na3 | 3.2951 | 2.1288 | |
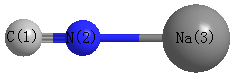 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
180.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
