return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H8 (Norbornadiene)

using model chemistry: CID/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CID/6-311G*
 hartrees
Energy at 0K-270.481931
Energy at 298.15K-270.491580
Nuclear repulsion energy289.220649
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3348 3111 9.53      
2 A1 3239 3009 12.21      
3 A1 3180 2954 44.02      
4 A1 1749 1625 0.06      
5 A1 1596 1483 1.26      
6 A1 1337 1242 4.21      
7 A1 1189 1104 0.31      
8 A1 999 928 4.75      
9 A1 951 884 12.47      
10 A1 828 769 0.64      
11 A1 796 740 65.54      
12 A1 447 415 7.33      
13 A2 3318 3083 0.00      
14 A2 1390 1292 0.00      
15 A2 1337 1243 0.00      
16 A2 1199 1114 0.00      
17 A2 984 915 0.00      
18 A2 968 899 0.00      
19 A2 753 700 0.00      
20 A2 473 440 0.00      
21 B1 3345 3108 22.98      
22 B1 3241 3011 39.28      
23 B1 1707 1586 5.01      
24 B1 1322 1228 0.93      
25 B1 1142 1061 1.23      
26 B1 1095 1017 0.84      
27 B1 975 906 2.81      
28 B1 722 671 40.27      
29 B1 539 501 7.10      
30 B2 3320 3085 15.12      
31 B2 3235 3006 69.24      
32 B2 1422 1321 29.09      
33 B2 1351 1255 0.00      
34 B2 1259 1170 0.44      
35 B2 1027 954 1.01      
36 B2 988 918 3.28      
37 B2 941 874 1.31      
38 B2 857 796 9.22      
39 B2 581 540 1.50      

Unscaled Zero Point Vibrational Energy (zpe) 29574.3 cm-1
Scaled (by 0.9291) Zero Point Vibrational Energy (zpe) 27477.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-311G*
ABC
0.14479 0.12137 0.10737

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.346
C2 0.000 1.115 0.274
C3 0.000 -1.115 0.274
C4 1.235 0.663 -0.519
C5 1.235 -0.663 -0.519
C6 -1.235 0.663 -0.519
C7 -1.235 -0.663 -0.519
H8 -0.893 0.000 1.965
H9 0.893 0.000 1.965
H10 0.000 2.144 0.613
H11 0.000 -2.144 0.613
H12 1.922 1.325 -1.017
H13 1.922 -1.325 -1.017
H14 -1.922 1.325 -1.017
H15 -1.922 -1.325 -1.017

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.54711.54712.33322.33322.33322.33321.08651.08652.26592.26593.32273.32273.32273.3227
C21.54712.22931.53532.30501.53532.30502.21382.21381.08413.27642.32533.36392.32533.3639
C31.54712.22932.30501.53532.30501.53532.21382.21383.27641.08413.36392.32533.36392.3253
C42.33321.53532.30501.32602.47032.80373.33732.59302.23623.26911.07702.16213.26463.7646
C52.33322.30501.53531.32602.80372.47033.33732.59303.26912.23622.16211.07703.76463.2646
C62.33321.53532.30502.47032.80371.32602.59303.33732.23623.26913.26463.76461.07702.1621
C72.33322.30501.53532.80372.47031.32602.59303.33733.26912.23623.76463.26462.16211.0770
H81.08652.21382.21383.33733.33732.59302.59301.78672.68732.68734.31034.31033.42193.4219
H91.08652.21382.21382.59302.59303.33733.33731.78672.68732.68733.42193.42194.31034.3103
H102.26591.08413.27642.23623.26912.23623.26912.68732.68734.28822.65074.28862.65074.2886
H112.26593.27641.08413.26912.23623.26912.23622.68732.68734.28824.28862.65074.28862.6507
H123.32272.32533.36391.07702.16213.26463.76464.31033.42192.65074.28862.65063.84494.6700
H133.32273.36392.32532.16211.07703.76463.26464.31033.42194.28862.65072.65064.67003.8449
H143.32272.32533.36393.26463.76461.07702.16213.42194.31032.65074.28863.84494.67002.6506
H153.32273.36392.32533.76463.26462.16211.07703.42194.31034.28862.65074.67003.84492.6506

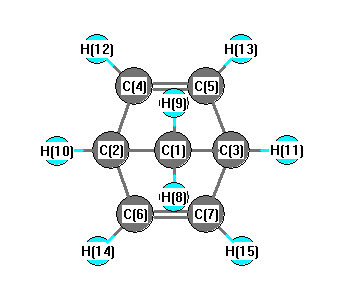
picture of Norbornadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 98.389 C1 C2 C6 98.389
C1 C2 H10 117.822 C1 C3 C5 98.389
C1 C3 C7 98.389 C1 C3 H11 117.822
C2 C1 C3 92.185 C2 C1 H8 113.247
C2 C1 H9 113.247 C2 C4 C5 107.107
C2 C4 H12 124.858 C2 C6 C7 107.107
C2 C6 H14 124.858 C3 C1 H8 113.247
C3 C1 H9 113.247 C3 C5 C4 107.107
C3 C5 H13 124.858 C3 C7 C6 107.107
C3 C7 H15 124.858 C4 C2 C6 107.125
C4 C2 H10 116.174 C4 C5 H13 127.948
C5 C3 C7 107.125 C5 C3 H11 116.174
C5 C4 H12 127.948 C6 C2 H10 116.174
C6 C7 H15 127.948 C7 C3 H11 116.174
C7 C6 H14 127.948 H8 C1 H9 110.620
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability