Jump to
S1C2
Energy calculated at CID/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.515829 |
| Energy at 298.15K | -133.521357 |
| HF Energy | -133.101095 |
| Nuclear repulsion energy | 71.122752 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3595 |
3340 |
1.47 |
|
|
|
| 2 |
A' |
3231 |
3002 |
13.75 |
|
|
|
| 3 |
A' |
3125 |
2904 |
40.70 |
|
|
|
| 4 |
A' |
3114 |
2893 |
30.41 |
|
|
|
| 5 |
A' |
1817 |
1688 |
59.83 |
|
|
|
| 6 |
A' |
1543 |
1434 |
11.75 |
|
|
|
| 7 |
A' |
1516 |
1408 |
32.88 |
|
|
|
| 8 |
A' |
1462 |
1358 |
13.11 |
|
|
|
| 9 |
A' |
1343 |
1248 |
29.47 |
|
|
|
| 10 |
A' |
1116 |
1037 |
25.86 |
|
|
|
| 11 |
A' |
965 |
897 |
5.42 |
|
|
|
| 12 |
A' |
506 |
470 |
18.82 |
|
|
|
| 13 |
A" |
3181 |
2956 |
20.38 |
|
|
|
| 14 |
A" |
1544 |
1434 |
11.02 |
|
|
|
| 15 |
A" |
1187 |
1103 |
7.18 |
|
|
|
| 16 |
A" |
1132 |
1052 |
16.27 |
|
|
|
| 17 |
A" |
714 |
664 |
61.57 |
|
|
|
| 18 |
A" |
189 |
176 |
2.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15640.1 cm
-1
Scaled (by 0.9291) Zero Point Vibrational Energy (zpe) 14531.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.029 |
-0.639 |
0.000 |
| C2 |
0.000 |
0.450 |
0.000 |
| N3 |
1.237 |
0.191 |
0.000 |
| H4 |
-0.537 |
-1.608 |
0.000 |
| H5 |
-1.671 |
-0.560 |
0.878 |
| H6 |
-1.671 |
-0.560 |
-0.878 |
| H7 |
-0.387 |
1.474 |
0.000 |
| H8 |
1.779 |
1.048 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4982 | 2.4137 | 1.0866 | 1.0911 | 1.0911 | 2.2082 | 3.2754 |
C2 | 1.4982 | | 1.2643 | 2.1267 | 2.1412 | 2.1412 | 1.0948 | 1.8765 | N3 | 2.4137 | 1.2643 | | 2.5268 | 3.1300 | 3.1300 | 2.0703 | 1.0133 | H4 | 1.0866 | 2.1267 | 2.5268 | | 1.7768 | 1.7768 | 3.0854 | 3.5234 | H5 | 1.0911 | 2.1412 | 3.1300 | 1.7768 | | 1.7567 | 2.5605 | 3.9063 | H6 | 1.0911 | 2.1412 | 3.1300 | 1.7768 | 1.7567 | | 2.5605 | 3.9063 | H7 | 2.2082 | 1.0948 | 2.0703 | 3.0854 | 2.5605 | 2.5605 | | 2.2077 | H8 | 3.2754 | 1.8765 | 1.0133 | 3.5234 | 3.9063 | 3.9063 | 2.2077 | |
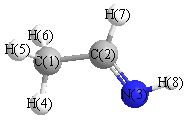 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.558 |
|
C1 |
C2 |
H7 |
115.900 |
| C2 |
C1 |
H4 |
109.699 |
|
C2 |
C1 |
H5 |
110.586 |
| C2 |
C1 |
H6 |
110.586 |
|
C2 |
N3 |
H8 |
110.472 |
| N3 |
C2 |
H7 |
122.542 |
|
H4 |
C1 |
H5 |
109.350 |
| H4 |
C1 |
H6 |
109.350 |
|
H5 |
C1 |
H6 |
107.226 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.514130 |
| Energy at 298.15K | -133.519611 |
| HF Energy | -133.099464 |
| Nuclear repulsion energy | 71.056260 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3555 |
3303 |
5.53 |
|
|
|
| 2 |
A' |
3206 |
2979 |
39.12 |
|
|
|
| 3 |
A' |
3180 |
2954 |
18.23 |
|
|
|
| 4 |
A' |
3115 |
2894 |
10.43 |
|
|
|
| 5 |
A' |
1816 |
1688 |
62.73 |
|
|
|
| 6 |
A' |
1549 |
1439 |
27.88 |
|
|
|
| 7 |
A' |
1494 |
1388 |
24.27 |
|
|
|
| 8 |
A' |
1468 |
1364 |
5.62 |
|
|
|
| 9 |
A' |
1348 |
1252 |
56.65 |
|
|
|
| 10 |
A' |
1113 |
1034 |
21.10 |
|
|
|
| 11 |
A' |
949 |
881 |
1.97 |
|
|
|
| 12 |
A' |
510 |
474 |
8.97 |
|
|
|
| 13 |
A" |
3186 |
2960 |
15.55 |
|
|
|
| 14 |
A" |
1539 |
1430 |
11.48 |
|
|
|
| 15 |
A" |
1202 |
1117 |
52.77 |
|
|
|
| 16 |
A" |
1128 |
1048 |
11.17 |
|
|
|
| 17 |
A" |
714 |
663 |
10.74 |
|
|
|
| 18 |
A" |
160 |
149 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15615.0 cm
-1
Scaled (by 0.9291) Zero Point Vibrational Energy (zpe) 14507.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.016 |
-0.627 |
0.000 |
| C2 |
0.000 |
0.483 |
0.000 |
| N3 |
1.260 |
0.382 |
0.000 |
| H4 |
-0.537 |
-1.605 |
0.000 |
| H5 |
-1.659 |
-0.552 |
0.878 |
| H6 |
-1.659 |
-0.552 |
-0.878 |
| H7 |
-0.405 |
1.495 |
0.000 |
| H8 |
1.536 |
-0.596 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5042 | 2.4889 | 1.0894 | 1.0908 | 1.0908 | 2.2080 | 2.5519 |
C2 | 1.5042 | | 1.2638 | 2.1562 | 2.1433 | 2.1433 | 1.0903 | 1.8772 | N3 | 2.4889 | 1.2638 | | 2.6790 | 3.1878 | 3.1878 | 2.0028 | 1.0161 | H4 | 1.0894 | 2.1562 | 2.6790 | | 1.7717 | 1.7717 | 3.1034 | 2.3057 | H5 | 1.0908 | 2.1433 | 3.1878 | 1.7717 | | 1.7553 | 2.5562 | 3.3138 | H6 | 1.0908 | 2.1433 | 3.1878 | 1.7717 | 1.7553 | | 2.5562 | 3.3138 | H7 | 2.2080 | 1.0903 | 2.0028 | 3.1034 | 2.5562 | 2.5562 | | 2.8533 | H8 | 2.5519 | 1.8772 | 1.0161 | 2.3057 | 3.3138 | 3.3138 | 2.8533 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.877 |
|
C1 |
C2 |
H7 |
115.728 |
| C2 |
C1 |
H4 |
111.471 |
|
C2 |
C1 |
H5 |
110.349 |
| C2 |
C1 |
H6 |
110.349 |
|
C2 |
N3 |
H8 |
110.371 |
| N3 |
C2 |
H7 |
116.395 |
|
H4 |
C1 |
H5 |
108.703 |
| H4 |
C1 |
H6 |
108.703 |
|
H5 |
C1 |
H6 |
107.142 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
