return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7 (n-Propyl radical)

using model chemistry: CID/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A"
Energy calculated at CID/6-311G*
 hartrees
Energy at 0K-118.039277
Energy at 298.15K 
HF Energy-117.652782
Nuclear repulsion energy75.902181
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3324 3089 18.66      
2 A 3220 2991 16.07      
3 A 3191 2965 36.32      
4 A 3187 2961 45.98      
5 A 3130 2908 20.40      
6 A 3113 2892 27.30      
7 A 3058 2841 37.32      
8 A 1570 1458 5.32      
9 A 1564 1453 8.10      
10 A 1550 1440 1.39      
11 A 1528 1419 2.00      
12 A 1482 1376 4.23      
13 A 1425 1324 1.21      
14 A 1334 1240 0.21      
15 A 1226 1139 0.57      
16 A 1128 1048 0.17      
17 A 1106 1028 1.26      
18 A 955 887 1.43      
19 A 921 855 0.21      
20 A 785 730 1.01      
21 A 469 436 49.09      
22 A 379 352 6.55      
23 A 261 243 0.16      
24 A 135 126 0.56      

Unscaled Zero Point Vibrational Energy (zpe) 20019.8 cm-1
Scaled (by 0.9291) Zero Point Vibrational Energy (zpe) 18600.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-311G*
ABC
1.09480 0.29968 0.26011

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.301 -0.295 -0.039
C2 0.075 0.557 0.057
C3 -1.222 -0.247 -0.040
H4 -2.095 0.402 0.033
H5 -1.277 -0.781 -0.989
H6 -1.285 -0.983 0.762
H7 0.080 1.113 1.005
H8 0.097 1.318 -0.727
H9 2.254 0.132 -0.317
H10 1.303 -1.298 0.365

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10
C11.49602.52273.46672.79012.79312.13532.12671.08051.0815
C21.49601.52862.17502.17102.17251.09801.09272.25132.2460
C32.52271.52861.09011.09011.09092.15242.15883.50672.7642
H43.46672.17501.09011.76401.76242.48532.49434.37063.8133
H52.79012.17101.09011.76401.76233.06602.52303.70852.9592
H62.79312.17251.09091.76241.76232.51303.07003.86402.6367
H72.13531.09802.15242.48533.06602.51301.74352.72632.7774
H82.12671.09272.15882.49432.52303.07001.74352.49503.0805
H91.08052.25133.50674.37063.70853.86402.72632.49501.8477
H101.08152.24602.76423.81332.95922.63672.77743.08051.8477

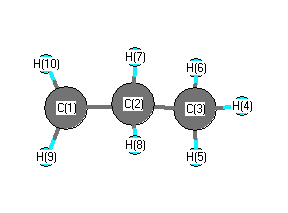
picture of n-Propyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 113.032 C1 C2 H7 109.850
C1 C2 H8 109.485 C2 C1 H9 120.953
C2 C1 H10 120.385 C2 C3 H4 111.210
C2 C3 H5 110.894 C2 C3 H6 110.963
C3 C2 H7 108.961 C3 C2 H8 109.764
H4 C3 H5 108.013 H4 C3 H6 107.811
H5 C3 H6 107.806 H7 C2 H8 105.471
H9 C1 H10 117.439
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability