return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(SH)CH2CH3 (2-Butanethiol)

using model chemistry: CISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CISD/6-31G*
 hartrees
Energy at 0K-555.397664
Energy at 298.15K-555.408672
HF Energy-554.804876
Nuclear repulsion energy237.664226
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3254 3012 27.20      
2 A 3252 3010 15.80      
3 A 3227 2988 46.25      
4 A 3223 2984 28.26      
5 A 3184 2948 22.50      
6 A 3170 2935 4.05      
7 A 3158 2924 15.54      
8 A 3153 2919 38.02      
9 A 3139 2907 18.61      
10 A 2823 2613 17.69      
11 A 1603 1484 5.70      
12 A 1592 1474 6.58      
13 A 1591 1473 4.47      
14 A 1589 1471 6.64      
15 A 1581 1464 1.37      
16 A 1515 1402 7.54      
17 A 1511 1399 2.71      
18 A 1475 1366 0.97      
19 A 1419 1314 12.63      
20 A 1391 1288 0.30      
21 A 1357 1256 3.08      
22 A 1259 1165 1.21      
23 A 1195 1106 13.12      
24 A 1191 1102 1.18      
25 A 1099 1017 0.15      
26 A 1072 993 6.68      
27 A 1028 952 2.80      
28 A 936 867 8.39      
29 A 910 842 3.80      
30 A 836 774 6.47      
31 A 662 613 3.96      
32 A 481 445 0.50      
33 A 399 370 1.25      
34 A 353 327 1.12      
35 A 269 249 0.19      
36 A 244 226 0.35      
37 A 232 215 1.31      
38 A 212 196 20.16      
39 A 114 105 0.16      

Unscaled Zero Point Vibrational Energy (zpe) 30348.9 cm-1
Scaled (by 0.9258) Zero Point Vibrational Energy (zpe) 28097.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CISD/6-31G*
ABC
0.15097 0.10136 0.06632

See section I.F.4 to change rotational constant units
Geometric Data calculated at CISD/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.986 -1.197 -0.006
H2 1.209 -0.991 1.297
C3 1.193 1.568 -0.016
H4 1.388 1.609 1.056
H5 0.765 2.525 -0.315
C6 0.231 0.433 -0.346
H7 0.071 0.411 -1.426
C8 -1.121 0.621 0.342
H9 -1.444 1.648 0.154
H10 -0.983 0.538 1.423
C11 -2.211 -0.337 -0.120
H12 -2.382 -0.241 -1.193
H13 -1.944 -1.370 0.086
H14 -3.151 -0.120 0.386
H15 2.144 1.439 -0.528

Atom - Atom Distances (Å)
  S1 H2 C3 H4 H5 C6 H7 C8 H9 H10 C11 H12 H13 H14 H15
S11.33742.77273.02783.74181.82882.33212.80563.74592.98893.31313.69682.93714.29342.9265
H21.33742.87592.61793.89322.38413.26642.99033.91322.67613.75924.43313.39894.53873.1798
C32.77272.87591.09061.09061.52412.14092.52602.64432.80483.90204.17564.29964.67811.0874
H43.02782.61791.09061.76242.16593.05452.79022.97322.62814.25754.76414.57474.90411.7634
H53.74183.89321.09061.76242.15942.48702.75912.42243.16564.13304.28054.76174.77731.7682
C61.82882.38411.52412.16592.15941.09121.52892.12922.14822.57012.82762.85833.50432.1693
H72.33213.26642.14093.05452.48701.09122.14282.51473.04002.73342.54863.08553.73462.4816
C82.80562.99032.52602.79022.75911.52892.14281.09291.09261.52282.16542.17032.16163.4768
H93.74593.91322.64432.97322.42242.12922.51471.09291.74782.14552.50273.06072.46893.6584
H102.98892.67612.80482.62813.16562.14823.04001.09261.74782.15723.06692.52072.49193.7944
C113.31313.75923.90204.25754.13302.57012.73341.52282.14552.15721.09061.08741.08944.7207
H123.69684.43314.17564.76414.28052.82762.54862.16542.50273.06691.09061.76141.76064.8729
H132.93713.39894.29964.57474.76172.85833.08552.17033.06072.52071.08741.76141.76344.9985
H144.29344.53874.67814.90414.77733.50433.73462.16162.46892.49191.08941.76061.76345.5951
H152.92653.17981.08741.76341.76822.16932.48163.47683.65843.79444.72074.87294.99855.5951

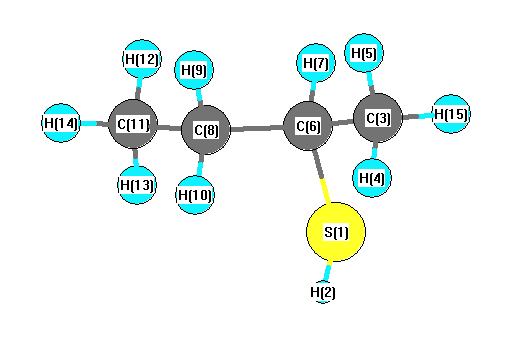
picture of 2-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C6 C3 111.251 S1 C6 H7 103.078
S1 C6 C8 113.044 H2 S1 C6 96.465
C3 C6 H7 108.765 C3 C6 C8 111.664
H4 C3 H5 107.799 H4 C3 C6 110.772
H4 C3 H15 108.120 H5 C3 C6 110.255
H5 C3 H15 108.553 C6 C3 H15 111.234
C6 C8 H9 107.436 C6 C8 H10 108.921
C6 C8 C11 114.745 H7 C6 C8 108.582
C8 C11 H12 110.827 C8 C11 H13 111.411
C8 C11 H14 110.593 H9 C8 H10 106.203
H9 C8 C11 109.112 H10 C8 C11 110.052
H12 C11 H13 107.944 H12 C11 H14 107.727
H13 C11 H14 108.205
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability