return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (Z)-)

using model chemistry: CISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CISD/6-31G*
 hartrees
Energy at 0K-195.731727
Energy at 298.15K-195.741912
HF Energy-195.142015
Nuclear repulsion energy171.631593
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3272 3029 62.29      
2 A 3256 3015 6.78      
3 A 3243 3002 27.58      
4 A 3235 2995 33.58      
5 A 3229 2989 31.88      
6 A 3208 2970 4.81      
7 A 3205 2968 25.39      
8 A 3154 2920 17.82      
9 A 3151 2917 36.04      
10 A 3146 2912 32.58      
11 A 1850 1713 1.32      
12 A 1603 1484 7.40      
13 A 1593 1475 4.87      
14 A 1592 1474 2.63      
15 A 1582 1464 3.66      
16 A 1579 1462 3.43      
17 A 1531 1418 4.82      
18 A 1508 1396 1.98      
19 A 1500 1389 1.83      
20 A 1423 1317 6.88      
21 A 1375 1273 0.09      
22 A 1353 1252 0.13      
23 A 1237 1145 0.41      
24 A 1161 1074 4.59      
25 A 1133 1049 1.21      
26 A 1112 1029 0.04      
27 A 1094 1013 2.01      
28 A 1062 984 0.29      
29 A 991 917 7.77      
30 A 910 843 1.21      
31 A 835 773 3.99      
32 A 765 708 30.93      
33 A 591 547 5.64      
34 A 493 457 1.14      
35 A 320 296 0.03      
36 A 272 251 0.03      
37 A 232 214 0.01      
38 A 122 113 0.24      
39 A 52 48 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 31482.3 cm-1
Scaled (by 0.9258) Zero Point Vibrational Energy (zpe) 29146.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CISD/6-31G*
ABC
0.37128 0.08628 0.07895

See section I.F.4 to change rotational constant units
Geometric Data calculated at CISD/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.172 -0.294 -0.416
H2 2.654 0.671 -0.572
H3 2.929 -0.991 -0.057
H4 1.814 -0.648 -1.382
C5 1.017 -0.176 0.579
H6 0.569 -1.155 0.736
H7 1.413 0.140 1.548
C8 -2.058 -0.677 -0.077
H9 -2.495 -0.919 -1.046
H10 -1.440 -1.517 0.230
H11 -2.884 -0.593 0.632
C12 -0.011 0.822 0.128
H13 0.365 1.833 0.009
C14 -1.294 0.612 -0.152
H15 -1.879 1.466 -0.474

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 H11 C12 H13 C14 H15
C11.09021.08981.08941.52912.15362.14964.26064.75103.86755.17152.51122.82273.59204.4174
H21.09021.76201.76052.17303.06502.51374.92565.40974.71045.80582.75922.63103.97034.6030
H31.08981.76201.76542.17422.49582.48104.99705.51454.41015.86713.45913.81464.51815.4159
H41.08941.76051.76542.16892.50893.06044.08614.33093.73385.11132.78733.19143.57194.3510
C51.52912.17302.17422.16891.08841.09313.18363.94092.82083.92301.50172.18702.54843.4917
H62.15363.06502.49582.50891.08841.74592.79053.55262.10273.49922.14783.08162.71643.7850
H72.14962.51372.48103.06041.09311.74593.91814.80893.55244.45362.12362.51633.23094.0848
C84.26064.92564.99704.08613.18362.79053.91811.09111.08651.09122.54513.49001.50032.1871
H94.75105.40975.51454.33093.94093.55264.80891.09111.76071.75333.25274.10742.14182.5288
H103.86754.71044.41013.73382.82082.10273.55241.08651.76071.76062.74243.81162.16753.0960
H115.17155.80585.86715.11133.92303.49924.45361.09121.75331.76063.24134.10212.14292.5437
C122.51122.75923.45912.78731.50172.14782.12362.54513.25272.74243.24131.08551.32962.0658
H132.82272.63103.81463.19142.18703.08162.51633.49004.10743.81164.10211.08552.06652.3253
C143.59203.97034.51813.57192.54842.71643.23091.50032.14182.16752.14291.32962.06651.0845
H154.41744.60305.41594.35103.49173.78504.08482.18712.52883.09602.54372.06582.32531.0845

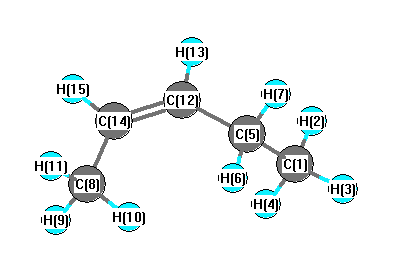
picture of 2-Pentene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.581 C1 C5 H7 108.994
C1 C5 C12 111.901 H2 C1 H3 107.853
H2 C1 H4 107.752 H2 C1 C5 111.014
H3 C1 H4 108.215 H3 C1 C5 111.136
H4 C1 C5 110.736 C5 C12 H13 114.461
C5 C12 C14 128.232 H6 C5 H7 106.319
H6 C5 C12 111.036 H7 C5 C12 108.831
C8 C14 C12 128.046 C8 C14 H15 114.634
H9 C8 H10 107.904 H9 C8 H11 106.907
H9 C8 C14 110.483 H10 C8 H11 107.889
H10 C8 C14 112.861 H11 C8 C14 110.565
C12 C14 H15 117.320 H13 C12 C14 117.302
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability