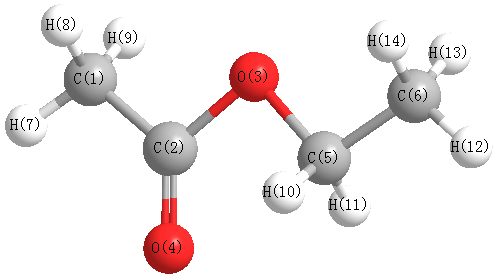
Vibrational Frequencies calculated at B3PW91/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3170 |
3034 |
5.45 |
55.84 |
0.67 |
0.80 |
| 2 |
A' |
3118 |
2985 |
19.92 |
67.39 |
0.69 |
0.81 |
| 3 |
A' |
3057 |
2926 |
3.16 |
138.40 |
0.00 |
0.01 |
| 4 |
A' |
3052 |
2922 |
13.94 |
127.55 |
0.04 |
0.07 |
| 5 |
A' |
3043 |
2913 |
15.64 |
161.22 |
0.00 |
0.01 |
| 6 |
A' |
1813 |
1736 |
278.85 |
7.54 |
0.24 |
0.38 |
| 7 |
A' |
1514 |
1449 |
7.80 |
1.01 |
0.75 |
0.86 |
| 8 |
A' |
1493 |
1429 |
2.97 |
6.83 |
0.75 |
0.86 |
| 9 |
A' |
1465 |
1402 |
15.36 |
5.67 |
0.61 |
0.76 |
| 10 |
A' |
1422 |
1361 |
6.02 |
1.21 |
0.74 |
0.85 |
| 11 |
A' |
1398 |
1339 |
55.12 |
0.17 |
0.70 |
0.82 |
| 12 |
A' |
1382 |
1323 |
18.58 |
1.03 |
0.42 |
0.59 |
| 13 |
A' |
1275 |
1221 |
405.81 |
1.07 |
0.60 |
0.75 |
| 14 |
A' |
1137 |
1089 |
12.12 |
7.29 |
0.11 |
0.19 |
| 15 |
A' |
1083 |
1037 |
104.20 |
1.78 |
0.75 |
0.86 |
| 16 |
A' |
1010 |
967 |
7.42 |
1.32 |
0.15 |
0.26 |
| 17 |
A' |
957 |
916 |
5.63 |
2.13 |
0.13 |
0.23 |
| 18 |
A' |
866 |
829 |
7.58 |
6.42 |
0.21 |
0.34 |
| 19 |
A' |
643 |
616 |
5.61 |
7.38 |
0.21 |
0.35 |
| 20 |
A' |
430 |
411 |
0.94 |
0.27 |
0.66 |
0.80 |
| 21 |
A' |
369 |
353 |
8.94 |
3.16 |
0.22 |
0.36 |
| 22 |
A' |
193 |
184 |
5.71 |
0.40 |
0.56 |
0.72 |
| 23 |
A" |
3125 |
2992 |
32.11 |
21.64 |
0.75 |
0.86 |
| 24 |
A" |
3124 |
2990 |
2.82 |
48.66 |
0.75 |
0.86 |
| 25 |
A" |
3090 |
2958 |
5.12 |
75.95 |
0.75 |
0.86 |
| 26 |
A" |
1481 |
1417 |
7.36 |
5.12 |
0.75 |
0.86 |
| 27 |
A" |
1471 |
1408 |
8.74 |
4.13 |
0.75 |
0.86 |
| 28 |
A" |
1294 |
1239 |
1.11 |
3.69 |
0.75 |
0.86 |
| 29 |
A" |
1174 |
1124 |
3.60 |
0.35 |
0.75 |
0.86 |
| 30 |
A" |
1060 |
1015 |
6.33 |
0.05 |
0.75 |
0.86 |
| 31 |
A" |
807 |
772 |
1.02 |
0.21 |
0.75 |
0.86 |
| 32 |
A" |
612 |
585 |
4.50 |
0.34 |
0.75 |
0.86 |
| 33 |
A" |
262 |
251 |
0.96 |
0.01 |
0.75 |
0.86 |
| 34 |
A" |
154 |
147 |
4.61 |
0.03 |
0.75 |
0.86 |
| 35 |
A" |
65 |
62 |
0.35 |
0.21 |
0.75 |
0.86 |
| 36 |
A" |
41 |
40 |
0.34 |
0.06 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 25823.4 cm
-1
Scaled (by 0.9573) Zero Point Vibrational Energy (zpe) 24720.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.321 |
|
|
|
| 2 |
C |
0.734 |
|
|
|
| 3 |
O |
-0.552 |
|
|
|
| 4 |
O |
-0.727 |
|
|
|
| 5 |
C |
0.176 |
|
|
|
| 6 |
C |
-0.413 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.137 |
|
|
|
| 11 |
H |
0.137 |
|
|
|
| 12 |
H |
0.130 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
| 14 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.309 |
2.068 |
0.000 |
2.091 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.400 |
-1.048 |
0.000 |
| y |
-1.048 |
-42.931 |
0.000 |
| z |
0.000 |
0.000 |
-35.530 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.831 |
-1.048 |
0.000 |
| y |
-1.048 |
-9.966 |
0.000 |
| z |
0.000 |
0.000 |
1.136 |
|
| Polar |
|---|
| 3z2-r2 | 2.271 |
|---|
| x2-y2 | 12.532 |
|---|
| xy | -1.048 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.200 |
0.598 |
0.000 |
| y |
0.598 |
8.757 |
0.000 |
| z |
0.000 |
0.000 |
6.811 |
<r2> (average value of r
2) Å
2
| <r2> |
200.537 |
| (<r2>)1/2 |
14.161 |