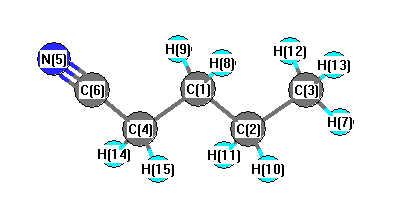
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3130 |
3020 |
26.11 |
|
|
|
| 2 |
A' |
3054 |
2947 |
32.67 |
|
|
|
| 3 |
A' |
3046 |
2939 |
3.81 |
|
|
|
| 4 |
A' |
3040 |
2934 |
20.33 |
|
|
|
| 5 |
A' |
3026 |
2920 |
23.94 |
|
|
|
| 6 |
A' |
2369 |
2286 |
13.70 |
|
|
|
| 7 |
A' |
1481 |
1430 |
7.49 |
|
|
|
| 8 |
A' |
1469 |
1418 |
0.54 |
|
|
|
| 9 |
A' |
1462 |
1410 |
0.95 |
|
|
|
| 10 |
A' |
1439 |
1389 |
3.83 |
|
|
|
| 11 |
A' |
1399 |
1350 |
0.79 |
|
|
|
| 12 |
A' |
1393 |
1344 |
1.83 |
|
|
|
| 13 |
A' |
1342 |
1295 |
2.25 |
|
|
|
| 14 |
A' |
1258 |
1214 |
3.00 |
|
|
|
| 15 |
A' |
1124 |
1085 |
2.89 |
|
|
|
| 16 |
A' |
1084 |
1046 |
0.16 |
|
|
|
| 17 |
A' |
1041 |
1005 |
0.80 |
|
|
|
| 18 |
A' |
952 |
918 |
0.43 |
|
|
|
| 19 |
A' |
911 |
879 |
2.57 |
|
|
|
| 20 |
A' |
544 |
525 |
0.93 |
|
|
|
| 21 |
A' |
384 |
371 |
0.70 |
|
|
|
| 22 |
A' |
280 |
270 |
0.98 |
|
|
|
| 23 |
A' |
125 |
121 |
5.03 |
|
|
|
| 24 |
A" |
3122 |
3013 |
45.16 |
|
|
|
| 25 |
A" |
3108 |
2999 |
14.09 |
|
|
|
| 26 |
A" |
3087 |
2979 |
2.93 |
|
|
|
| 27 |
A" |
3061 |
2954 |
8.32 |
|
|
|
| 28 |
A" |
1468 |
1417 |
7.85 |
|
|
|
| 29 |
A" |
1319 |
1273 |
0.13 |
|
|
|
| 30 |
A" |
1300 |
1255 |
0.33 |
|
|
|
| 31 |
A" |
1225 |
1182 |
0.10 |
|
|
|
| 32 |
A" |
1121 |
1082 |
0.06 |
|
|
|
| 33 |
A" |
924 |
892 |
0.95 |
|
|
|
| 34 |
A" |
789 |
761 |
0.06 |
|
|
|
| 35 |
A" |
739 |
713 |
5.19 |
|
|
|
| 36 |
A" |
393 |
379 |
0.15 |
|
|
|
| 37 |
A" |
248 |
239 |
0.12 |
|
|
|
| 38 |
A" |
117 |
113 |
0.42 |
|
|
|
| 39 |
A" |
88 |
85 |
3.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28729.6 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 27724.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.080 |
|
|
|
| 2 |
C |
-0.109 |
|
|
|
| 3 |
C |
-0.070 |
|
|
|
| 4 |
C |
-0.020 |
|
|
|
| 5 |
N |
-0.140 |
|
|
|
| 6 |
C |
-0.079 |
|
|
|
| 7 |
H |
0.044 |
|
|
|
| 8 |
H |
0.055 |
|
|
|
| 9 |
H |
0.055 |
|
|
|
| 10 |
H |
0.039 |
|
|
|
| 11 |
H |
0.039 |
|
|
|
| 12 |
H |
0.047 |
|
|
|
| 13 |
H |
0.047 |
|
|
|
| 14 |
H |
0.086 |
|
|
|
| 15 |
H |
0.086 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.652 |
1.877 |
0.000 |
4.106 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.998 |
-7.791 |
0.000 |
| y |
-7.791 |
-41.833 |
0.000 |
| z |
0.000 |
0.000 |
-36.559 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.802 |
-7.791 |
0.000 |
| y |
-7.791 |
-0.054 |
0.000 |
| z |
0.000 |
0.000 |
7.856 |
|
| Polar |
|---|
| 3z2-r2 | 15.713 |
|---|
| x2-y2 | -5.165 |
|---|
| xy | -7.791 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
258.604 |
| (<r2>)1/2 |
16.081 |