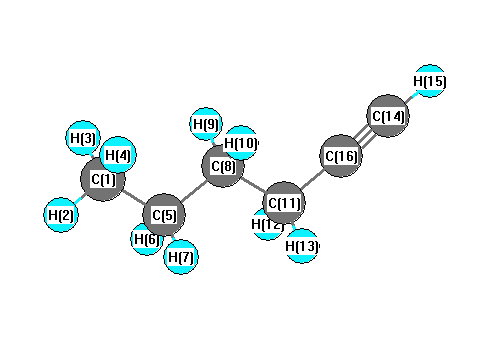
Vibrational Frequencies calculated at B3PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3474 |
3341 |
70.10 |
|
|
|
| 2 |
A' |
3105 |
2986 |
34.65 |
|
|
|
| 3 |
A' |
3040 |
2923 |
50.70 |
|
|
|
| 4 |
A' |
3031 |
2915 |
22.19 |
|
|
|
| 5 |
A' |
3021 |
2905 |
7.62 |
|
|
|
| 6 |
A' |
3017 |
2901 |
16.66 |
|
|
|
| 7 |
A' |
2225 |
2139 |
12.72 |
|
|
|
| 8 |
A' |
1505 |
1447 |
8.41 |
|
|
|
| 9 |
A' |
1492 |
1434 |
1.09 |
|
|
|
| 10 |
A' |
1483 |
1426 |
0.56 |
|
|
|
| 11 |
A' |
1465 |
1409 |
1.98 |
|
|
|
| 12 |
A' |
1409 |
1355 |
3.10 |
|
|
|
| 13 |
A' |
1391 |
1337 |
0.68 |
|
|
|
| 14 |
A' |
1352 |
1300 |
2.12 |
|
|
|
| 15 |
A' |
1271 |
1222 |
3.22 |
|
|
|
| 16 |
A' |
1126 |
1083 |
3.04 |
|
|
|
| 17 |
A' |
1072 |
1031 |
0.16 |
|
|
|
| 18 |
A' |
1033 |
993 |
1.35 |
|
|
|
| 19 |
A' |
954 |
918 |
0.09 |
|
|
|
| 20 |
A' |
908 |
873 |
2.66 |
|
|
|
| 21 |
A' |
673 |
647 |
45.04 |
|
|
|
| 22 |
A' |
519 |
499 |
8.97 |
|
|
|
| 23 |
A' |
381 |
367 |
0.03 |
|
|
|
| 24 |
A' |
272 |
262 |
1.41 |
|
|
|
| 25 |
A' |
119 |
115 |
0.05 |
|
|
|
| 26 |
A" |
3100 |
2981 |
58.87 |
|
|
|
| 27 |
A" |
3083 |
2964 |
22.27 |
|
|
|
| 28 |
A" |
3052 |
2934 |
0.01 |
|
|
|
| 29 |
A" |
3042 |
2925 |
6.53 |
|
|
|
| 30 |
A" |
1494 |
1437 |
7.66 |
|
|
|
| 31 |
A" |
1326 |
1275 |
0.08 |
|
|
|
| 32 |
A" |
1309 |
1258 |
0.44 |
|
|
|
| 33 |
A" |
1237 |
1189 |
0.03 |
|
|
|
| 34 |
A" |
1128 |
1085 |
0.20 |
|
|
|
| 35 |
A" |
931 |
895 |
0.21 |
|
|
|
| 36 |
A" |
788 |
758 |
0.53 |
|
|
|
| 37 |
A" |
732 |
704 |
3.72 |
|
|
|
| 38 |
A" |
665 |
640 |
46.81 |
|
|
|
| 39 |
A" |
356 |
343 |
7.80 |
|
|
|
| 40 |
A" |
246 |
237 |
0.04 |
|
|
|
| 41 |
A" |
115 |
111 |
0.00 |
|
|
|
| 42 |
A" |
90 |
86 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31014.6 cm
-1
Scaled (by 0.9616) Zero Point Vibrational Energy (zpe) 29823.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.281 |
|
|
|
| 2 |
H |
0.094 |
|
|
|
| 3 |
H |
0.088 |
|
|
|
| 4 |
H |
0.088 |
|
|
|
| 5 |
C |
-0.161 |
|
|
|
| 6 |
H |
0.082 |
|
|
|
| 7 |
H |
0.082 |
|
|
|
| 8 |
C |
-0.137 |
|
|
|
| 9 |
H |
0.092 |
|
|
|
| 10 |
H |
0.092 |
|
|
|
| 11 |
C |
-0.150 |
|
|
|
| 12 |
H |
0.100 |
|
|
|
| 13 |
H |
0.100 |
|
|
|
| 14 |
C |
-0.215 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
| 16 |
C |
-0.001 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.770 |
0.587 |
0.000 |
0.968 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.331 |
0.714 |
0.000 |
| y |
0.714 |
-39.983 |
0.000 |
| z |
0.000 |
0.000 |
-38.824 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.073 |
0.714 |
0.000 |
| y |
0.714 |
-2.405 |
0.000 |
| z |
0.000 |
0.000 |
-0.667 |
|
| Polar |
|---|
| 3z2-r2 | -1.335 |
|---|
| x2-y2 | 3.652 |
|---|
| xy | 0.714 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
264.966 |
| (<r2>)1/2 |
16.278 |