return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Pentadiene, (Z)-)

using model chemistry: MP2=FULL/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G(2df,p)
 hartrees
Energy at 0K-194.798099
Energy at 298.15K-194.805588
HF Energy-193.979706
Nuclear repulsion energy156.157965
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3321 3121 6.75      
2 A' 3249 3053 15.18      
3 A' 3233 3039 14.24      
4 A' 3224 3030 3.13      
5 A' 3219 3025 2.61      
6 A' 3205 3013 0.15      
7 A' 3110 2923 14.58      
8 A' 1733 1629 4.15      
9 A' 1663 1563 0.98      
10 A' 1531 1439 7.04      
11 A' 1489 1399 10.54      
12 A' 1441 1354 3.98      
13 A' 1406 1322 0.77      
14 A' 1333 1253 0.80      
15 A' 1305 1227 0.92      
16 A' 1204 1132 1.30      
17 A' 1074 1009 0.02      
18 A' 986 927 9.99      
19 A' 911 856 1.35      
20 A' 616 579 4.79      
21 A' 387 364 3.32      
22 A' 225 211 0.26      
23 A" 3181 2990 9.69      
24 A" 1525 1434 5.62      
25 A" 1075 1011 1.47      
26 A" 1055 992 11.35      
27 A" 1002 942 1.68      
28 A" 938 882 34.16      
29 A" 819 770 5.93      
30 A" 654 615 21.74      
31 A" 364 342 0.10      
32 A" 143 134 0.00      
33 A" 119 112 0.56      

Unscaled Zero Point Vibrational Energy (zpe) 25369.1 cm-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 23844.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G(2df,p)
ABC
0.53515 0.08882 0.07725

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.382 0.159 0.000
C2 1.096 -0.212 0.000
C3 0.000 0.736 0.000
C4 -1.310 0.447 0.000
C5 -1.939 -0.906 0.000
H6 2.663 1.204 0.000
H7 3.183 -0.566 0.000
H8 0.859 -1.269 0.000
H9 0.289 1.782 0.000
H10 -2.003 1.282 0.000
H11 -1.209 -1.711 0.000
H12 -2.578 -1.032 0.875
H13 -2.578 -1.032 -0.875

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.33852.45103.70384.45031.08221.08002.08712.64884.52674.04915.17535.1753
C21.33851.44872.49503.11322.11202.11651.08292.15093.44032.75003.86453.8645
C32.45101.44871.34172.54062.70403.43862.18071.08542.07622.72943.24563.2456
C43.70382.49501.34171.49234.04504.60592.76642.08281.08482.16102.13532.1353
C54.45033.11322.54061.49235.06275.13272.82173.49112.18951.08651.09101.0910
H61.08222.11202.70404.04505.06271.84473.06052.44374.66674.84705.76465.7646
H71.08002.11653.43864.60595.13271.84472.42703.72665.50514.53865.84505.8450
H82.08711.08292.18072.76642.82173.06052.42703.10343.83412.11553.55483.5548
H92.64882.15091.08542.08283.49112.44373.72663.10342.34553.80084.11074.1107
H104.52673.44032.07621.08482.18954.66675.50513.83412.34553.09712.53982.5398
H114.04912.75002.72942.16101.08654.84704.53862.11553.80083.09711.76111.7611
H125.17533.86453.24562.13531.09105.76465.84503.55484.11072.53981.76111.7510
H135.17533.86453.24562.13531.09105.76465.84503.55484.11072.53981.76111.7510

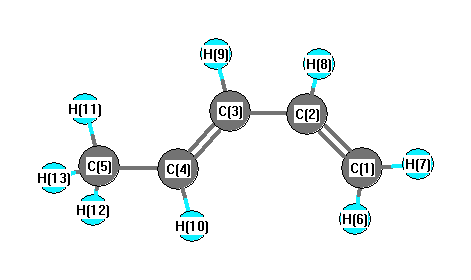
picture of 1,3-Pentadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.081 C1 C2 H8 118.693
C2 C1 H6 121.130 C2 C1 H7 121.753
C2 C3 C4 126.753 C2 C3 H9 115.412
C3 C2 H8 118.227 C3 C4 C5 127.311
C3 C4 H10 117.264 C4 C3 H9 117.835
C4 C5 H11 112.909 C4 C5 H12 110.533
C4 C5 H13 110.533 C5 C4 H10 115.425
H6 C1 H7 117.117 H11 C5 H12 107.947
H11 C5 H13 107.947 H12 C5 H13 106.728
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability