Jump to
S1C2
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -1071.665520 |
| Energy at 298.15K | -1071.668101 |
| HF Energy | -1070.725224 |
| Nuclear repulsion energy | 265.340451 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3217 |
3023 |
1.75 |
|
|
|
| 2 |
A' |
3030 |
2848 |
32.76 |
|
|
|
| 3 |
A' |
1805 |
1696 |
103.13 |
|
|
|
| 4 |
A' |
1420 |
1335 |
2.16 |
|
|
|
| 5 |
A' |
1235 |
1161 |
5.65 |
|
|
|
| 6 |
A' |
1101 |
1034 |
24.82 |
|
|
|
| 7 |
A' |
816 |
767 |
19.77 |
|
|
|
| 8 |
A' |
450 |
423 |
3.75 |
|
|
|
| 9 |
A' |
325 |
306 |
21.53 |
|
|
|
| 10 |
A' |
262 |
246 |
1.96 |
|
|
|
| 11 |
A" |
1265 |
1189 |
23.08 |
|
|
|
| 12 |
A" |
1032 |
970 |
19.65 |
|
|
|
| 13 |
A" |
797 |
749 |
98.28 |
|
|
|
| 14 |
A" |
298 |
280 |
3.16 |
|
|
|
| 15 |
A" |
86 |
81 |
6.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8568.9 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 8053.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.340 |
0.205 |
0.000 |
| C2 |
-0.931 |
1.032 |
0.000 |
| H3 |
1.218 |
0.844 |
0.000 |
| Cl4 |
0.340 |
-0.782 |
1.459 |
| Cl5 |
0.340 |
-0.782 |
-1.459 |
| O6 |
-0.924 |
2.235 |
0.000 |
| H7 |
-1.850 |
0.421 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5169 | 1.0852 | 1.7616 | 1.7616 | 2.3914 | 2.2013 |
C2 | 1.5169 | | 2.1575 | 2.6526 | 2.6526 | 1.2032 | 1.1033 | H3 | 1.0852 | 2.1575 | | 2.3540 | 2.3540 | 2.5540 | 3.0971 | Cl4 | 1.7616 | 2.6526 | 2.3540 | | 2.9186 | 3.5817 | 2.8940 | Cl5 | 1.7616 | 2.6526 | 2.3540 | 2.9186 | | 3.5817 | 2.8940 | O6 | 2.3914 | 1.2032 | 2.5540 | 3.5817 | 3.5817 | | 2.0368 | H7 | 2.2013 | 1.1033 | 3.0971 | 2.8940 | 2.8940 | 2.0368 | |
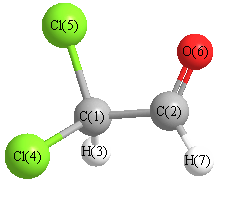 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.665 |
|
C1 |
C2 |
H7 |
113.361 |
| C2 |
C1 |
H3 |
110.930 |
|
C2 |
C1 |
Cl4 |
107.777 |
| C2 |
C1 |
Cl5 |
107.777 |
|
H3 |
C1 |
Cl4 |
109.244 |
| H3 |
C1 |
Cl5 |
109.244 |
|
Cl4 |
C1 |
Cl5 |
111.866 |
| O6 |
C2 |
H7 |
123.974 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -1071.663551 |
| Energy at 298.15K | |
| HF Energy | -1070.722468 |
| Nuclear repulsion energy | 268.285857 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
2981 |
3.89 |
66.62 |
0.23 |
0.37 |
| 2 |
A |
3020 |
2838 |
43.29 |
111.61 |
0.30 |
0.46 |
| 3 |
A |
1811 |
1702 |
82.30 |
8.60 |
0.43 |
0.60 |
| 4 |
A |
1413 |
1329 |
12.12 |
4.71 |
0.57 |
0.73 |
| 5 |
A |
1290 |
1212 |
13.59 |
3.26 |
0.70 |
0.82 |
| 6 |
A |
1245 |
1170 |
11.38 |
6.50 |
0.72 |
0.84 |
| 7 |
A |
1056 |
993 |
14.88 |
1.71 |
0.39 |
0.56 |
| 8 |
A |
961 |
904 |
5.96 |
4.98 |
0.63 |
0.77 |
| 9 |
A |
860 |
809 |
72.76 |
5.04 |
0.73 |
0.84 |
| 10 |
A |
675 |
634 |
25.51 |
9.85 |
0.06 |
0.12 |
| 11 |
A |
643 |
604 |
29.84 |
4.52 |
0.53 |
0.70 |
| 12 |
A |
354 |
332 |
1.36 |
2.12 |
0.20 |
0.34 |
| 13 |
A |
284 |
267 |
3.37 |
3.83 |
0.63 |
0.77 |
| 14 |
A |
226 |
212 |
2.63 |
0.97 |
0.68 |
0.81 |
| 15 |
A |
97 |
91 |
8.54 |
1.44 |
0.73 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 8553.1 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 8039.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.106 |
-0.013 |
0.530 |
| C2 |
-0.691 |
-1.273 |
0.215 |
| H3 |
0.243 |
0.078 |
1.606 |
| Cl4 |
1.714 |
-0.244 |
-0.175 |
| Cl5 |
-0.685 |
1.426 |
-0.057 |
| O6 |
-1.758 |
-1.284 |
-0.334 |
| H7 |
-0.173 |
-2.188 |
0.551 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5241 | 1.0887 | 1.7716 | 1.7440 | 2.4162 | 2.1936 |
C2 | 1.5241 | | 2.1526 | 2.6449 | 2.7135 | 1.2007 | 1.1039 | H3 | 1.0887 | 2.1526 | | 2.3328 | 2.3338 | 3.1025 | 2.5346 | Cl4 | 1.7716 | 2.6449 | 2.3328 | | 2.9260 | 3.6286 | 2.8051 | Cl5 | 1.7440 | 2.7135 | 2.3338 | 2.9260 | | 2.9282 | 3.7013 | O6 | 2.4162 | 1.2007 | 3.1025 | 3.6286 | 2.9282 | | 2.0288 | H7 | 2.1936 | 1.1039 | 2.5346 | 2.8051 | 3.7013 | 2.0288 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.505 |
|
C1 |
C2 |
H7 |
112.185 |
| C2 |
C1 |
H3 |
109.833 |
|
C2 |
C1 |
Cl4 |
106.508 |
| C2 |
C1 |
Cl5 |
112.085 |
|
H3 |
C1 |
Cl4 |
106.855 |
| H3 |
C1 |
Cl5 |
108.737 |
|
Cl4 |
C1 |
Cl5 |
112.671 |
| O6 |
C2 |
H7 |
123.310 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
