return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4O2 (1,3-dioxetane)

using model chemistry: MP2=FULL/6-311G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at MP2=FULL/6-311G**
 hartrees
Energy at 0K-228.540896
Energy at 298.15K-228.546588
Nuclear repulsion energy130.955178
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3073 2915 0.00      
2 Ag 1613 1530 0.00      
3 Ag 1146 1088 0.00      
4 Ag 894 848 0.00      
5 Au 1157 1098 0.00      
6 B1g 1377 1306 0.00      
7 B1g 1048 994 0.00      
8 B1u 3125 2965 139.27      
9 B1u 1198 1137 23.02      
10 B1u 140 133 15.15      
11 B2g 3123 2962 0.00      
12 B2g 1143 1084 0.00      
13 B2u 1482 1406 20.96      
14 B2u 982 932 100.06      
15 B3g 1094 1038 0.00      
16 B3u 3059 2902 201.92      
17 B3u 1576 1495 17.51      
18 B3u 1107 1050 240.09      

Unscaled Zero Point Vibrational Energy (zpe) 14169.1 cm-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 13440.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G**
ABC
0.46356 0.44642 0.24958

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G**

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.975 0.000 0.000
C2 0.975 0.000 0.000
O3 0.000 1.038 0.000
O4 0.000 -1.038 0.000
H5 -1.595 0.000 0.904
H6 1.595 0.000 0.904
H7 -1.595 0.000 -0.904
H8 1.595 0.000 -0.904

Atom - Atom Distances (Å)
  C1 C2 O3 O4 H5 H6 H7 H8
C11.95001.42411.42411.09612.72451.09612.7245
C21.95001.42411.42412.72451.09612.72451.0961
O31.42411.42412.07612.10692.10692.10692.1069
O41.42411.42412.07612.10692.10692.10692.1069
H51.09612.72452.10692.10693.19051.80743.6669
H62.72451.09612.10692.10693.19053.66691.8074
H71.09612.72452.10692.10691.80743.66693.1905
H82.72451.09612.10692.10693.66691.80743.1905

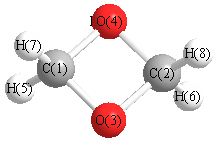
picture of 1,3-dioxetane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O3 C2 86.412 C1 O4 C2 86.412
O3 C1 O4 93.588 O3 C1 H5 112.794
O3 C1 H7 112.794 O3 C2 O4 93.588
O3 C2 H6 112.794 O3 C2 H8 112.794
O4 C1 H5 112.794 O4 C1 H7 112.794
O4 C2 H6 112.794 O4 C2 H8 112.794
H5 C1 H7 111.072 H6 C2 H8 111.072
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability