Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -277.598549 |
| Energy at 298.15K | |
| HF Energy | -276.939570 |
| Nuclear repulsion energy | 125.430917 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3117 |
2963 |
0.00 |
|
|
|
| 2 |
Ag |
1527 |
1451 |
0.00 |
|
|
|
| 3 |
Ag |
1474 |
1401 |
0.00 |
|
|
|
| 4 |
Ag |
1128 |
1072 |
0.00 |
|
|
|
| 5 |
Ag |
1097 |
1043 |
0.00 |
|
|
|
| 6 |
Ag |
465 |
442 |
0.00 |
|
|
|
| 7 |
Au |
3200 |
3042 |
61.47 |
|
|
|
| 8 |
Au |
1245 |
1183 |
4.07 |
|
|
|
| 9 |
Au |
823 |
782 |
0.11 |
|
|
|
| 10 |
Au |
131 |
124 |
12.66 |
|
|
|
| 11 |
Bg |
3177 |
3019 |
0.00 |
|
|
|
| 12 |
Bg |
1309 |
1244 |
0.00 |
|
|
|
| 13 |
Bg |
1191 |
1132 |
0.00 |
|
|
|
| 14 |
Bu |
3120 |
2965 |
61.96 |
|
|
|
| 15 |
Bu |
1536 |
1459 |
1.73 |
|
|
|
| 16 |
Bu |
1367 |
1299 |
17.73 |
|
|
|
| 17 |
Bu |
1113 |
1058 |
183.54 |
|
|
|
| 18 |
Bu |
275 |
261 |
18.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13646.9 cm
-1
Scaled (by 0.9504) Zero Point Vibrational Energy (zpe) 12970.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.424 |
0.629 |
0.000 |
| C2 |
-0.424 |
-0.629 |
0.000 |
| F3 |
-0.424 |
1.727 |
0.000 |
| F4 |
0.424 |
-1.727 |
0.000 |
| H5 |
1.062 |
0.662 |
0.897 |
| H6 |
1.062 |
0.662 |
-0.897 |
| H7 |
-1.062 |
-0.662 |
0.897 |
| H8 |
-1.062 |
-0.662 |
-0.897 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5169 | 1.3872 | 2.3556 | 1.1012 | 1.1012 | 2.1629 | 2.1629 |
C2 | 1.5169 | | 2.3556 | 1.3872 | 2.1629 | 2.1629 | 1.1012 | 1.1012 | F3 | 1.3872 | 2.3556 | | 3.5560 | 2.0366 | 2.0366 | 2.6297 | 2.6297 | F4 | 2.3556 | 1.3872 | 3.5560 | | 2.6297 | 2.6297 | 2.0366 | 2.0366 | H5 | 1.1012 | 2.1629 | 2.0366 | 2.6297 | | 1.7937 | 2.5027 | 3.0791 | H6 | 1.1012 | 2.1629 | 2.0366 | 2.6297 | 1.7937 | | 3.0791 | 2.5027 | H7 | 2.1629 | 1.1012 | 2.6297 | 2.0366 | 2.5027 | 3.0791 | | 1.7937 | H8 | 2.1629 | 1.1012 | 2.6297 | 2.0366 | 3.0791 | 2.5027 | 1.7937 | |
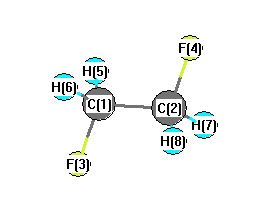 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.325 |
|
C1 |
C2 |
H7 |
110.399 |
| C1 |
C2 |
H8 |
110.399 |
|
C2 |
C1 |
F3 |
108.325 |
| C2 |
C1 |
H5 |
110.399 |
|
C2 |
C1 |
H6 |
110.399 |
| F3 |
C1 |
H5 |
109.318 |
|
F3 |
C1 |
H6 |
109.318 |
| F4 |
C2 |
H7 |
109.318 |
|
F4 |
C2 |
H8 |
109.318 |
| H5 |
C1 |
H6 |
109.061 |
|
H7 |
C2 |
H8 |
109.061 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -277.598998 |
| Energy at 298.15K | |
| HF Energy | -276.939191 |
| Nuclear repulsion energy | 127.574168 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3169 |
3012 |
20.29 |
|
|
|
| 2 |
A |
3108 |
2954 |
38.24 |
|
|
|
| 3 |
A |
1503 |
1428 |
0.57 |
|
|
|
| 4 |
A |
1467 |
1394 |
20.50 |
|
|
|
| 5 |
A |
1320 |
1254 |
1.37 |
|
|
|
| 6 |
A |
1161 |
1103 |
69.25 |
|
|
|
| 7 |
A |
1146 |
1089 |
6.94 |
|
|
|
| 8 |
A |
895 |
850 |
27.46 |
|
|
|
| 9 |
A |
324 |
308 |
0.26 |
|
|
|
| 10 |
A |
152 |
145 |
3.44 |
|
|
|
| 11 |
B |
3181 |
3023 |
46.06 |
|
|
|
| 12 |
B |
3097 |
2943 |
16.28 |
|
|
|
| 13 |
B |
1499 |
1425 |
6.21 |
|
|
|
| 14 |
B |
1418 |
1347 |
13.63 |
|
|
|
| 15 |
B |
1276 |
1213 |
6.05 |
|
|
|
| 16 |
B |
1119 |
1063 |
43.19 |
|
|
|
| 17 |
B |
920 |
874 |
35.97 |
|
|
|
| 18 |
B |
497 |
473 |
15.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13624.3 cm
-1
Scaled (by 0.9504) Zero Point Vibrational Energy (zpe) 12948.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.264 |
0.706 |
0.512 |
| C2 |
-0.264 |
-0.706 |
0.512 |
| F3 |
-0.264 |
1.420 |
-0.550 |
| F4 |
0.264 |
-1.420 |
-0.550 |
| H5 |
-0.021 |
1.204 |
1.455 |
| H6 |
1.362 |
0.696 |
0.423 |
| H7 |
0.021 |
-1.204 |
1.455 |
| H8 |
-1.362 |
-0.696 |
0.423 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5067 | 1.3852 | 2.3766 | 1.1034 | 1.1019 | 2.1432 | 2.1491 |
C2 | 1.5067 | | 2.3766 | 1.3852 | 2.1432 | 2.1491 | 1.1034 | 1.1019 | F3 | 1.3852 | 2.3766 | | 2.8891 | 2.0312 | 2.0286 | 3.3147 | 2.5756 | F4 | 2.3766 | 1.3852 | 2.8891 | | 3.3147 | 2.5756 | 2.0312 | 2.0286 | H5 | 1.1034 | 2.1432 | 2.0312 | 3.3147 | | 1.7989 | 2.4083 | 2.5446 | H6 | 1.1019 | 2.1491 | 2.0286 | 2.5756 | 1.7989 | | 2.5446 | 3.0599 | H7 | 2.1432 | 1.1034 | 3.3147 | 2.0312 | 2.4083 | 2.5446 | | 1.7989 | H8 | 2.1491 | 1.1019 | 2.5756 | 2.0286 | 2.5446 | 3.0599 | 1.7989 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.464 |
|
C1 |
C2 |
H7 |
109.418 |
| C1 |
C2 |
H8 |
109.970 |
|
C2 |
C1 |
F3 |
110.464 |
| C2 |
C1 |
H5 |
109.418 |
|
C2 |
C1 |
H6 |
109.970 |
| F3 |
C1 |
H5 |
108.883 |
|
F3 |
C1 |
H6 |
108.769 |
| F4 |
C2 |
H7 |
108.883 |
|
F4 |
C2 |
H8 |
108.769 |
| H5 |
C1 |
H6 |
109.312 |
|
H7 |
C2 |
H8 |
109.312 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
