return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H6O2 (1,2-Ethanediol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-229.716208
Energy at 298.15K-229.723366
HF Energy-228.977176
Nuclear repulsion energy130.758787
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3874 3669 0.00      
2 Ag 3060 2899 0.00      
3 Ag 1569 1486 0.00      
4 Ag 1534 1453 0.00      
5 Ag 1324 1254 0.00      
6 Ag 1125 1066 0.00      
7 Ag 1029 974 0.00      
8 Ag 492 466 0.00      
9 Au 3129 2964 101.53      
10 Au 1258 1191 2.20      
11 Au 857 812 3.75      
12 Au 245 232 284.99      
13 Au 131 124 43.99      
14 Bg 3103 2939 0.00      
15 Bg 1338 1268 0.00      
16 Bg 1188 1125 0.00      
17 Bg 227 215 0.00      
18 Bu 3875 3670 50.33      
19 Bu 3065 2903 103.48      
20 Bu 1578 1494 5.10      
21 Bu 1445 1369 7.38      
22 Bu 1220 1156 132.86      
23 Bu 1112 1053 189.44      
24 Bu 301 285 25.87      

Unscaled Zero Point Vibrational Energy (zpe) 19039.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18032.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.02007 0.13496 0.12483

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.755 0.000
C2 0.000 -0.755 0.000
O3 1.359 -1.163 0.000
O4 -1.359 1.163 0.000
H5 1.386 -2.121 0.000
H6 -1.386 2.121 0.000
H7 -0.533 -1.110 0.890
H8 -0.533 -1.110 -0.890
H9 0.533 1.110 0.890
H10 0.533 1.110 -0.890

Atom - Atom Distances (Å)
  C1 C2 O3 O4 H5 H6 H7 H8 H9 H10
C11.50932.35011.41913.19211.94612.13382.13381.09661.0966
C21.50931.41912.35011.94613.19211.09661.09662.13382.1338
O32.35011.41913.57700.95884.27972.09212.09212.57632.5763
O41.41912.35013.57704.27970.95882.57632.57632.09212.0921
H53.19211.94610.95884.27975.06722.34472.34473.45783.4578
H61.94613.19214.27970.95885.06723.45783.45782.34472.3447
H72.13381.09662.09212.57632.34473.45781.77972.46293.0386
H82.13381.09662.09212.57632.34473.45781.77973.03862.4629
H91.09662.13382.57632.09213.45782.34472.46293.03861.7797
H101.09662.13382.57632.09213.45782.34473.03862.46291.7797

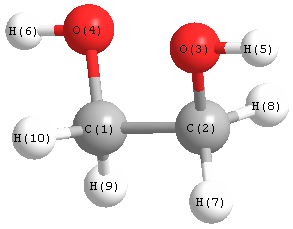
picture of 1,2-Ethanediol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 106.703 C1 C2 H7 108.900
C1 C2 H8 108.900 C1 O4 H6 108.293
C2 C1 O4 106.703 C2 C1 H9 108.900
C2 C1 H10 108.900 C2 O3 H5 108.293
O3 C2 H7 111.892 O3 C2 H8 111.892
O4 C1 H9 111.892 O4 C1 H10 111.892
H7 C2 H8 108.472 H9 C1 H10 108.472
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability