return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H2O3 (Maleic Anhydride)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-378.538446
Energy at 298.15K-378.541666
HF Energy-377.312626
Nuclear repulsion energy274.713223
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3298 3124 0.56      
2 A1 1900 1800 42.66      
3 A1 1642 1555 3.65      
4 A1 1288 1220 159.05      
5 A1 1085 1028 25.38      
6 A1 885 838 6.07      
7 A1 634 601 0.67      
8 A1 406 384 9.74      
9 A2 931 882 0.00      
10 A2 760 720 0.00      
11 A2 239 226 0.00      
12 B1 846 801 79.64      
13 B1 635 602 0.10      
14 B1 161 152 1.85      
15 B2 3277 3104 0.77      
16 B2 1847 1749 455.12      
17 B2 1341 1270 0.51      
18 B2 1077 1020 94.00      
19 B2 923 874 152.25      
20 B2 705 668 38.48      
21 B2 560 530 6.48      

Unscaled Zero Point Vibrational Energy (zpe) 12219.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 11573.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.22852 0.08195 0.06032

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 0.972
C2 0.000 1.128 0.160
C3 0.000 -1.128 0.160
O4 0.000 2.241 0.598
O5 0.000 -2.241 0.598
C6 0.000 0.670 -1.258
C7 0.000 -0.670 -1.258
H8 0.000 1.362 -2.089
H9 0.000 -1.362 -2.089

Atom - Atom Distances (Å)
  O1 C2 C3 O4 O5 C6 C7 H8 H9
O11.38981.38982.27182.27182.32792.32793.35053.3505
C21.38982.25641.19603.39761.49022.28972.26173.3557
C31.38982.25643.39761.19602.28971.49023.35572.2617
O42.27181.19603.39764.48192.43203.45222.82814.4949
O52.27183.39761.19604.48193.45222.43204.49492.8281
C62.32791.49022.28972.43203.45221.33941.08202.1951
C72.32792.28971.49023.45222.43201.33942.19511.0820
H83.35052.26173.35572.82814.49491.08202.19512.7234
H93.35053.35572.26174.49492.82812.19511.08202.7234

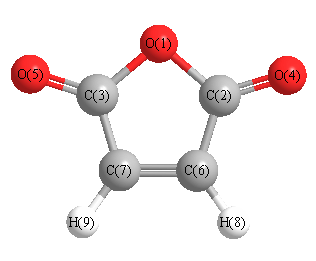
picture of Maleic Anhydride state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 O4 122.768 O1 C2 C6 107.810
O1 C3 O5 122.768 O1 C3 C7 107.810
C2 O1 C3 108.537 C2 C6 C7 107.921
C2 C6 H8 122.318 C3 C7 C6 107.921
C3 C7 H9 122.318 O4 C2 C6 129.421
O5 C3 C7 129.421 C6 C7 H9 129.761
C7 C6 H8 129.761
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability