Jump to
S1C2
S2C1
S2C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -1817.521794 |
| Energy at 298.15K | -1817.523151 |
| Nuclear repulsion energy | 67.216087 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3146 |
2980 |
41.08 |
|
|
|
| 2 |
A1 |
1360 |
1288 |
30.76 |
|
|
|
| 3 |
A1 |
543 |
515 |
24.34 |
|
|
|
| 4 |
B1 |
622 |
589 |
112.06 |
|
|
|
| 5 |
B2 |
3250 |
3078 |
27.19 |
|
|
|
| 6 |
B2 |
555 |
525 |
29.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4737.8 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 4487.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.438 |
| C2 |
0.000 |
0.000 |
-1.494 |
| H3 |
0.000 |
0.909 |
-2.092 |
| H4 |
0.000 |
-0.909 |
-2.092 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.9329 | 2.6890 | 2.6890 |
C2 | 1.9329 | | 1.0878 | 1.0878 | H3 | 2.6890 | 1.0878 | | 1.8173 | H4 | 2.6890 | 1.0878 | 1.8173 | |
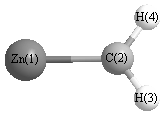 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Zn1 |
C2 |
H3 |
123.349 |
|
Zn1 |
C2 |
H4 |
123.349 |
| H3 |
C2 |
H4 |
113.301 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -1817.521794 |
| Energy at 298.15K | -1817.523151 |
| Nuclear repulsion energy | 67.216087 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -1817.464125 |
| Energy at 298.15K | -1817.465783 |
| Nuclear repulsion energy | 70.907686 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3229 |
3058 |
0.93 |
|
|
|
| 2 |
A1 |
1271 |
1204 |
182.58 |
|
|
|
| 3 |
A1 |
623 |
590 |
4.19 |
|
|
|
| 4 |
B1 |
1269 |
1202 |
896.20 |
|
|
|
| 5 |
B2 |
3433 |
3252 |
2.91 |
|
|
|
| 6 |
B2 |
736 |
697 |
50.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5280.5 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 5001.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.409 |
| C2 |
0.000 |
0.000 |
-1.428 |
| H3 |
0.000 |
0.990 |
-1.847 |
| H4 |
0.000 |
-0.990 |
-1.847 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.8372 | 2.4638 | 2.4638 |
C2 | 1.8372 | | 1.0748 | 1.0748 | H3 | 2.4638 | 1.0748 | | 1.9795 | H4 | 2.4638 | 1.0748 | 1.9795 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -1817.503246 |
| Energy at 298.15K | -1817.504773 |
| Nuclear repulsion energy | 68.508504 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2920 |
2765 |
108.27 |
|
|
|
| 2 |
A' |
1455 |
1378 |
4.78 |
|
|
|
| 3 |
A' |
826 |
782 |
165.39 |
|
|
|
| 4 |
A' |
507 |
481 |
1.07 |
|
|
|
| 5 |
A" |
2991 |
2833 |
128.66 |
|
|
|
| 6 |
A" |
759 |
719 |
14.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4729.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 4478.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
-0.034 |
-0.419 |
0.000 |
| C2 |
-0.034 |
1.507 |
0.000 |
| H3 |
0.613 |
1.762 |
0.868 |
| H4 |
0.613 |
1.762 |
-0.868 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.9260 | 2.4352 | 2.4352 |
C2 | 1.9260 | | 1.1123 | 1.1123 | H3 | 2.4352 | 1.1123 | | 1.7366 | H4 | 2.4352 | 1.1123 | 1.7366 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
