return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

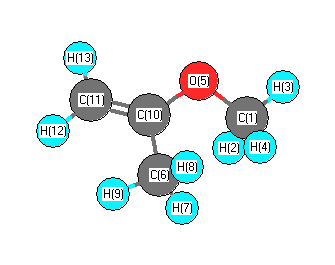
All results from a given calculation for CH2C(CH3)OCH3 (1-Propene, 2-methoxy-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-231.829415
Energy at 298.15K-231.838042
HF Energy-230.998189
Nuclear repulsion energy180.245375
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3315 3139 5.93      
2 A 3209 3039 0.91      
3 A 3204 3035 20.27      
4 A 3193 3024 15.25      
5 A 3158 2991 10.94      
6 A 3143 2977 45.51      
7 A 3082 2919 17.13      
8 A 3062 2900 46.98      
9 A 1728 1636 82.36      
10 A 1553 1471 7.63      
11 A 1545 1463 10.78      
12 A 1529 1449 12.90      
13 A 1519 1438 8.00      
14 A 1516 1436 6.91      
15 A 1457 1380 18.08      
16 A 1431 1355 19.31      
17 A 1330 1260 151.01      
18 A 1233 1167 0.42      
19 A 1200 1136 3.40      
20 A 1141 1081 99.15      
21 A 1080 1022 1.36      
22 A 1038 983 3.82      
23 A 953 903 5.12      
24 A 832 788 16.58      
25 A 816 773 53.34      
26 A 757 717 8.15      
27 A 545 516 6.26      
28 A 477 451 4.66      
29 A 421 398 1.03      
30 A 324 307 1.15      
31 A 260 246 1.56      
32 A 205 194 1.09      
33 A 93 88 8.01      

Unscaled Zero Point Vibrational Energy (zpe) 25172.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23840.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.26801 0.14439 0.10183

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.800 -0.071 0.228
H2 1.700 -0.098 1.319
H3 2.649 -0.683 -0.069
H4 1.975 0.959 -0.096
O5 0.666 -0.647 -0.405
C6 -0.628 1.411 -0.041
H7 -0.062 1.847 0.787
H8 -0.228 1.825 -0.971
H9 -1.666 1.726 0.062
C10 -0.541 -0.087 -0.058
C11 -1.579 -0.905 0.161
H12 -2.569 -0.509 0.345
H13 -1.442 -1.978 0.124

Atom - Atom Distances (Å)
  C1 H2 H3 H4 O5 C6 H7 H8 H9 C10 C11 H12 H13
C11.09521.08831.09361.42052.85742.73063.02433.90812.35863.48154.39313.7630
H21.09521.78041.78682.08313.08932.67723.55754.02952.62993.56994.39803.8512
H31.08831.78041.77532.01173.88953.80573.92224.94443.24564.24095.23824.2959
H41.09361.78681.77532.09392.64222.39062.52353.72432.72484.02144.79584.5110
O51.42052.08312.01172.09392.45802.85762.68873.35981.37492.33023.32442.5493
C62.85743.08933.88952.64222.45801.09301.09391.08991.50122.51222.75833.4900
H72.73062.67723.80572.39062.85761.09301.76561.76462.16423.20413.46904.1199
H83.02433.55753.92222.52352.68871.09391.76561.77362.14223.24983.55904.1402
H93.90814.02954.94443.72433.35981.08991.76461.77362.13772.63452.42793.7119
C102.35862.62993.24562.72481.37491.50122.16422.14222.13771.33972.11082.1027
C113.48153.56994.24094.02142.33022.51223.20413.24982.63451.33971.08191.0828
H124.39314.39805.23824.79583.32442.75833.46903.55902.42792.11081.08191.8648
H133.76303.85124.29594.51102.54933.49004.11994.14023.71192.10271.08281.8648

picture of 1-Propene, 2-methoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O5 C10 115.070 H2 C1 H3 109.252
H2 C1 H4 109.439 H2 C1 O5 111.139
H3 C1 H4 108.906 H3 C1 O5 105.858
H4 C1 O5 112.137 O5 C10 C6 117.368
O5 C10 C11 118.269 C6 C10 C11 124.230
H7 C6 H8 107.682 H7 C6 H9 107.869
H7 C6 C10 112.110 H8 C6 H9 108.619
H8 C6 C10 110.285 H9 C6 C10 110.163
C10 C11 H12 120.935 C10 C11 H13 120.078
H12 C11 H13 118.960
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability