return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,3-Hexadiene, (Z)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.946891
Energy at 298.15K-233.956686
HF Energy-233.025073
Nuclear repulsion energy216.357216
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3284 3110 17.22      
2 A' 3261 3088 1.04      
3 A' 3180 3012 8.54      
4 A' 3176 3008 28.69      
5 A' 3168 3000 35.43      
6 A' 3152 2986 3.21      
7 A' 3086 2923 18.21      
8 A' 3056 2895 25.63      
9 A' 1725 1634 4.85      
10 A' 1652 1565 1.06      
11 A' 1555 1473 9.79      
12 A' 1513 1433 6.78      
13 A' 1493 1414 13.37      
14 A' 1451 1374 5.96      
15 A' 1420 1345 5.03      
16 A' 1415 1340 0.16      
17 A' 1344 1273 0.50      
18 A' 1326 1255 6.30      
19 A' 1197 1134 0.66      
20 A' 1100 1042 4.54      
21 A' 1068 1012 1.77      
22 A' 951 901 3.00      
23 A' 830 786 3.93      
24 A' 722 684 1.75      
25 A' 427 405 0.33      
26 A' 278 263 2.09      
27 A' 199 189 0.34      
28 A" 3186 3017 23.43      
29 A" 3090 2927 15.27      
30 A" 1547 1465 9.76      
31 A" 1333 1262 0.29      
32 A" 1128 1068 3.85      
33 A" 1012 959 21.28      
34 A" 953 903 3.30      
35 A" 884 837 43.80      
36 A" 832 788 3.70      
37 A" 758 718 5.15      
38 A" 616 583 33.10      
39 A" 392 372 0.20      
40 A" 305 289 0.06      
41 A" 92 87 0.35      
42 A" 55 52 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 31605.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29933.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.23133 0.06727 0.05314

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.056 1.268 0.000
C2 -1.231 0.554 0.000
H3 0.024 2.201 0.000
C4 0.000 1.111 0.000
H5 1.405 -0.589 0.000
C6 1.322 0.490 0.000
H7 3.430 0.752 0.000
H8 2.425 2.307 0.000
C9 2.452 1.221 0.000
H10 -2.371 -0.989 0.869
H11 -2.371 -0.989 -0.869
C12 -1.710 -0.873 0.000
H13 -1.214 -2.969 0.000
H14 -0.057 -1.980 -0.887
H15 -0.057 -1.980 0.887
C16 -0.692 -2.008 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 C6 H7 H8 C9 H10 H11 C12 H13 H14 H15 C16
H11.09112.27992.06203.92803.46615.51024.59994.50872.43952.43952.16854.31993.91573.91573.5488
C21.09112.07121.35122.87352.55374.66544.05473.74352.10642.10641.50483.52282.93012.93012.6182
H32.27992.07121.09093.11352.14733.70152.40322.61884.08344.08343.52945.31624.27494.27494.2700
C42.06201.35121.09092.20551.46003.44872.70402.45493.28473.28472.61884.25623.21583.21583.1948
H53.92802.87353.11352.20551.08282.42863.07042.09123.89573.89573.12803.53822.20392.20392.5322
C63.46612.55372.14731.46001.08282.12452.12541.34624.07234.07233.32404.28862.96452.96453.2091
H75.51024.66543.70153.44872.42862.12451.85161.08436.11906.11905.39065.95004.51724.51724.9608
H84.59994.05472.40322.70403.07042.12541.85161.08645.88445.88445.21626.40875.03215.03215.3232
C94.50873.74352.61882.45492.09121.34621.08431.08645.37685.37684.65935.56704.16264.16264.5073
H102.43952.10644.08343.28473.89574.07236.11905.88445.37681.73771.09812.45233.06932.51762.1476
H112.43952.10644.08343.28473.89574.07236.11905.88445.37681.73771.09812.45232.51763.06932.1476
C122.16851.50483.52942.61883.12803.32405.39065.21624.65931.09811.09812.15422.17842.17841.5249
H134.31993.52285.31624.25623.53824.28865.95006.40875.56702.45232.45232.15421.76131.76131.0929
H143.91572.93014.27493.21582.20392.96454.51725.03214.16263.06932.51762.17841.76131.77351.0913
H153.91572.93014.27493.21582.20392.96454.51725.03214.16262.51763.06932.17841.76131.77351.0913
C163.54882.61824.27003.19482.53223.20914.96085.32324.50732.14762.14761.52491.09291.09131.0913

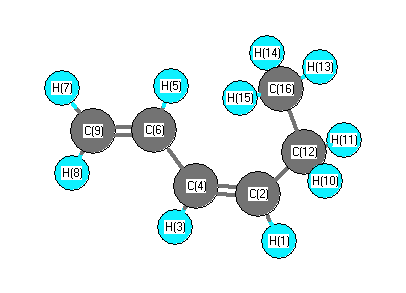
picture of 1,3-Hexadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 114.776 H1 C2 C12 112.334
C2 C4 H3 115.602 C2 C4 C6 130.521
C2 C12 H10 107.007 C2 C12 H11 107.007
C2 C12 C16 119.572 H3 C4 C6 113.877
C4 C2 C12 132.890 C4 C6 H5 119.563
C4 C6 C9 121.993 H5 C6 C9 118.444
C6 C9 H7 121.498 C6 C9 H8 121.417
H7 C9 H8 117.085 H10 C12 H11 104.599
H10 C12 C16 108.829 H11 C12 C16 108.829
C12 C16 H13 109.650 C12 C16 H14 111.675
C12 C16 H15 111.675 H13 C16 H14 107.485
H13 C16 H15 107.485 H14 C16 H15 108.695
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability