return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6N2 (3-Aminopropionitrile)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-226.862328
Energy at 298.15K-226.869430
HF Energy-226.025098
Nuclear repulsion energy154.464032
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3559 3371 0.29      
2 A' 3112 2948 24.50      
3 A' 3094 2931 3.31      
4 A' 2205 2089 0.02      
5 A' 1743 1651 37.48      
6 A' 1532 1451 6.90      
7 A' 1512 1432 3.45      
8 A' 1418 1343 13.21      
9 A' 1321 1251 0.66      
10 A' 1150 1089 7.92      
11 A' 1055 999 54.07      
12 A' 967 916 11.71      
13 A' 878 831 221.73      
14 A' 522 494 1.63      
15 A' 382 362 7.68      
16 A' 162 153 6.45      
17 A" 3659 3465 0.91      
18 A" 3174 3006 19.66      
19 A" 3145 2979 0.14      
20 A" 1429 1353 0.32      
21 A" 1343 1272 0.57      
22 A" 1189 1126 0.05      
23 A" 995 942 0.59      
24 A" 765 725 1.50      
25 A" 379 359 0.24      
26 A" 313 297 53.19      
27 A" 110 104 1.23      

Unscaled Zero Point Vibrational Energy (zpe) 20555.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19468.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.82546 0.07596 0.07222

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 2.539 -0.035 0.819
H2 2.539 -0.035 -0.819
N3 2.227 -0.542 0.000
H4 0.490 -1.265 0.877
H5 0.490 -1.265 -0.877
C6 0.781 -0.682 0.000
H7 0.270 1.239 0.881
H8 0.270 1.239 -0.881
C9 0.000 0.649 0.000
N10 -2.599 0.216 0.000
C11 -1.446 0.434 0.000

Atom - Atom Distances (Å)
  H1 H2 N3 H4 H5 C6 H7 H8 C9 N10 C11
H11.63741.01202.39002.93012.04422.60213.10752.75365.20844.0948
H21.63741.01202.93012.39002.04423.10752.60212.75365.20844.0948
N31.01201.01202.07552.07551.45252.78792.78792.52484.88463.7999
H42.39002.93012.07551.75491.09262.51353.06772.16183.53652.7214
H52.93012.39002.07551.75491.09263.06772.51352.16183.53652.7214
C62.04422.04421.45251.09261.09262.17442.17441.54343.49722.4910
H72.60213.10752.78792.51353.06772.17441.76241.09443.17122.0907
H83.10752.60212.78793.06772.51352.17441.76241.09443.17122.0907
C92.75362.75362.52482.16182.16181.54341.09441.09442.63481.4620
N105.20845.20844.88463.53653.53653.49723.17123.17122.63481.1734
C114.09484.09483.79992.72142.72142.49102.09072.09071.46201.1734

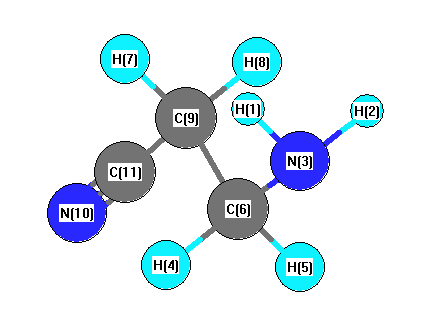
picture of 3-Aminopropionitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 N3 H2 107.989 H1 N3 C6 110.818
H2 N3 C6 110.818 N3 C6 H4 108.446
N3 C6 H5 108.446 N3 C6 C9 114.828
H4 C6 H5 106.846 H4 C6 C9 108.992
H5 C6 C9 108.992 C6 C9 H7 109.873
C6 C9 H8 109.873 C6 C9 C11 111.937
H7 C9 H8 107.260 H7 C9 C11 108.888
H8 C9 C11 108.888 C9 C11 N10 177.750
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability