return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (cis-2-butenal)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-230.666953
Energy at 298.15K-230.672770
Nuclear repulsion energy155.107356
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3204 3034 16.64      
2 A' 3196 3027 3.11      
3 A' 3175 3007 9.99      
4 A' 3074 2912 12.71      
5 A' 2973 2816 163.73      
6 A' 1767 1674 55.82      
7 A' 1713 1623 122.37      
8 A' 1527 1446 23.71      
9 A' 1453 1376 6.22      
10 A' 1446 1369 3.09      
11 A' 1341 1270 18.89      
12 A' 1338 1267 6.11      
13 A' 1163 1102 5.82      
14 A' 1046 990 11.63      
15 A' 912 864 30.00      
16 A' 749 709 37.13      
17 A' 399 378 3.05      
18 A' 210 199 4.81      
19 A" 3147 2981 11.20      
20 A" 1516 1436 10.88      
21 A" 1089 1031 1.00      
22 A" 1027 973 0.13      
23 A" 1004 951 43.32      
24 A" 759 719 0.23      
25 A" 244 231 6.27      
26 A" 199 188 0.21      
27 A" 142 135 5.55      

Unscaled Zero Point Vibrational Energy (zpe) 19906.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18853.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.63479 0.08719 0.07776

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.475 0.555 0.000
C2 0.000 0.699 0.000
C3 0.804 -0.377 0.000
C4 2.297 -0.335 0.000
O5 -2.064 -0.512 0.000
H6 -2.047 1.504 0.000
H7 0.408 1.708 0.000
H8 0.323 -1.355 0.000
H9 2.673 0.690 0.000
H10 2.696 -0.852 0.878
H11 2.696 -0.852 -0.878

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.48172.46213.87491.21801.10872.20812.62294.14944.48854.4885
C21.48171.34372.51892.39252.19981.08832.07952.67263.23223.2322
C32.46211.34371.49302.87103.41622.12241.08972.15132.13912.1391
C43.87492.51891.49304.36394.71732.78222.22141.09141.09421.0942
O51.21802.39252.87104.36392.01573.32212.53164.88604.85214.8521
H61.10872.19983.41624.71732.01572.46393.71404.78955.36875.3687
H72.20811.08832.12242.78223.32212.46393.06422.48273.54393.5439
H82.62292.07951.08972.22142.53163.71403.06423.11432.57962.5796
H94.14942.67262.15131.09144.88604.78952.48273.11431.77411.7741
H104.48853.23222.13911.09424.85215.36873.54392.57961.77411.7554
H114.48853.23222.13911.09424.85215.36873.54392.57961.77411.7554

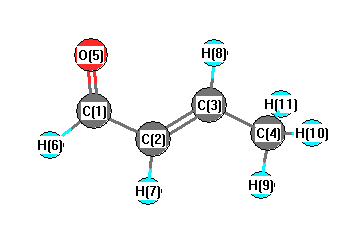
picture of cis-2-butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.174 C1 C2 H7 117.630
C2 C1 O5 124.510 C2 C1 H6 115.497
C2 C3 C4 125.153 C2 C3 H8 117.040
C3 C2 H7 121.196 C3 C4 H9 111.756
C3 C4 H10 110.593 C3 C4 H11 110.593
C4 C3 H8 117.807 O5 C1 H6 119.993
H9 C4 H10 108.532 H9 C4 H11 108.532
H10 C4 H11 106.668
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-311G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.230      
2 C -0.407      
3 C -0.088      
4 C -0.639      
5 O -0.425      
6 H 0.181      
7 H 0.215      
8 H 0.250      
9 H 0.224      
10 H 0.230      
11 H 0.230      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000