return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4 ((Z)-Hexa-1,5-diyne-3-ene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-230.316618
Energy at 298.15K-230.317140
HF Energy-229.421257
Nuclear repulsion energy165.846849
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3489 3305 63.99      
2 A1 3217 3047 9.73      
3 A1 2116 2004 7.06      
4 A1 1630 1544 2.09      
5 A1 1252 1186 0.37      
6 A1 902 855 4.99      
7 A1 595 563 25.60      
8 A1 425 403 3.14      
9 A1 100 94 0.17      
10 A2 899 851 0.00      
11 A2 568 538 0.00      
12 A2 324 307 0.00      
13 A2 166 158 0.00      
14 B1 755 715 18.89      
15 B1 505 478 120.53      
16 B1 274 259 0.69      
17 B2 3489 3304 48.02      
18 B2 3200 3030 2.10      
19 B2 2132 2019 3.97      
20 B2 1426 1350 1.26      
21 B2 1043 988 8.34      
22 B2 728 690 10.15      
23 B2 580 549 55.74      
24 B2 228 216 3.51      

Unscaled Zero Point Vibrational Energy (zpe) 15021.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 14227.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.22893 0.08707 0.06308

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.678 1.115
C2 0.000 -0.678 1.115
C3 0.000 1.468 -0.066
C4 0.000 -1.468 -0.066
H5 0.000 1.199 2.069
H6 0.000 -1.199 2.069
C7 0.000 2.166 -1.069
C8 0.000 -2.166 -1.069
H9 0.000 2.765 -1.949
H10 0.000 -2.765 -1.949

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 C7 C8 H9 H10
C11.35661.42082.44981.08752.10642.64203.58553.70714.6092
C21.35662.44981.42082.10641.08753.58552.64204.60923.7071
C31.42082.44982.93562.15253.41701.22133.76912.28634.6326
C42.44981.42082.93563.41702.15253.76911.22134.63262.2863
H51.08752.10642.15253.41702.39893.28344.60124.31265.6449
H62.10641.08753.41702.15252.39894.60123.28345.64494.3126
C72.64203.58551.22133.76913.28344.60124.33121.06515.0086
C83.58552.64203.76911.22134.60123.28344.33125.00861.0651
H93.70714.60922.28634.63264.31265.64491.06515.00865.5301
H104.60923.70714.63262.28635.64494.31265.00861.06515.5301

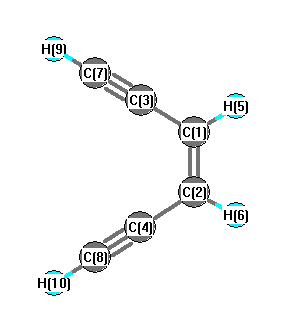
picture of (Z)-Hexa-1,5-diyne-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 123.755 C1 C2 H6 118.635
C1 C3 C7 178.909 C2 C1 C3 123.755
C2 C1 H5 118.635 C2 C4 C8 178.909
C3 C1 H5 117.610 C3 C7 H9 179.404
C4 C2 H6 117.610 C4 C8 H10 179.404
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability