return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (1-Butanethiol, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-594.936090
Energy at 298.15K-594.949094
HF Energy-593.891765
Nuclear repulsion energy307.452726
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3177 3009 27.60      
2 A 3174 3006 17.18      
3 A 3166 2999 33.71      
4 A 3163 2996 31.36      
5 A 3158 2991 20.46      
6 A 3117 2952 17.01      
7 A 3089 2926 27.75      
8 A 3079 2916 22.93      
9 A 3074 2912 32.05      
10 A 3069 2906 5.71      
11 A 3063 2901 9.80      
12 A 2685 2543 11.63      
13 A 1551 1469 12.55      
14 A 1544 1463 14.64      
15 A 1539 1458 7.48      
16 A 1534 1453 2.00      
17 A 1524 1443 1.62      
18 A 1513 1433 8.24      
19 A 1451 1374 7.99      
20 A 1450 1373 12.80      
21 A 1418 1343 1.93      
22 A 1406 1332 2.68      
23 A 1356 1285 0.20      
24 A 1350 1279 16.20      
25 A 1311 1242 3.08      
26 A 1265 1198 7.09      
27 A 1212 1148 0.41      
28 A 1177 1115 3.27      
29 A 1131 1071 3.37      
30 A 1084 1026 0.88      
31 A 1058 1002 2.58      
32 A 1003 950 5.90      
33 A 971 920 2.22      
34 A 925 876 4.08      
35 A 901 853 1.96      
36 A 818 775 3.72      
37 A 815 772 2.58      
38 A 748 709 1.29      
39 A 466 442 0.20      
40 A 418 395 0.11      
41 A 395 374 1.11      
42 A 269 255 0.41      
43 A 251 238 0.17      
44 A 214 203 0.19      
45 A 207 196 1.23      
46 A 146 138 24.34      
47 A 111 105 0.34      
48 A 77 73 5.89      

Unscaled Zero Point Vibrational Energy (zpe) 35812.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 33917.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.15203 0.05116 0.04069

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.042 1.879 0.037
H2 -1.038 2.084 -0.360
H3 -0.075 2.044 1.119
H4 0.644 2.612 -0.396
S5 -2.191 -0.397 -0.132
H6 -2.618 -1.579 0.345
C7 -0.457 -0.588 0.402
H8 -0.388 -0.476 1.489
H9 -0.123 -1.590 0.129
C10 0.427 0.458 -0.271
H11 0.368 0.296 -1.356
C12 2.494 -1.082 -0.159
H13 3.573 -1.077 0.013
H14 2.072 -1.872 0.465
H15 2.325 -1.352 -1.205
C16 1.890 0.285 0.155
H17 2.477 1.061 -0.350
H18 1.972 0.492 1.230

Atom - Atom Distances (Å)
  C1 H2 H3 H4 S5 H6 C7 H8 H9 C10 H11 C12 H13 H14 H15 C16 H17 H18
C11.09171.09481.09333.13494.32302.52852.78873.47111.52772.14813.90364.66984.32754.19342.50752.67692.7209
H21.09171.76581.76392.74544.05082.83893.22473.81732.19052.48304.74755.60285.09964.88153.47483.66113.7581
H31.09481.76581.77003.46444.49362.75452.56643.76632.16713.06184.24244.92574.51344.76312.80713.10372.5705
H41.09331.76391.77004.14285.36233.47683.76214.30302.16772.52174.13744.72724.78384.38032.69562.40112.9827
S53.13492.74543.46444.14281.34391.82452.42592.40122.75812.92074.73475.80574.55024.73914.14734.89524.4693
H64.32304.05084.49365.36231.34392.37842.73862.50453.71563.91545.16096.22054.70095.18594.88195.78035.1129
C72.52852.83892.75453.47681.82452.37841.09501.09041.52642.13403.04384.07782.83683.30232.51593.44832.7841
H82.78873.22472.56643.76212.42592.73861.09501.77772.15313.04373.37424.26913.00793.92242.74713.73512.5639
H93.47113.81733.76634.30302.40122.50451.09041.77772.15822.45022.68133.73342.23862.79832.75093.74363.1522
C101.52772.19052.16712.16772.75813.71561.52642.15312.15821.09882.57993.51172.94602.78401.53302.13772.1538
H112.14812.48303.06182.52172.92073.91542.13403.04372.45021.09882.80203.74563.30482.56272.14432.45823.0490
C123.90364.74754.24244.13744.73475.16093.04383.37422.68132.57992.80201.09271.09181.09391.52712.15112.1628
H134.66985.60284.92574.72725.80576.22054.07784.26913.73343.51173.74561.09271.75801.76562.16972.42922.5503
H144.32755.09964.51344.78384.55024.70092.83683.00792.23862.94603.30481.09181.75801.76782.18733.07102.4873
H154.19344.88154.76314.38034.73915.18593.30233.92242.79832.78402.56271.09391.76561.76782.17222.56383.0746
C162.50753.47482.80712.69564.14734.88192.51592.74712.75091.53302.14431.52712.16972.18732.17221.09601.0976
H172.67693.66113.10372.40114.89525.78033.44833.73513.74362.13772.45822.15112.42923.07102.56381.09601.7532
H182.72093.75812.57052.98274.46935.11292.78412.56393.15222.15383.04902.16282.55032.48733.07461.09761.7532

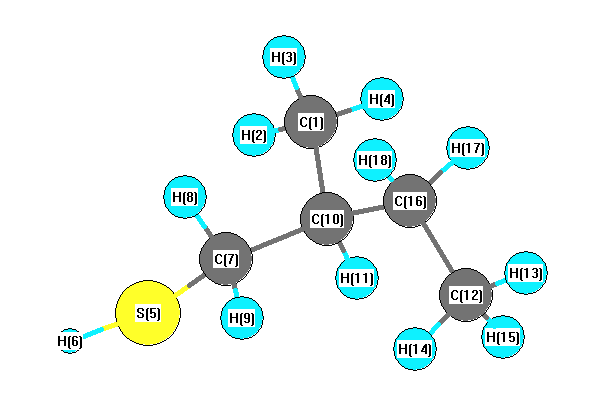
picture of 1-Butanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C10 C7 111.767 C1 C10 H11 108.641
C1 C10 C16 110.023 H2 C1 H3 107.727
H2 C1 H4 107.668 H2 C1 C10 112.430
H3 C1 H4 107.987 H3 C1 C10 110.362
H4 C1 C10 110.505 S5 C7 H8 109.859
S5 C7 H9 108.278 S5 C7 C10 110.479
H6 S5 C7 96.110 C7 C10 H11 107.644
C7 C10 C16 110.647 H8 C7 H9 108.863
H8 C7 C10 109.332 H9 C7 C10 110.003
C10 C16 C12 114.933 C10 C16 H17 107.647
C10 C16 H18 108.793 H11 C10 C16 107.994
C12 C16 H17 109.079 C12 C16 H18 109.896
H13 C12 H14 107.180 H13 C12 H15 107.698
H13 C12 C16 110.738 H14 C12 H15 107.960
H14 C12 C16 112.210 H15 C12 C16 110.861
H17 C16 H18 106.108
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability